


Introduction
In natural ecosystems, several research endeavors have revealed that petroleum hydrocarbons undergo biodegradation, primarily facilitated by diverse communities of both bacteria and fungi [1-6]. Fungi, as eukaryotic organisms, stand out as highly proficient entities for breaking down oil hydrocarbon compounds [7]. A multitude of studies have uncovered a diverse array of fungal species that possess the ability to make crude oil their main source of energy and carbon. Among these species, Trichoderma, Mortierella, Aspergillus spp., Alternaria, Talaromyces, Cephalosporium, Penicillium, Geotrichum, Fusarium, and Cladosporium. Notably, strains of Aspergillus and Penicillium that come from environments contaminated with oil display significant promise in effectively decomposing drilling wastes and tackling oil spill issues [8, 9].
The importance of fungal isolation and identification from diverse environmental sources remains evident [8]. This practice is crucial for expanding our understanding and recognition of additional species, refining scientific classifications, assessing their ecological impacts, and providing strains that can be harnessed for ecological restoration, biological control, and industrial applications [10]. Conversely, intentional introductions of fungi with the capability to degrade or utilize hydrocarbons have probably been infrequent [11]. The advancement of high-throughput sequencing (HTS) technologies has significantly enhanced our ability to identify fungi and uncover their ecological roles in a wide range of ecosystems. This is achieved through the reanalysis of existing datasets and the application of these advanced sequencing techniques. An investigation has shown that operational taxonomic units (OTUs) are more effective than amplified sequence variants (ASVs) when it comes to capturing the full spectrum of fungal diversity, a conclusion that is especially pronounced with longer genetic markers. Furthermore, when it comes to the taxonomic classification of fungi and other eukaryotes, the analysis of the entire ITS region, as opposed to just the ITS2 subregion, provides more precise results [12].
The molecular identification of hydrocarbon-degrading fungi (HDF) at the species level has been a topic of extensive discussion and continues to be the focus of numerous ongoing research studies [13]. It has been highlighted that molecular identification methods reliant on the extraction of total fungal DNA offer a distinct barcode, enabling the precise determination and identification of various fungal isolates at the species level. This molecular identification approach has evolved into an indispensable tool for mycologists engaged in the study of fungal taxonomy, molecular evolution, population genetics, and interactions between fungi and plants [14]. The molecular identification of fungi can be significantly enhanced by employing the sequencing of a PCR-amplified segment of the 18S rRNA genes using universal primers designed for fungal species [15]. Efforts to isolate and identify robust indigenous fungal strains capable of degrading hydrocarbons represent a commendable endeavor, especially when considering the restoration of hydrocarbon-contaminated soils through bioremediation techniques [16]. Therefore, this study is focused on the isolation and precise identification of hydrocarbon-degrading fungal isolates, verified at the species level through the application of molecular strategies. These isolates are obtained from oil-laden soils found at various geographical locations within the Kurdistan Region of Iraq (KRI) oilfields.
The significance of this study is underscored by its proactive approach into tackling the environmental and economic issues stemming from oil pollution in the KRI. The comprehensive identification and comprehension of hydrocarbon-degrading fungi across diverse regions within the KRI oilfields are pivotal for the successful implementation of bioremediation initiatives. Furthermore, the application of molecular techniques amplifies our capacity to oversee and safeguard these ecosystems, rendering this research exceptionally important for advancing environmental preservation and sustainable development in the region.
Experimental
Sample Collection
In this study, soil samples heavily contaminated with oil, referred to as oil-laden soils, were collected from a total of 41 drilling waste pits. These pits were dispersed across five significant oil and gas fields, each situated in diverse geographical regions within the KRI (Fig. 1). To obtain these samples, a small hand auger was employed. For each sample, approximately 100 grams of oil-laden soil were extracted, with varying depths ranging from surface level to beyond 15 centimeters. A representative sample was created by combining 3 -4 individual samples collected from an area covering several square meters. These representative samples were then carefully placed into sterile nylon bags.
Isolation, Macroscopic and Microscopic Identification of Fungi
The soil enrichment technique using Potato Dextrose Agar (PDA) was used for the isolation of fungi. Then pure isolates were tested for their ability to grow on Bushnell Haas Media (BHS), composed of K2HPO4 (1 g/l), MgSO4.7H2O (0.2 g/l), KH2PO4 (1 g/l), CaCl2 (0.02 g/l), NaNO3 (1 g/l), and FeCl2 (0.05 g/l) at 30ºC for one week. About 0.2 g of chloramphenicol was added to culture media as an antibiotic to prevent bacterial growth. The plates were incubated for 5-7 days at 23±2ºC using a Gallenkamp 210 refrigerated incubator, and then counting of the plates was carried out [13-14]. The visual characteristics of the fungal growth on the culture plates were examined to assess their gross morphology. This examination encompassed various morphological attributes such as color, shape, size, hyphal structures, conidia, conidiophores, and the arrangement of spores. For reference and classification, the Manual of Fungi Atlas served as the guide for this morphological analysis [17]. The confirmation of fungal identities involved a dual approach, combining cultural techniques with the cross-referencing of established reference text keys that contained authenticated representative samples [18]. The presence and frequency (expressed as a percentage) of each fungus were determined by applying the provided formula [19].
Molecular Identification
Approximately 0.5 grams of fungal hyphae were extracted from a tube containing pure fungal isolates. These hyphae were subjected to specific treatments. Firstly, they were placed in 100 μl of a lyticase solution and incubated at a temperature of 30ºC for 60 minutes. To break down proteins within the crude sample, 20 μl of proteinase K was introduced, and the mixture was then incubated at 55ºC for 90 minutes. Subsequently, the sample was further incubated for two hours at 65ºC. Finally, a portion of about 10 μl from these treated samples was employed for the polymerase chain reaction (PCR) amplification [13]. In the PCR process, 25 μl reactions were prepared, utilizing universal primers ITS1 (5´-TCC GTA GGT GAA CCT TGC GG-3´) and ITS2 (5´-GCT GCG TTC TTC ATC GAT GC-3´) [13- 16]. The M primers included 1.5 mM MgCl2, 10 μM dNTPs, and 1x buffer. The PCR was programmed for 35 cycles with the following amplification conditions: initial denaturation at 94ºC for 1 minute, annealing at 55.5ºC for 2 minutes, extension at 72ºC for 2 minutes, final extension at 72ºC for 10 minutes, and finally, a cooling step at 4ºC at the end of the last cycle. To verify the purity of DNA, electrophoresis was conducted using a 2% agarose gel, and quantification was carried out using the Quant-iTTM HS ds-DNA assay kit (Invitrogen, Paisley, United Kingdom) method, coupled with the QuBitTM Fluorometer [20].
PCR Product Sequencing and BLAST Analysis
To analyze the PCR product sequences, the highly effective fungal strains, which were amplified using the ITS1 and ITS2 primers, were compared with similar sequences available in the GenBank database, specifically those related to Aspergillus and Penicillium commune. These sequences were subjected to a homology search using the Basic Local Alignment Search Tool (BLAST) software provided by the National Centre for Biotechnology Information (NCBI). By assessing the degree of homology, the fungi were identified, and a phylogenetic tree was constructed using the NCBI platform. The evolutionary distances between these sequences were computed using the Maximum Likelihood approach, and these distances were expressed in terms of the number of base substitutions per site. This entire process of evolutionary analysis and phylogenetic tree construction was carried out using MEGA Version-4, following the methodology outlined in the provided reference [20, 21].
Results and Discussion
Isolation and Identification of Fungi
The study investigated 68 fungal isolates obtained from diverse geographical locations within the KRI oilfields (Fig. 1). The findings revealed a significant predominance of hydrocarbon-degrading fungi (HDF) in comparison to the total heterogeneous fungi (THF) within oil-contaminated soil samples. HDF constituted a substantial 92% of the isolates, while THF accounted for only 8% (Fig. 2a). This high occurrence of HDF bears ecological implications, signifying the adaptability of these fungi to the challenging environment of oil-contaminated soil, likely due to their ability to utilize hydrocarbons as a carbon source for growth [5]. This ecological adaptation positions them as crucial contributors to the natural mitigation of hydrocarbon pollutants in the environment [15]. Furthermore, the prevalence of specific HDF genera is noteworthy, with Aspergillus spp. comprising 63% and Penicillium sp. 34% of HDF isolates (Fig. 2b). However, Aspergillus spp. showed the maximum ability to utilize crude oil, giving the highest percent degradation of 91.76%, followed by Penicillium sp., indicating an 81.87% degradation calculated after two months of bioremediation. These genera are recognized for their proficiency in degrading hydrocarbons and have been extensively studied in bioremediation initiatives [15, 16]. Consequently, their abundance in the oil-laden soil samples underscores their potential for biotechnological applications in bioremediation and environmental clean-up [22-24]. No previous reports are available on hydrocarbon-degrading fungi from Iraq. But in a previous study, the reference [4] has reported the presence of two crude oil-degrading fungi strains from the Rumaila oil field in Iraq, namely Penicillium sp. RMA1 and RMA2, which are known to utilize a variety of hydrocarbons as the sole source of carbon and energy.
Fig. 1.
Location map showing tentative locations of the 41 drilling waste pits sampled for oil-laden soils.
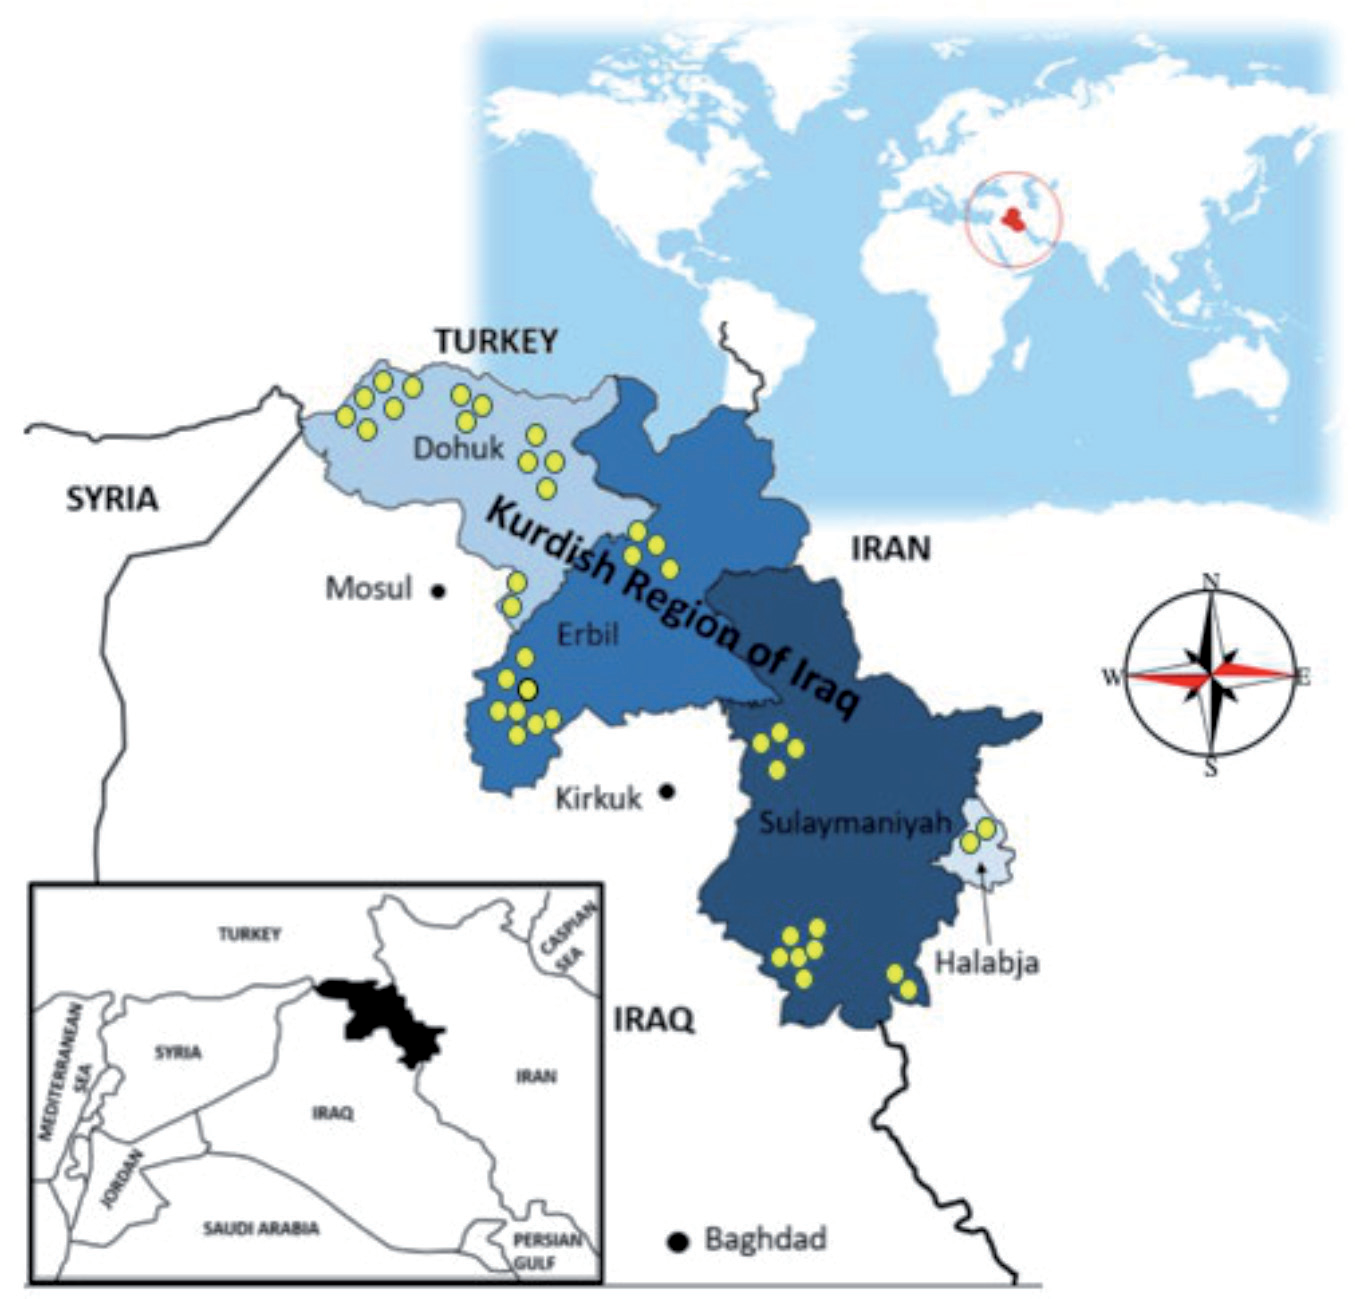
Fig. 2.
Frequency of occurrence % a), along mean percentage occurrence b) of HDF and THF isolates in the oil-laden soils.
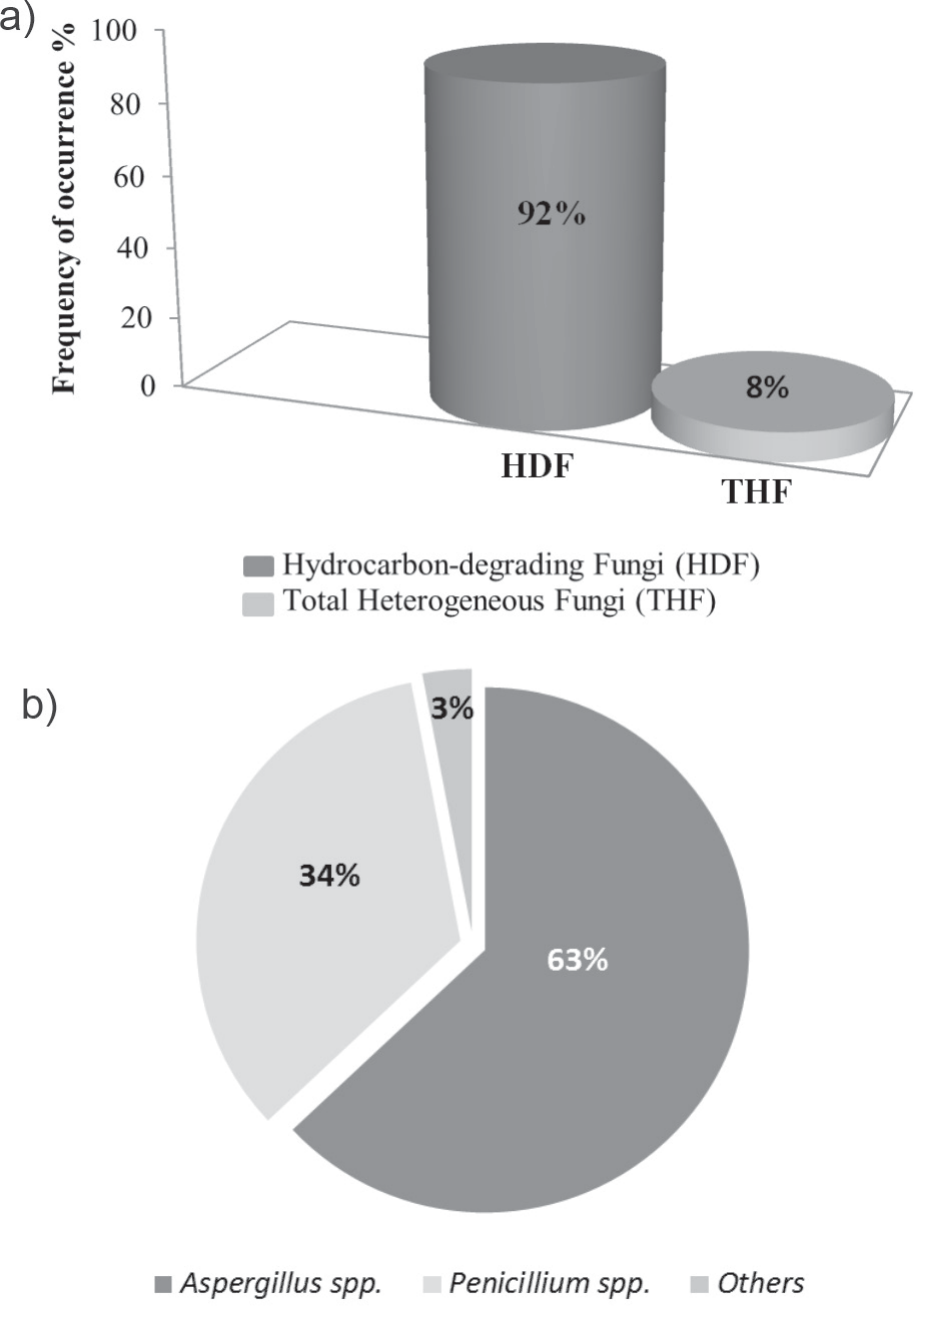
Following the screening of 68 fungal isolates gathered from various geographic locations within the KRI oilfields, it became evident that in soil samples laden with oil, the presence of hydrocarbon-degrading fungi (HDF) significantly outweighed the total heterogeneous fungi (THF). Specifically, HDF had a frequency of occurrence of 92%, while THF accounted for only 8% (Fig. 2a). Moreover, within the 92% frequency of HDF isolates, the distribution of these isolates was as follows: 63% corresponded to Aspergillus spp., 34% to Penicillium sp., and a mere 3% to other types of isolates (Fig. 2b). The findings derived from Fig. 2a) and Fig. 2b) led to the conclusion that HDF isolates, as opposed to THF strains, exhibited greater efficiency in utilizing crude oil as their exclusive carbon source and energy source [5]. These observations align with earlier conclusions made by previous researchers in a similar context [4-16]. Furthermore, it was obvious that the four identified most potent HDF isolates were followed by an order as A. fumigatus> P. chrysogenum> A. niger> A. flavus concerning their ability to degrade/ utilize total petroleum hydrocarbons (TPH) in oil-laden soil samples, as depicted in Fig. 3, additionally, they were followed by an equal order in their occurrence percent at the different studied geographical locations within the studied area (i.e., KRI oilfields).
Plate (1): Aspergillus fumigatus Size: 200-400. Stipe’s color: grayish near apex. Surface: smooth walled. Vesicle serration: uni-seriate pyriform. Metula covering: 2/3. Shape: globuse small in columns. Conidia surface: smooth or spiny.
Plate (2): Aspergillus flavus Size: 400-800. Stipe’s color: pale brown roughened. Surface: quietly spherical. Vesicle serration: bi-seriate. Metula covering: 3/4. Shape: glubose ellipsoid. Conidia surface: smooth finely roughened.
Plate (3): Aspergillus niger Size: 400-3000. Stipe’s color: slightly brown. Surface: smooth walled. Vesicle serration: bi-seriate large size. Metula covering: entirely. Shape: glubose. Conidia surface: very rough irregular.
Plate (4): Penicillium chrysogenum Stipe’s: short, smooth. Penicilli: treverticillate. Phialides: ampulliform. Collula: short. Conidia: spherical to ellipsoidal smooth, greenish.
Fig. 3.
Total petroleum hydrocarbon (TPH) removal rate/degradation (%) via applied fungal strains during the studied period (n = 5).
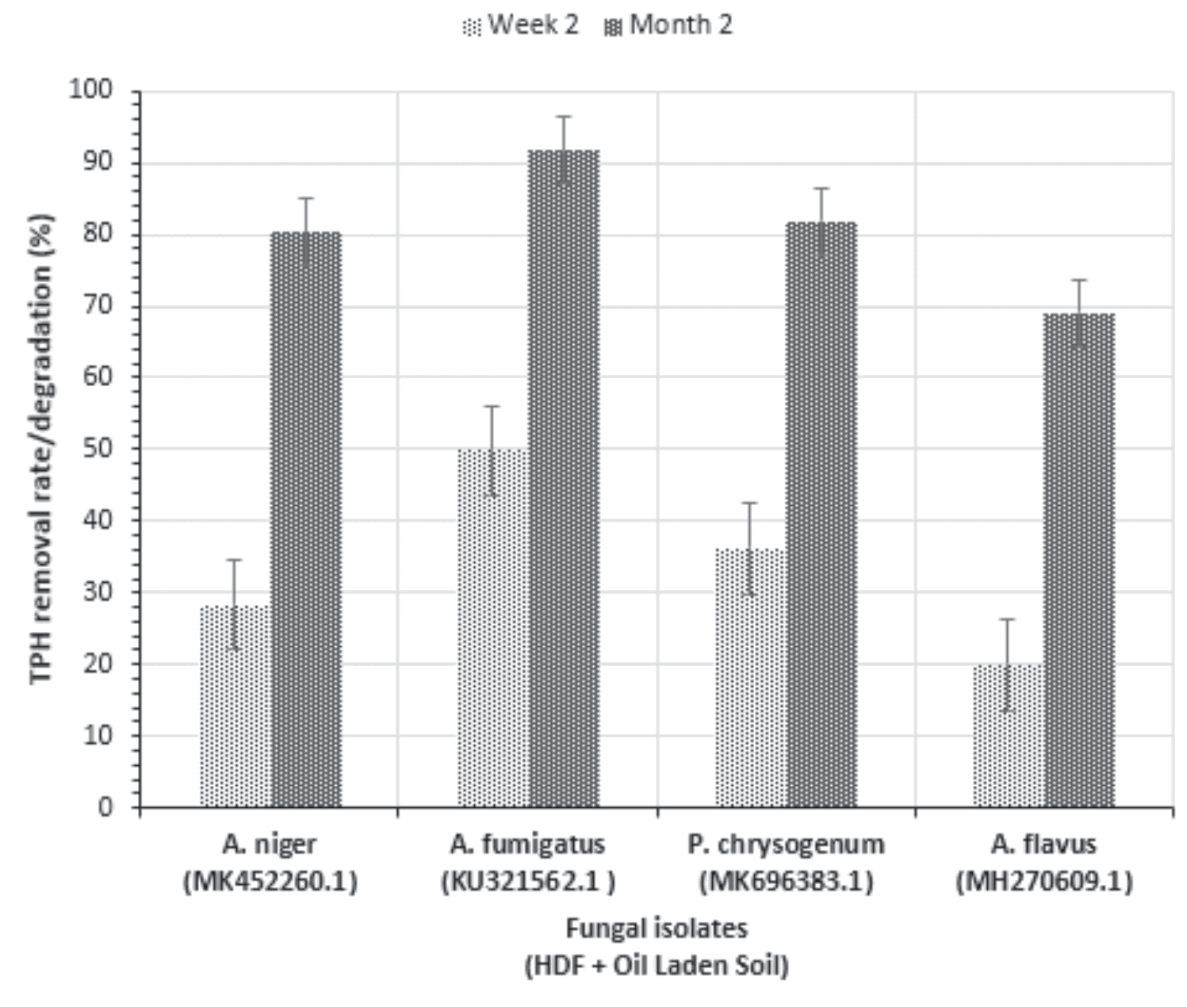
Fig. 4.
The partial sequencing a) and pair wise alignment partial genes b) results for A. fumigatus Strain 004.
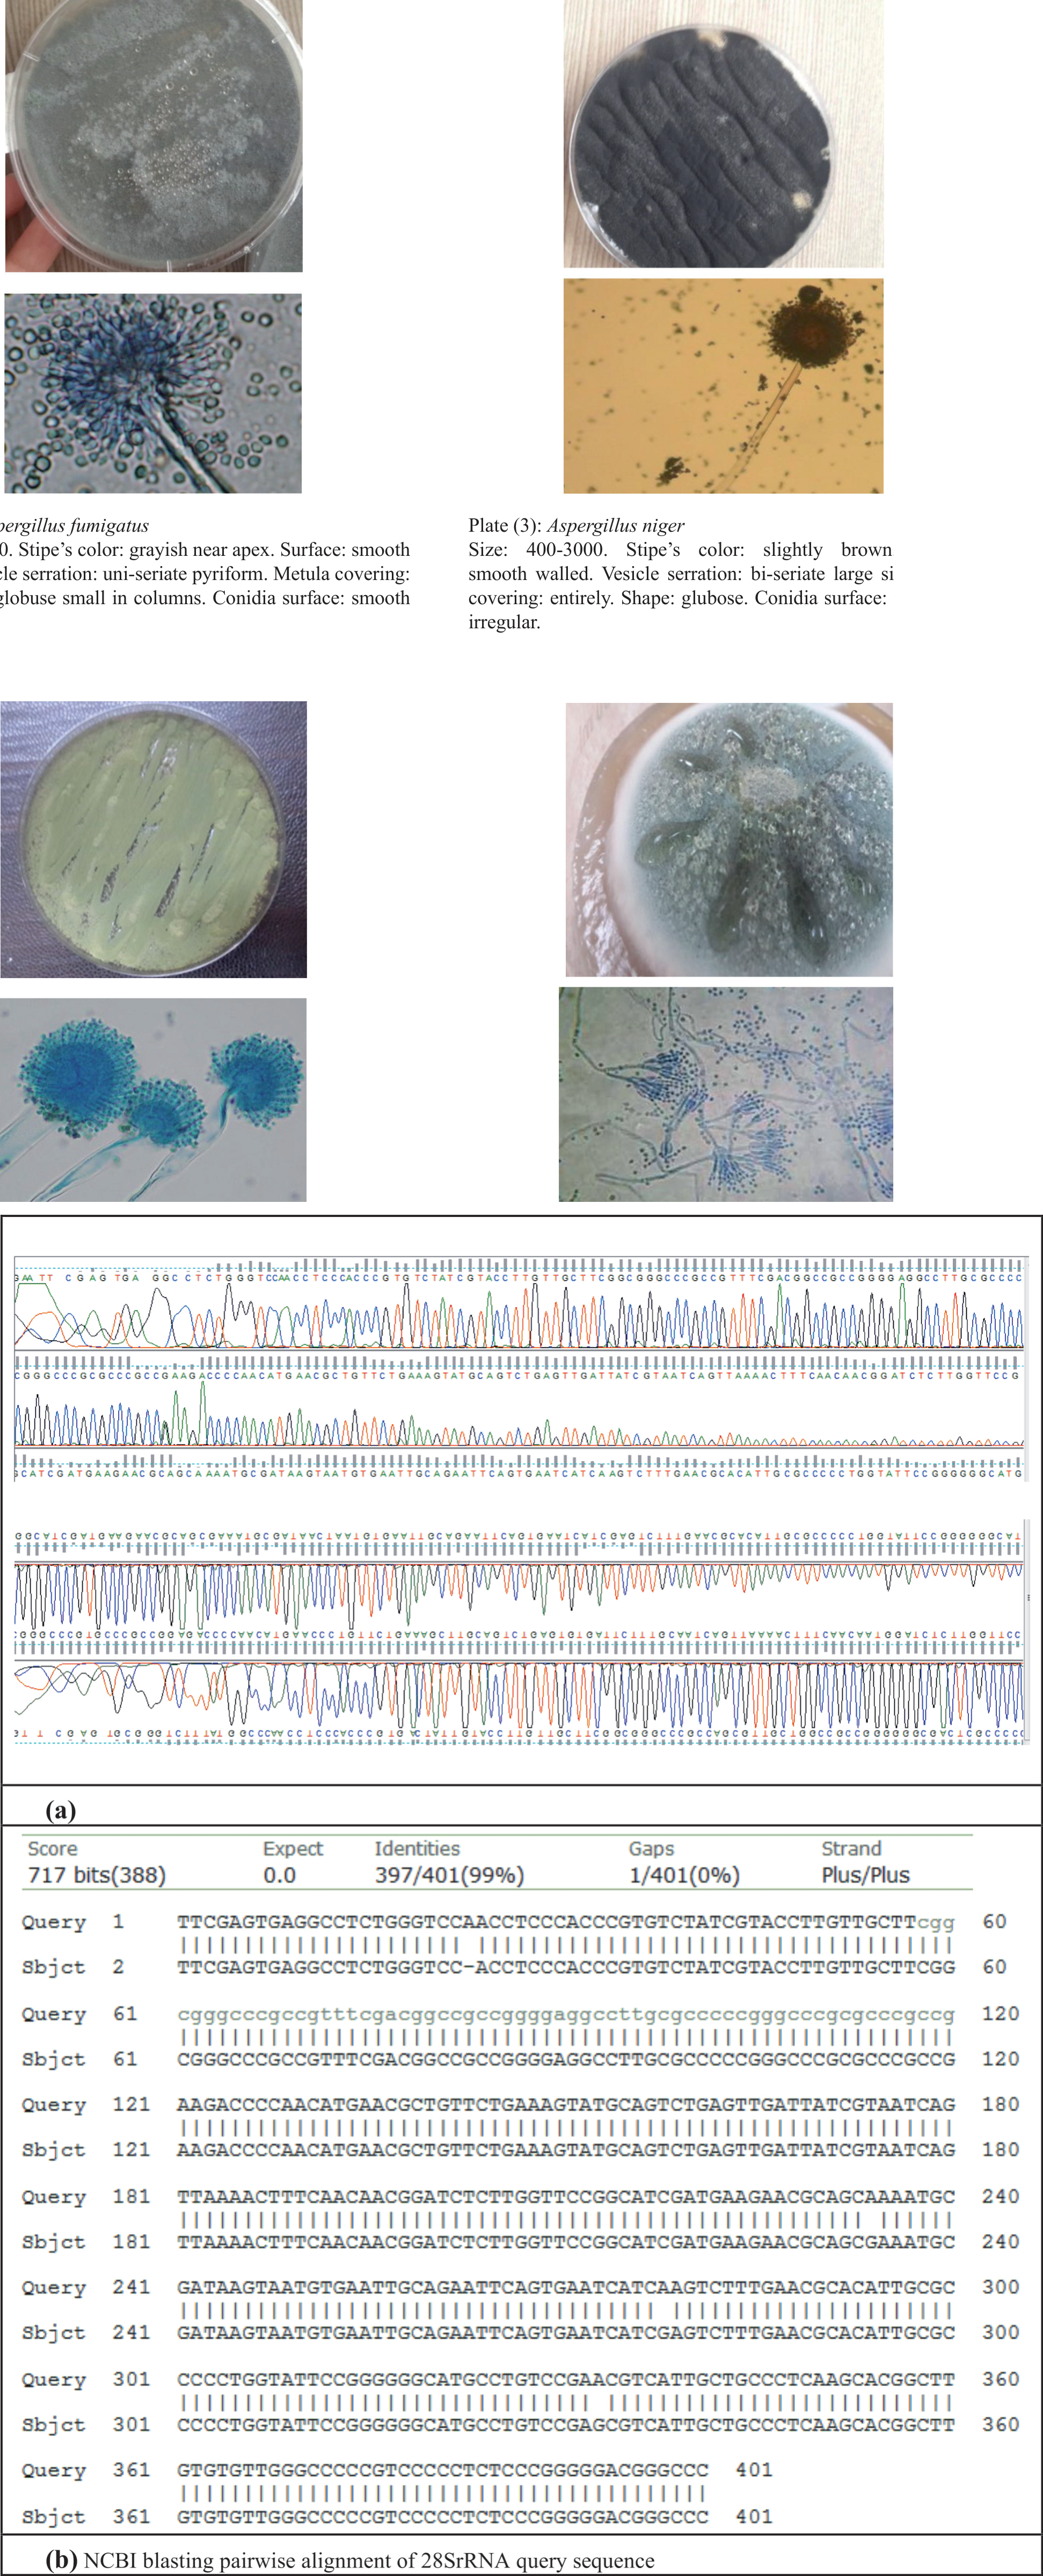
Table 1.
Identification of fungus, along GenBank accessed number and GenBank species identification.
Table 2.
Identification of most potent isolated HDFs, along accession numbers submitted in NCBI.
Morphology (Macroscopic and Microscopic) Features of Isolated Fungi
The examination of the isolated fungi was conducted by assessing their cultural, microscopic, and morphological attributes. Out of 68 fungal strains, only four fungal species were identified as the most potent hydrocarbon degrading/utilizing isolates, and they were selected for further identification, as A. fumigatus, A. flavus, A. niger, and P. chrysogenum. To avoid repetition, microscopic morphological features of the identified species are shown in self-explanatory plates (1, 2, 3, and 4), respectively.
Taxonomically, the four fungal isolates belong to the Ascomycota Phylum, Eurotiomycetes (class). The family Trichocomaceae and the order Eurotiales have been recognized in the context of fungal taxonomy [17-19]. Numerous researchers have examined a range of factors that can influence the effectiveness of petroleum hydrocarbon degradation by various fungal species in soil environments [25-31]. A diverse array of fungi was isolated from various geographic regions and substrates and subsequently identified using both conventional and contemporary methodologies [4, 12- 16]. These observations may hold true for the fungi under investigation in this study. In our research, we adopted a multifaceted approach that included both macroscopic and microscopic examinations, as well as molecular techniques, to isolate and identify native fungal strains proficient in hydrocarbon degradation in oil-contaminated soils. Existing research has indicated that, while morphological characteristics can effectively differentiate isolates into their respective phyla and, in many instances, the correct genera, it is always advisable to use a combination of several methods. This is because the characterization and identification of unknown fungal isolates can be fraught with errors when relying solely on a single method [5, 18, 19, 32].
Molecular Identification and DNA Extraction from Isolated Fungi
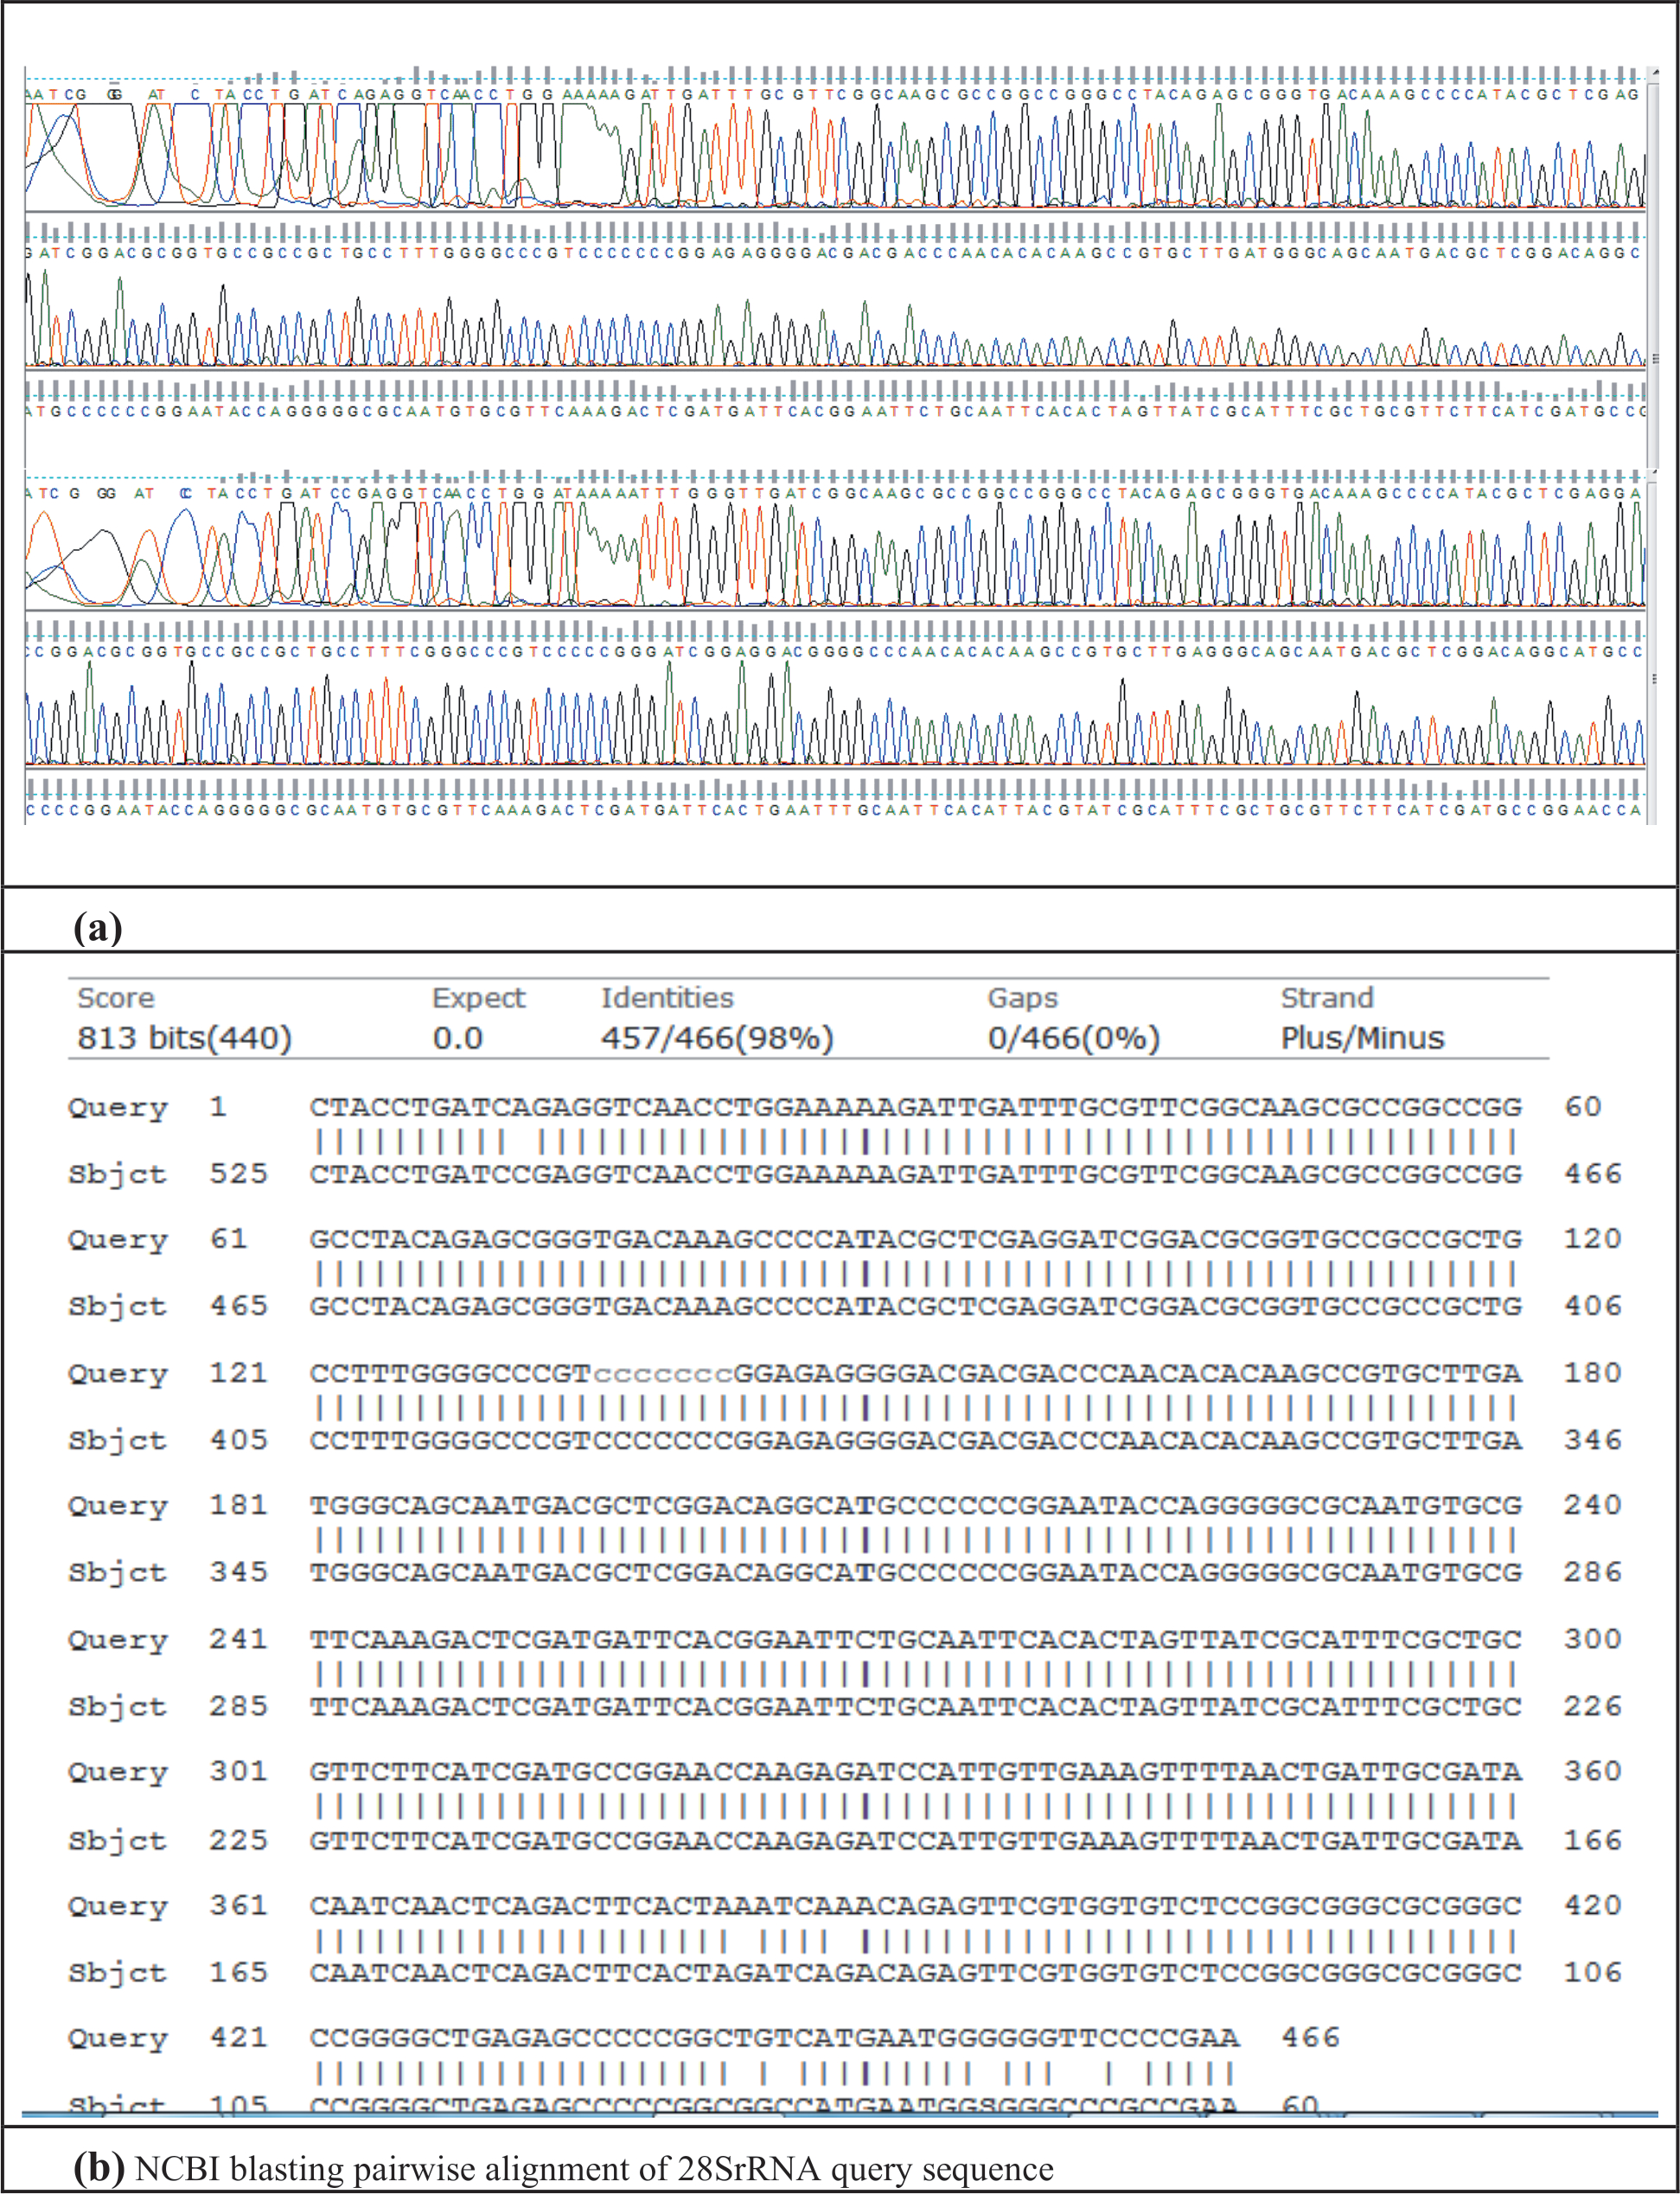
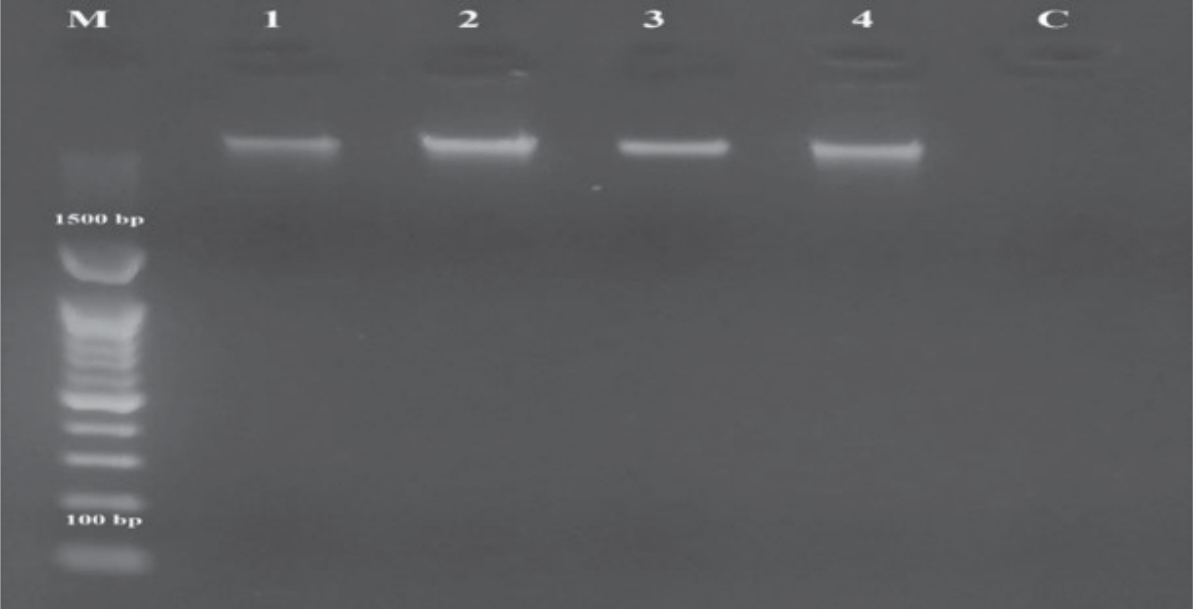
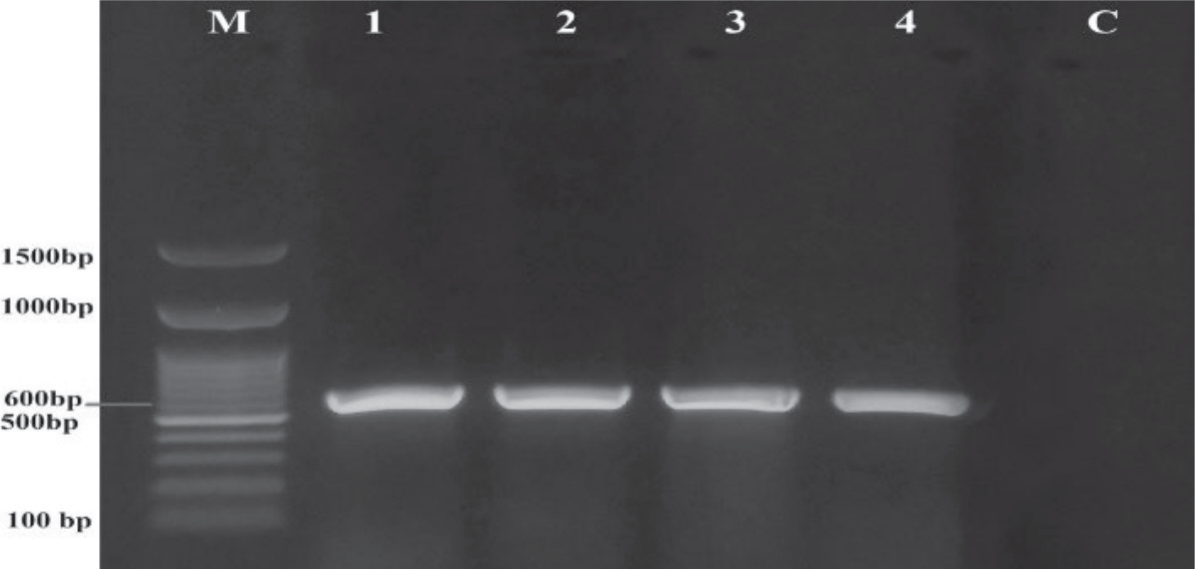
Specifically, designed primers were employed to amplify the Internal Transcribed Spacer (ITS1) and (ITS2) regions, and the resulting PCR products were visualized through electrophoresis on a 2% agarose gel stained with ethidium bromide. The molecular identification of these isolates was achieved using ITS1 and ITS2 primers, revealing that they belong to Aspergillus sp. (including A. niger, A. fumigatus, and A. flavus) and Penicillium sp. (specifically P. chrysogenum). Partial sequencing results and pairwise alignment of partial genes were performed for A. fumigatus, A. flavus, A. niger, and P. chrysogenum, as illustrated in Figs. (4a, 4b, 5a, 5b, 6a, 6b, and 7a, 7b), respectively. Additional comprehensive data, including BLASTN results with hit identifiers and scoring for 28SrRNA sequences of these fungi, can be obtained from the corresponding author. A phylogenetic tree was constructed based on these four sequences, comparing them against nine other related sequences from the GenBank database using MEGA Version-4. The analysis revealed that these fungal species were closely related to many other Aspergillus and Penicillium species present in the GenBank database, as summarized in Table 1, which was updated based on the current results (Table 2). Fig. 8 illustrates the DNA extraction product obtained from fungal isolates and the control, focusing on the internal transcribed spacer (ITS) gene in conjunction with a 600-100 bp marker. Meanwhile, Fig. 9 depicts the polymerase chain reaction (PCR) amplified products derived from different fungi, including A. niger, A. fumigatus, P. chrysogenum, and A. flavus.
Fig. 5.
The partial sequencing(a) and pair wise alignment partial genes b) results for A. flavus ND103.
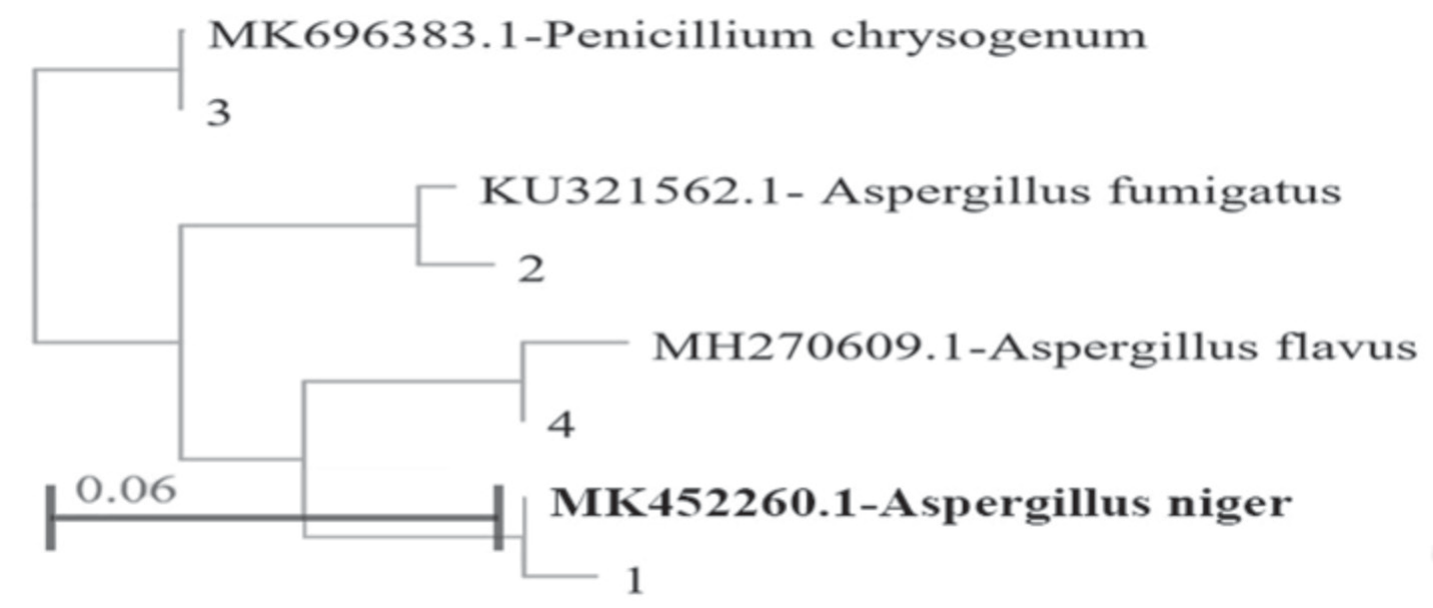
A consensus among many taxonomists in the field is that utilizing DNA coding serves as a valuable tool for the identification and cataloging of fungal species [14, 20]. Molecular methods like PCR and DNA sequencing have demonstrated a high level of specificity and sensitivity in the detection of various species of organisms [14]. Molecular techniques, specifically the innovation of the polymerase chain reaction (PCR), have brought about a significant transformation in the field of molecular biology and the molecular diagnosis of fungi. It’s worth noting that, as of now, there hasn’t been a comprehensive examination employing molecular techniques for the classification and diagnosis of Aspergillus spp. and Penicillium spp. in Iraq [4]. The research conducted by the reference [4] involving the extraction of genomic DNA and PCR amplification of marker genes for the identification of a substantial number of soil and airborne fungi may provide supportive evidence for the DNA sequence and PCR amplification findings in our current study. They observed that similar coordination steps can be observed for the fungal species investigated in our work. However, Fig. 8 and Fig. 9 illustrate that the primer subsequence of A. fumigatus and, to some extent, P. chrysogenum, along with two other fungi, results in an approximately 600 bp DNA fragment in the PCR process. The primer for A. fumigatus has been designed to be specific to the fungal system. These specific primers exhibited strong amplification capabilities for a section of the 18SrDNA sequences of A. fumigatus and P. chrysogenum. These sequences were aligned for different fungal species as follows: A. niger (Lane 1), A. fumigatus (Lane 2), P. chrysogenum (Lane 3), and A. flavus (Lane 4) (Fig. 9). In addition, Fig. 10 demonstrates that the construction of phylogenetic trees and alignment reconstructions were carried out using the BUILD Function of the Environment for Tree Exploration (ETE) V3.0.0b32, as conducted on the GenomeNet platform [4]. The alignment process was conducted using the MAFFT program with default settings, which involved multiple alignments through the application of a fast Fourier transform [33-36]. Furthermore, the tree was constructed using the default parameters of Fast Tree V2.1.8. The internal transcribed spacer (ITS) region of the filamentous fungi strains was organized and subsequently added to the GenBank sequence database under the accession number MK452260.1, as evident in the optimal evolutionary tree representing the primary strains. This evolutionary tree illustrates the potent fungal isolates based on the ITS sequence. The findings revealed that both morphometric data and molecular methods were effective in the identification of A. fumigatus (GenBank accession KU321562.1), A. flavus (GenBank accession MH270609.1), A. niger (GenBank accession MK452260.1), and P. chrysogenum (GenBank accession MK696383.1). Utilizing ITS rDNA sequence similarity (95%) to establish species boundaries operationally, a total of 277 fungal species were identified from 1,403 endophytic strains isolated from various common plants across arctic, boreal, temperate, and tropical regions, representing a wide range of phylogenetically diverse plant taxa [37].
Fig. 6.
The partial sequencing a) and pair wise alignment partial genes b) results for A. niger Strain F3.
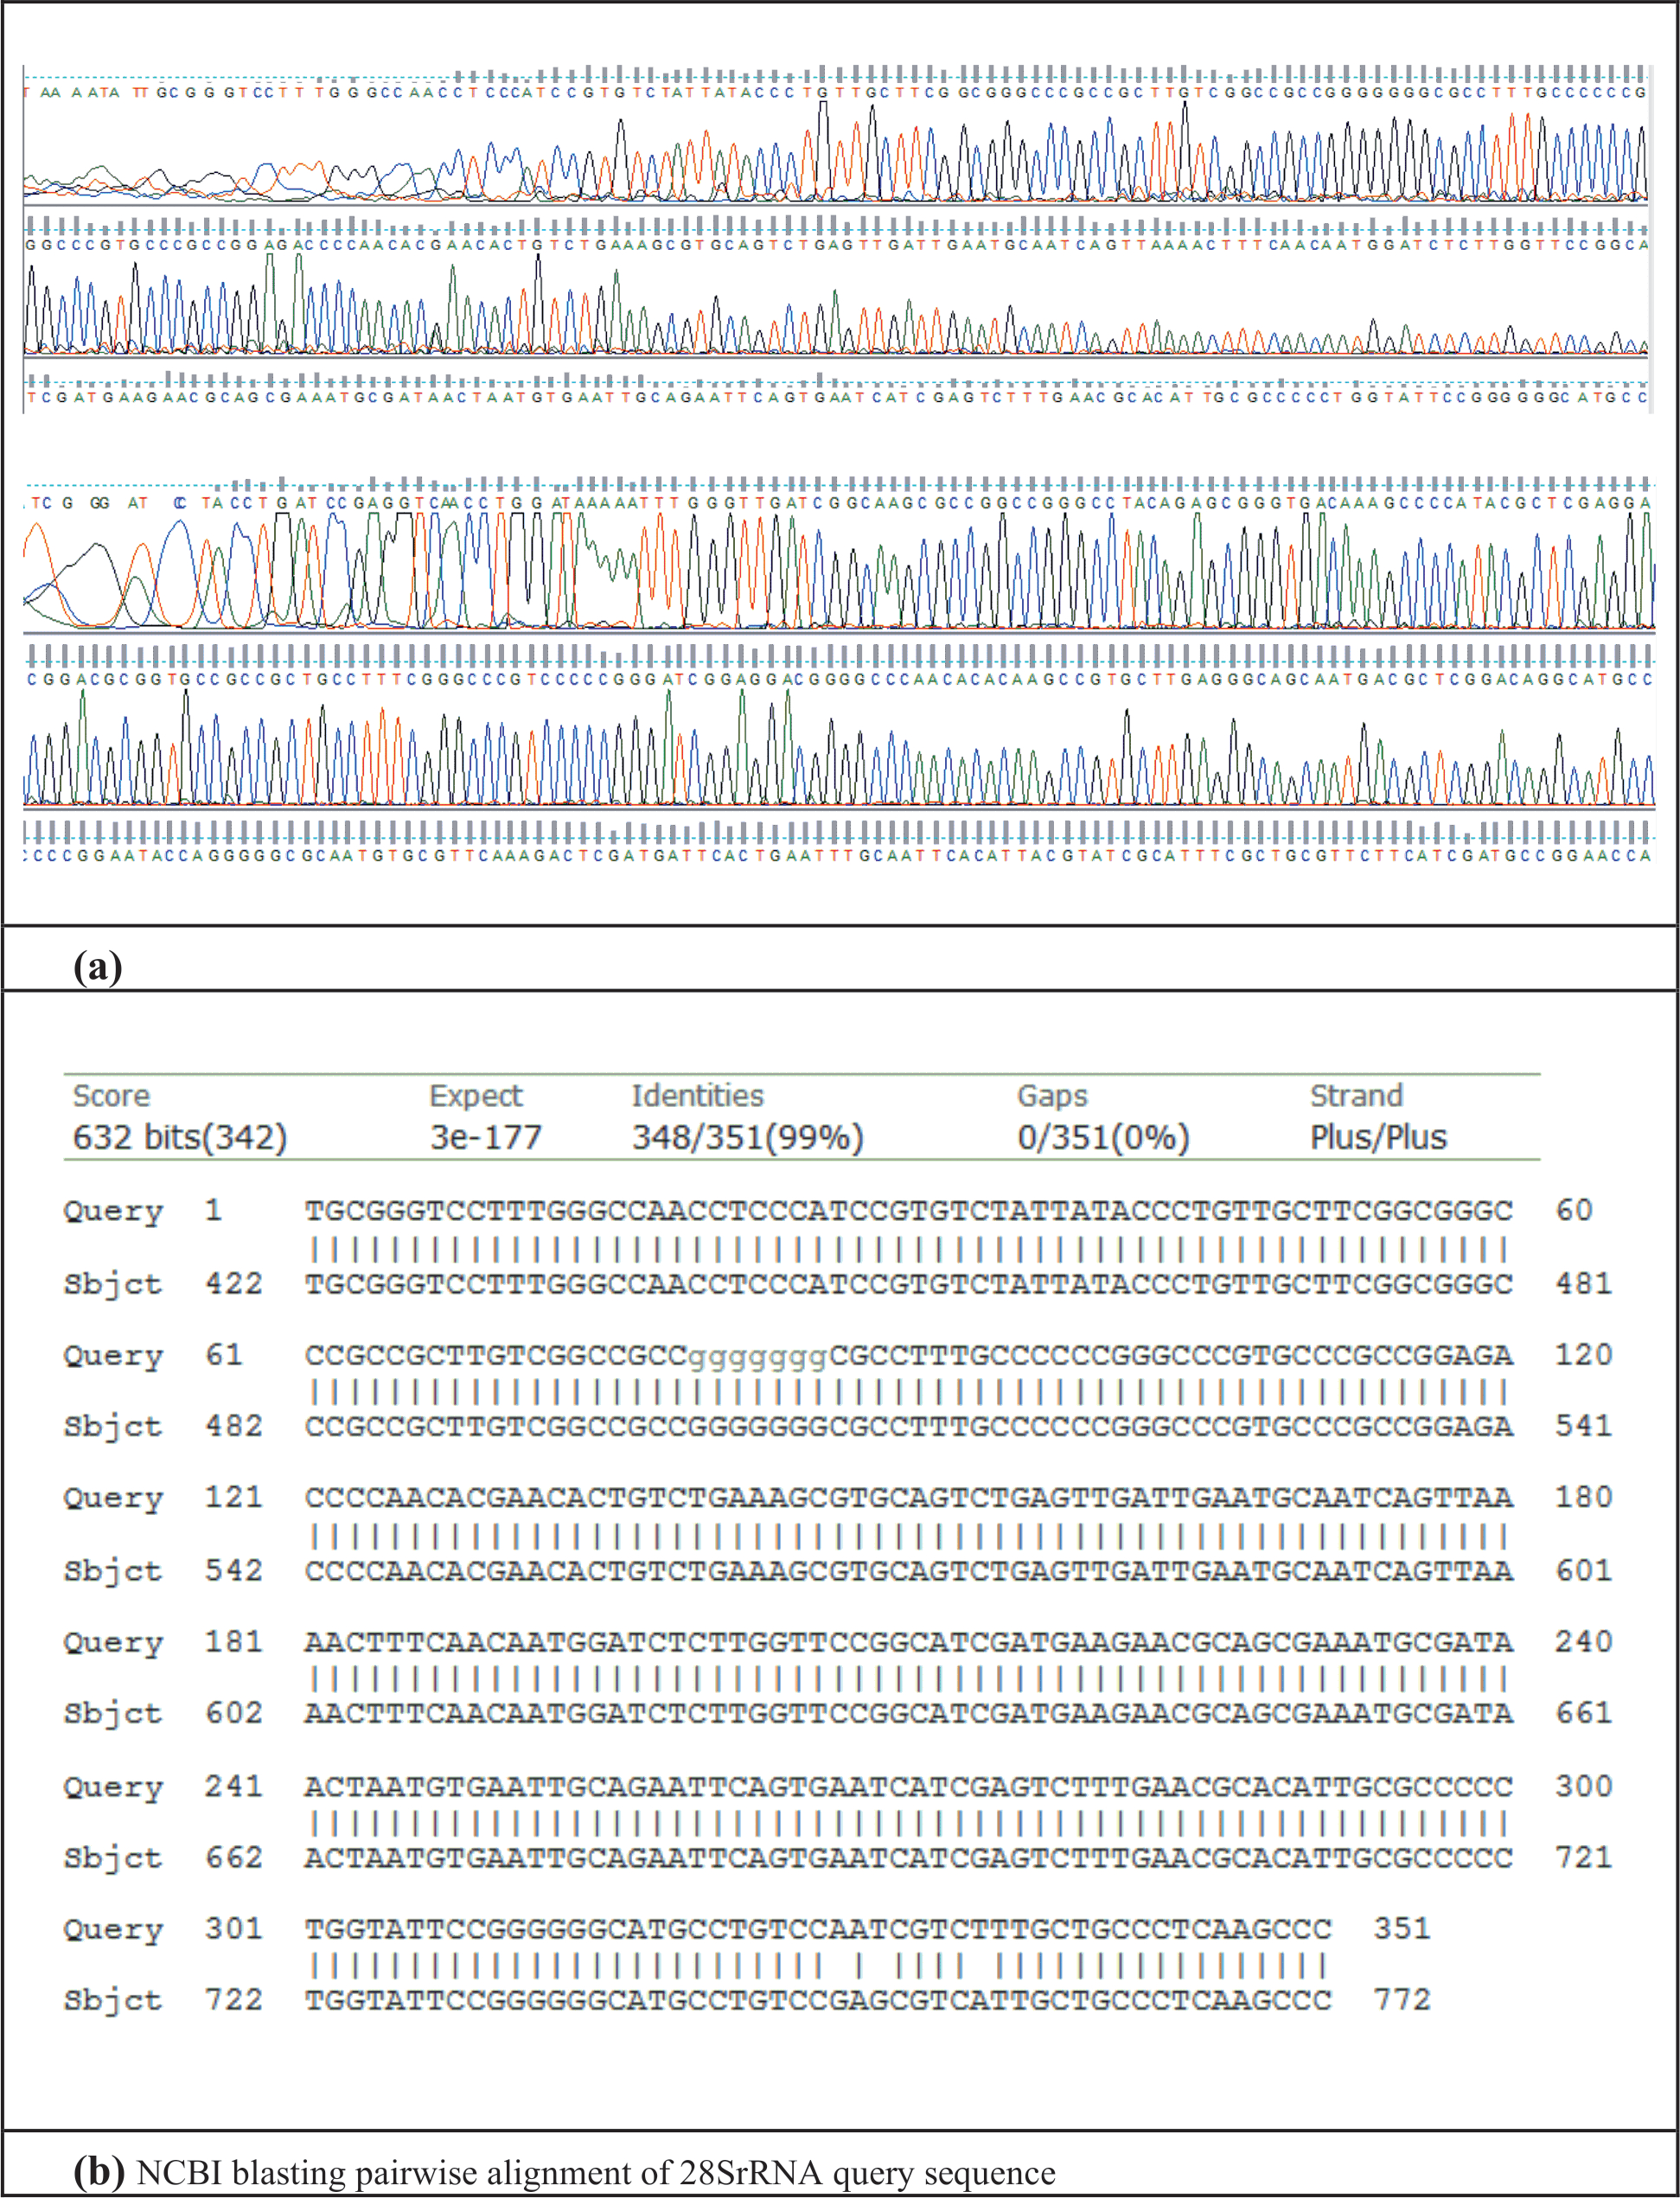
Fig. 7.
The partial sequencing a) and pair wise alignment partial genes b) results for P. chrysogenum Strain CBS132208.
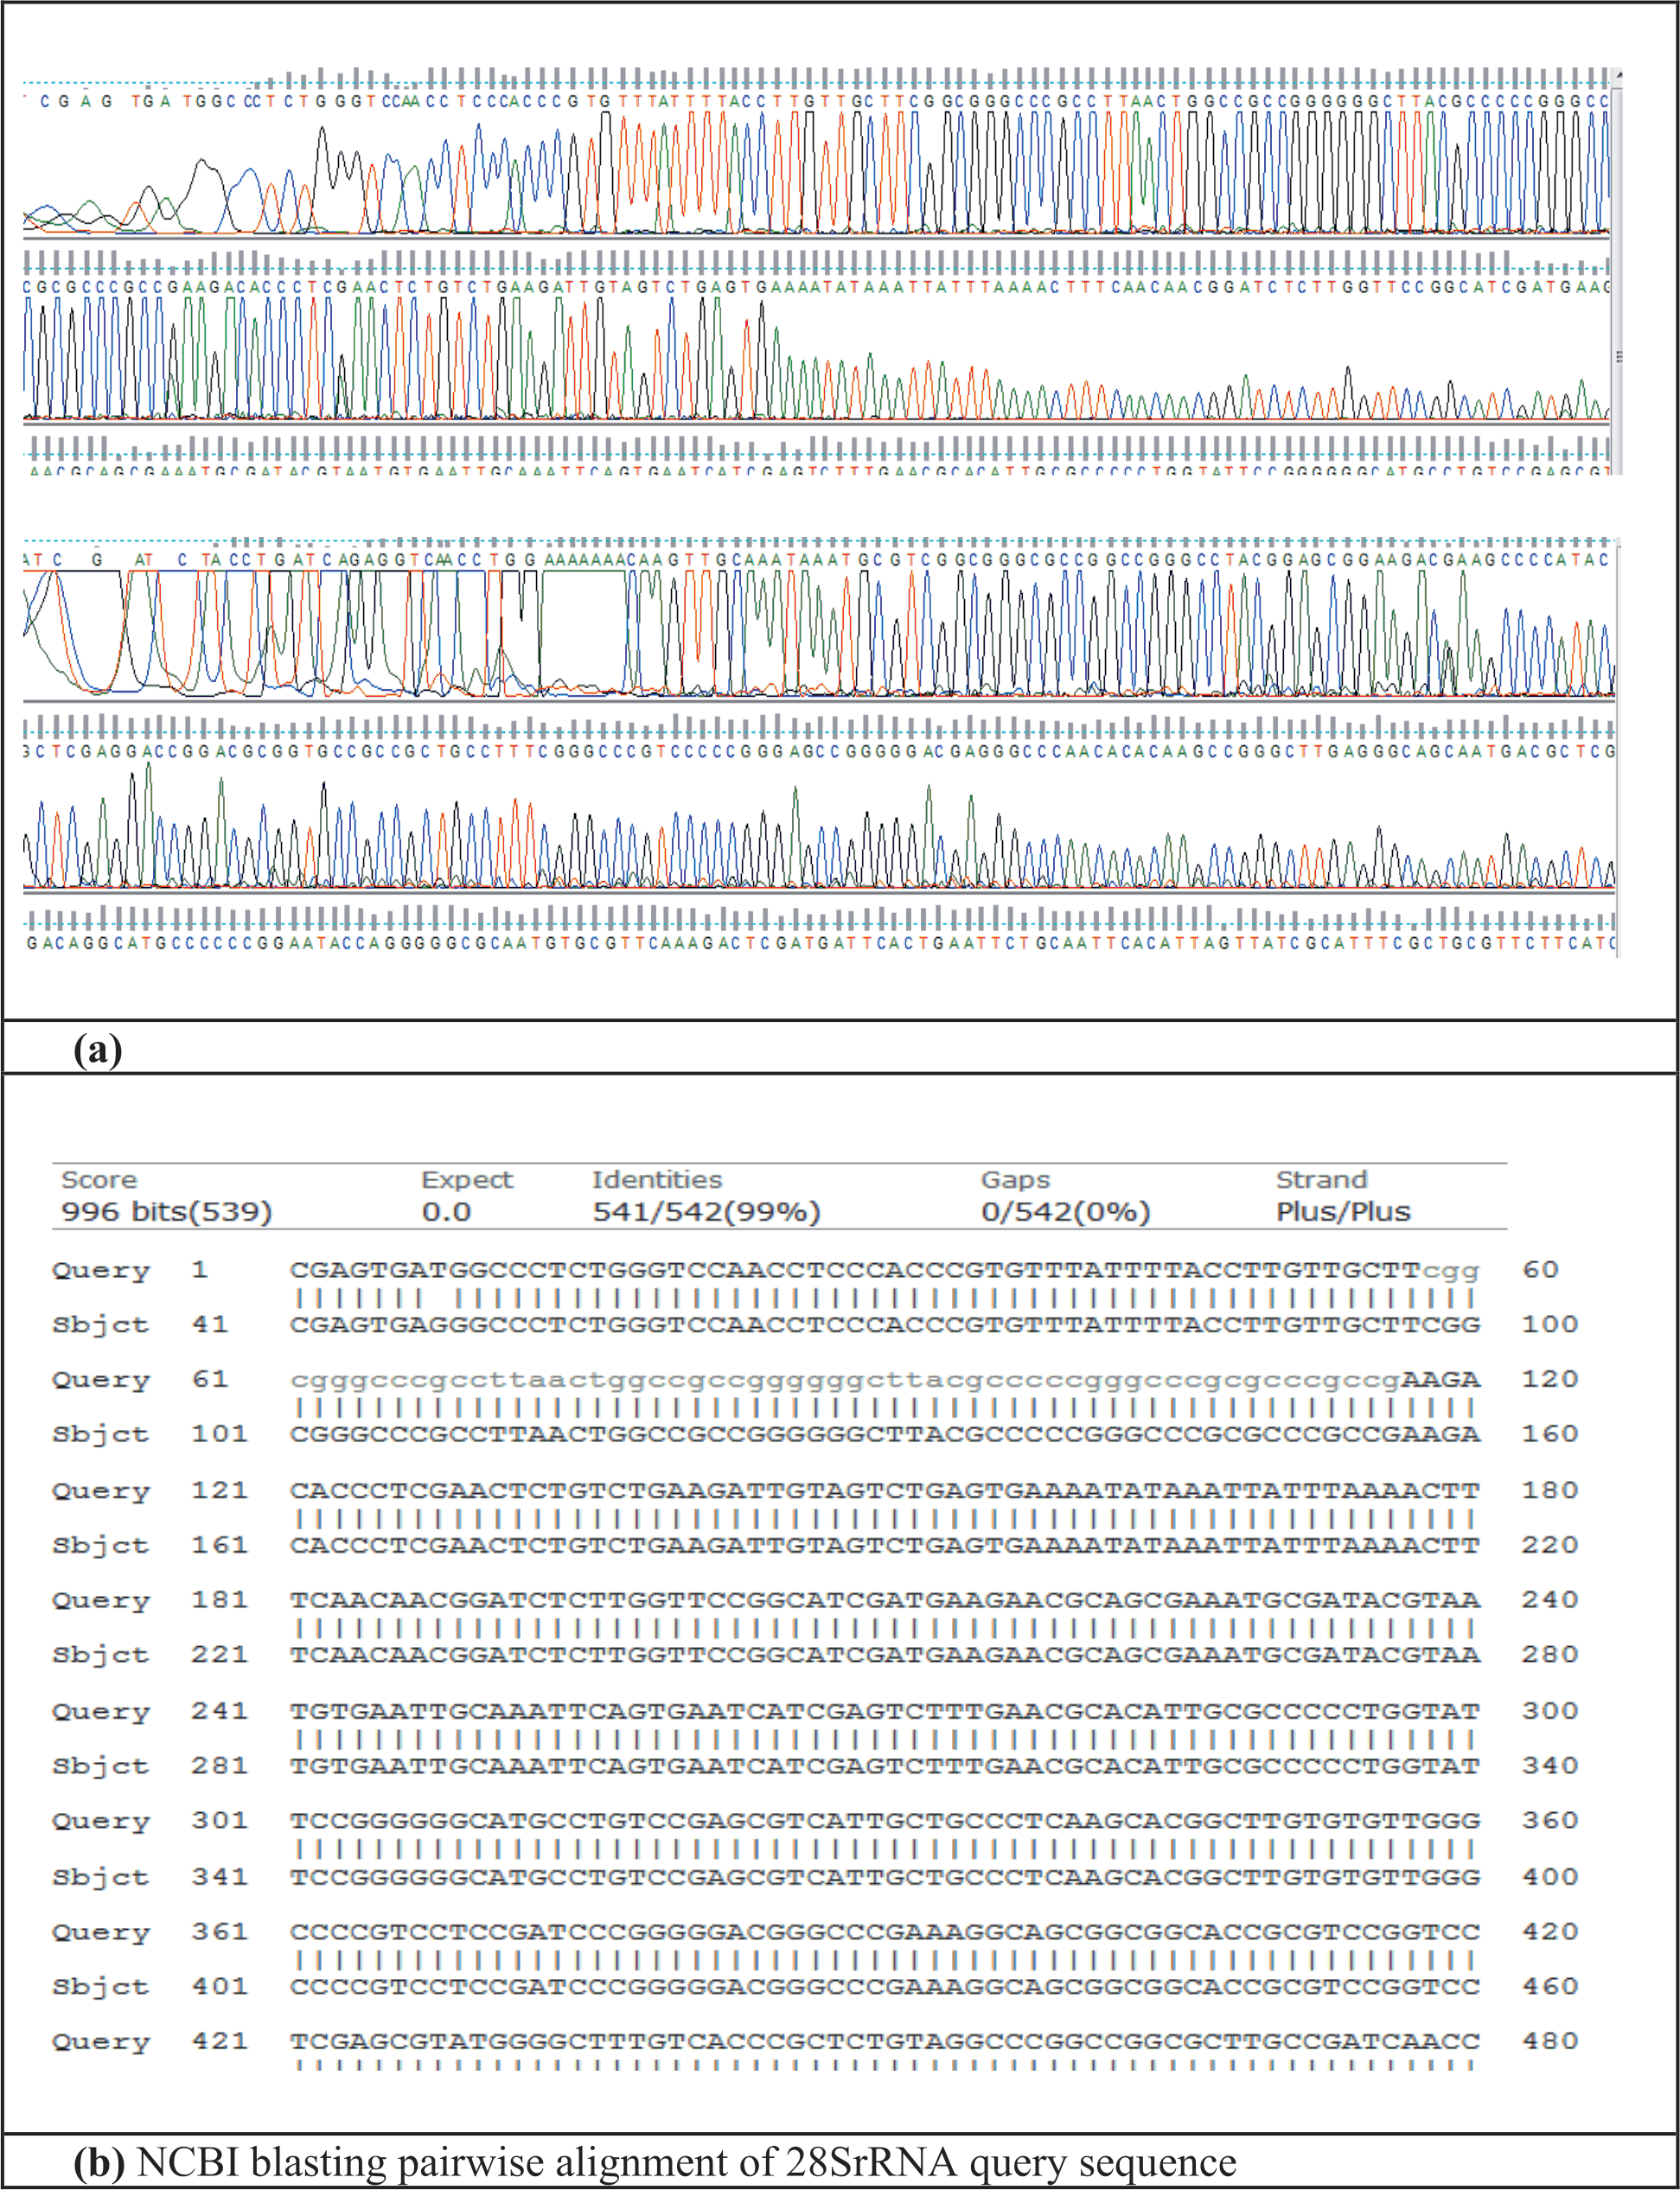
Fig. 8.
DNA extraction product from fungal isolates and control for ITS gene along with 600-100 bp marker. Where; M = markers; 1, 2, 3 & 4 = isolated fungi and C = control.
Dendrogram Method
The results of the dendrogram analysis, aimed at identifying the most potent isolated fungal species, are as follows: A. niger with GenBank accession MK452260.1, A. fumigatus with GenBank accession KU321562.1, P. chrysogenum with GenBank accession MK696383.1, and A. flavus with GenBank accession MH270609.1, as depicted in Fig. 10 and Table 2.
Conclusions
In this study, two crude oil-degrading strains, Aspergillus and Penicillium, were isolated from KRI oilfields. These strains exhibited high levels of crude oil degradation, which can possibly be used for bioremediation. Incorporating molecular techniques into the traditional taxonomy based on morphology provides immense value in distinguishing between different fungal species and their variations. This highlights the increasing importance of identifying genes that can differentiate fungal organisms at various taxonomic levels without the need for prior cultivation. DNA sequences and other genetic markers provide comprehensive datasets that are not contingent on cultivation and are unaffected by physiological variations. These genetic markers consistently unveil a wealth of identification information within the genetic data, allowing for the adjustment of the level of detail by selecting the appropriate genes.

