Introduction
Water is essential for life because it supports and sustains all forms of life. Without water, no living thing on the planet could survive. Moreover, water is crucial for residential and commercial purposes in human civilization [1]. The presence of both organic and inorganic substances in water provides an ideal environment for microbial growth. Prokaryotes distribution could be differing in aquatic regions according to different environmental parameters in different habitats [2]. Prokaryotes can be found in a wide range of environments, including water, land, deep-sea sediments, etc. Prokaryotic communities are essential in various ecosystems, including both man-made environments such as wastewater treatment plants and natural habitats like rivers. These communities are involved in critical processes such as nutrient cycling, carbon fixation, and decomposition of organic matter, which maintain ecosystem health and function [3]. For example, in wastewater treatment plants, prokaryotic communities are responsible for the removal of organic and inorganic pollutants from wastewater, making them crucial in ensuring the quality of treated water before discharge into the environment [4].
The characterization of prokaryotic communities is critical in understanding their structure and function in these environments. In recent years, advances in sequencing technologies have revolutionized the field of microbial ecology, allowing for the identification and quantification of prokaryotic communities in both man-made and natural habitats. Advanced high-throughput sequencing technologies, particularly next-generation sequencing (NGS), have revolutionized our ability to analyze entire prokaryotic communities within a sample. This has facilitated a more thorough and nuanced understanding of the structure and diversity of these communities [5].
Several studies have characterized prokaryotic communities in both man-made and natural habitats. For example, a study by Jiang et al. [6] investigated the prokaryotic community composition in a wastewater treatment plant using high-throughput sequencing (HTS). The study revealed the dominance of Proteobacteria, Bacteroidetes, and Actinobacteria phyla, which are commonly found in such environments. Another study by Liu et al. [7] analyzed the prokaryotic community composition in the Yangtze River, China, using 16S rRNA gene sequencing. The study identified a high diversity of prokaryotes, with Proteobacteria and Bacteroidetes being the dominant phyla.
Moreover, comparisons between prokaryotic communities in man-made and natural habitats have highlighted significant differences in their diversity and composition. For instance, a study by Li et al. [8] compared the prokaryotic community compositions in a wastewater treatment plant and a river using metagenomic sequencing. The study identified a higher diversity of prokaryotic communities in the river, with Actinobacteria and Cyanobacteria dominating. In contrast, the wastewater treatment plant had a lower diversity of prokaryotic communities, dominated by Proteobacteria and Bacteroidetes.
Microorganisms are the backbone of anaerobic processes and sewage treatment systems, where they are in charge of removing pathogens and contaminants, recovering nutrients, and creating safe water [9]. During wastewater treatment, prokaryotes are primarily responsible for eliminating carbon (C) molecules and biologically active chemicals in soluble forms [10, 11]. Additionally, prokaryotic bacteria can be employed to boost the effectiveness of treating wastewater, particularly in terms of nitrogen removal [12]. Heterotrophic aerobic Gram-negative bacteria are very powerful in the degradation process of xenobiotic and aerobic sewage treatment. Cyanobacteria and oxygenic prokaryotic phototrophs are employed to effectively remove nitrogen and phosphorus from wastewater used in the food processing industry. Additionally, sulfur-oxidizing chemolithotrophic bacteria are utilized to mitigate the presence of harmful H2S in water and wastewater by oxidizing reduced sulfur compounds. Iron-oxidizing bacteria can oxidize Fe(II) in acidic or neutral conditions. They eliminate iron from water for drinking [13]. HTS technology is an advanced technology applied to characterize various prokaryotes in water and wastewater, which could help to understand their roles in water treatment [14, 15].
The Nile River, as Egypt's primary freshwater source, is integral for drinking, agricultural, and industrial activities. Consequently, the river's water quality directly impacts the health of the local population. The main source of water pollution of the Nile River is wastewater [16]. Few studies about prokaryotes diversity in natural water in Egypt limit the understanding of the microbial ecosystem’s aspects within these habitats. In environmental remediation, prokaryotes can be employed to remove contaminants like agrochemicals that seep into water from the soil. Biological treatment is another method that can be used to get rid of some hazardous metals from water, like selenium and arsenic compounds. Additionally, prokaryotes can be used to eliminate oil spills [17, 18]. There is limited information on the characterization of prokaryotic community compositions in man-made (wastewater treatment plant) and natural habitats (Nile River) in Egypt [19- 21]. While studies have been conducted in other parts of the world, the specific environmental conditions and anthropogenic activities in Egypt may result in unique microbial communities. Additionally, there is a lack of comprehensive studies that compare the prokaryotic communities in both man-made and natural habitats in Egypt. Thus this study aimed to apply HTS technology to characterize and compare the prokaryotic composition in surface water of the Nile River as well as the inlet and outlet of Zenin wastewater treatment plant.
Material and Methods
Study Area and Sample Collection
Three habitats, including the inlet and outlet of Zenin wastewater treatment plant (WWTP) and an urban location at the Nile River, were screened using next-generation high-throughput 16S rRNA sequencing. Zenin WWTP is based on activated sludge technology and the actual capacity of the station is from 400000 to 500000 m3/day. The station discharges its effluent in Libyan discharge then to Al Rahway discharge and finally to the Rosetta branch. Triplicate samples were collected from each habitat and filtered by 0.2 μm Isopora membrane filters (Millipore, Bedford, MA, USA) for prokaryotic community analysis. The surface water sample volume was about 1000 mL and the wastewater sample volume was about 500 mL for inlet and outlet samples. The filters were stored at −80ºC until DNA extraction.
DNA Extraction, 16S rRNA Gene Full-Length High-Throughput Sequencing
Environmental DNA was extracted from this study’s samples using the DNeasy PowerLyzer PowerSoil Kit (QIAGEN, USA), and quantified using the Qubit 2.0 fluorometer with Qubit dsDNA BR assay kit (Life Technologies, Grand Island, NY, USA). The full length of 16S rRNA genes were amplified by using the primer pair; 27F (5’-AGRGTTYGATYMTGGCTCAG-3’) and 1492R (5’-RGYTACCTTGTTACGACTT-3’) [22]. PCR reactions were performed in 25 μL in triplicate containing 5 μL TransStart FastPfu Buffer (5×), 2 μL dNTPs (2.5 mM), 0.5 μL TransStart FastPfu DNA Polymerase (2.5 units/μL, TransGen Biotech, Beijing, China) and 0.4 μM of forward and reverse primers, and 10 ng of template DNA. The PCR amplification cycle was set as 95ºC for 5 min for initial denaturation, followed by 30 cycles of 95ºC for 30 s, 55ºC for 45 s, and 72ºC for 90 s, and followed by a final extension at 72ºC for 10 min. PCR reactions were performed for each sample, and the PCR products were purified using the QIAquick@ Gel Extraction Kit (Qiagen, Santa Clarita, CA, USA). Sequencing libraries were generated using SMRTbellTM Template Prep Kit (Pacific Biosciences, Menlo Park, CA, USA) following the manufacturer’s recommendations, and then sequenced on a PacBio Sequel II platform (Creative-proteomics, NY, USA).
Sequence Analysis
The full-length 16S rRNA gene sequences were quality trimmed by using PacBio SMRT portal v2.3.0. Briefly, sequences not meeting the following criteria were discarded: (i) a minimum pass ≥3; (ii) a minimum predicted accuracy ≥90%; (iii) a sequence length <1340 bp or >1640 bp. The high-quality reads were used for chimera-checking and clustered into operational taxonomic units (OTUs) at the cutoff of 97% identity using UPARSE v7.0.1001 [23]. Each OTU’s representative sequence was classified using the RDP classifier with the SILVA database v138 [24] at a confidence threshold of 80%. In order to standardize the uneven sequencing effort, all samples were randomly subsampled to the smallest library sizes with 6800 reads for prokaryotic communities and respectively.
Physicochemical Analyses
Environmental variables were determined according to the International Standard Methods for Water and Wastewater [25]. Briefly, an AD 360 DO meter (Adwa Instruments, Inc, Europe) was used to determine the temperature, dissolved oxygen (DO) and DO saturation of the water samples in situ. pH was measured using bench pH meter Hach (Hach Sension 1) method 4500 H+B. Electric conductivity (EC) and total dissolved solids (TDS) were measured by Hach (Hach Sension 5) conductivity meter method 2510 B. Chemical oxygen demand (COD) was measured according to dichromate method 5210-D using digestion for two hours at 150ºC by spectrophotometer Hach DR 5000. Total Kjeldahl Nitrogen (TKN) was measured using mercuric sulfate digestion method followed by titration method (4500-Norg), ammonium nitrogen (NH4-N) was measured by Ion Metrohm Omnis Titrator & 940 Professional Ic Vario Ion Chromatography System, nitrite nitrogen (NO2-N) was detected according to a colorimetric method (4500-B) and nitrate nitrogen (NO3-N) by modified sodium salicylate method according to Scheiner [26]. Total nitrogen was calculated as the sum of TKN, NO2-N, NO3-N, and NH4-N. Total phosphorous (TP) was measured according to the method (4500-C) [27].
Statistical Analysis
Principal coordinate analysis (PCoA) based on the Bray-Curtis distance index was used to map prokaryotes in different environments. Distance-based redundancy analysis (dbRDA) was used to show the relationship between the prokaryotic community (response group) and physicochemical parameters (explanatory group) in different habitats. The permutational multivariate analysis of variance (PERMANOVA) and the analysis of similarity (ANOSIM) were used to test the significance of differences for the prokaryotic communities among different habitats. Statistical analyses and visualization were performed using R v4.1.0 (https://www.r-project. org/) and PRIMER v.7.0.21 (Quest Research Limited, Auckland, New Zealand).
Results and Discussion
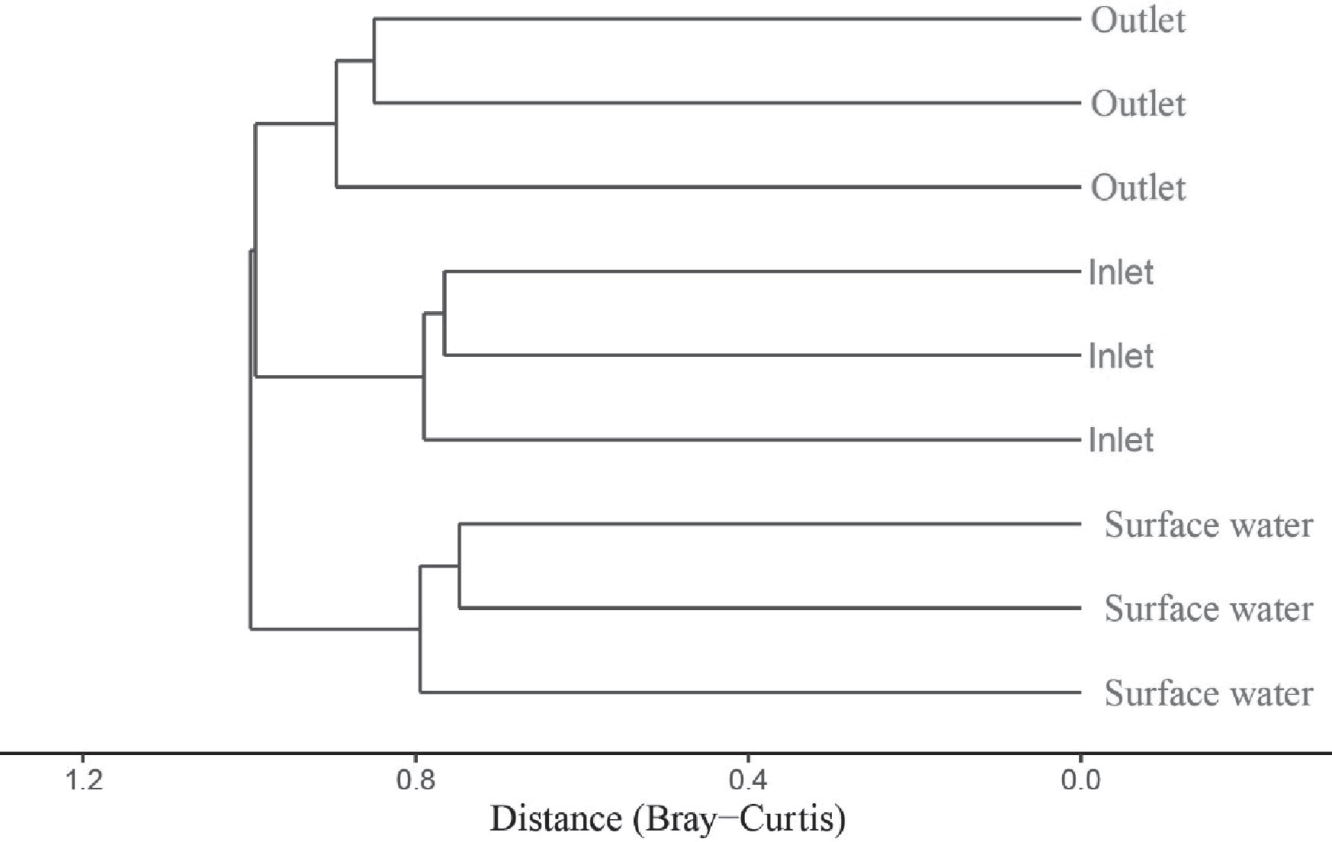
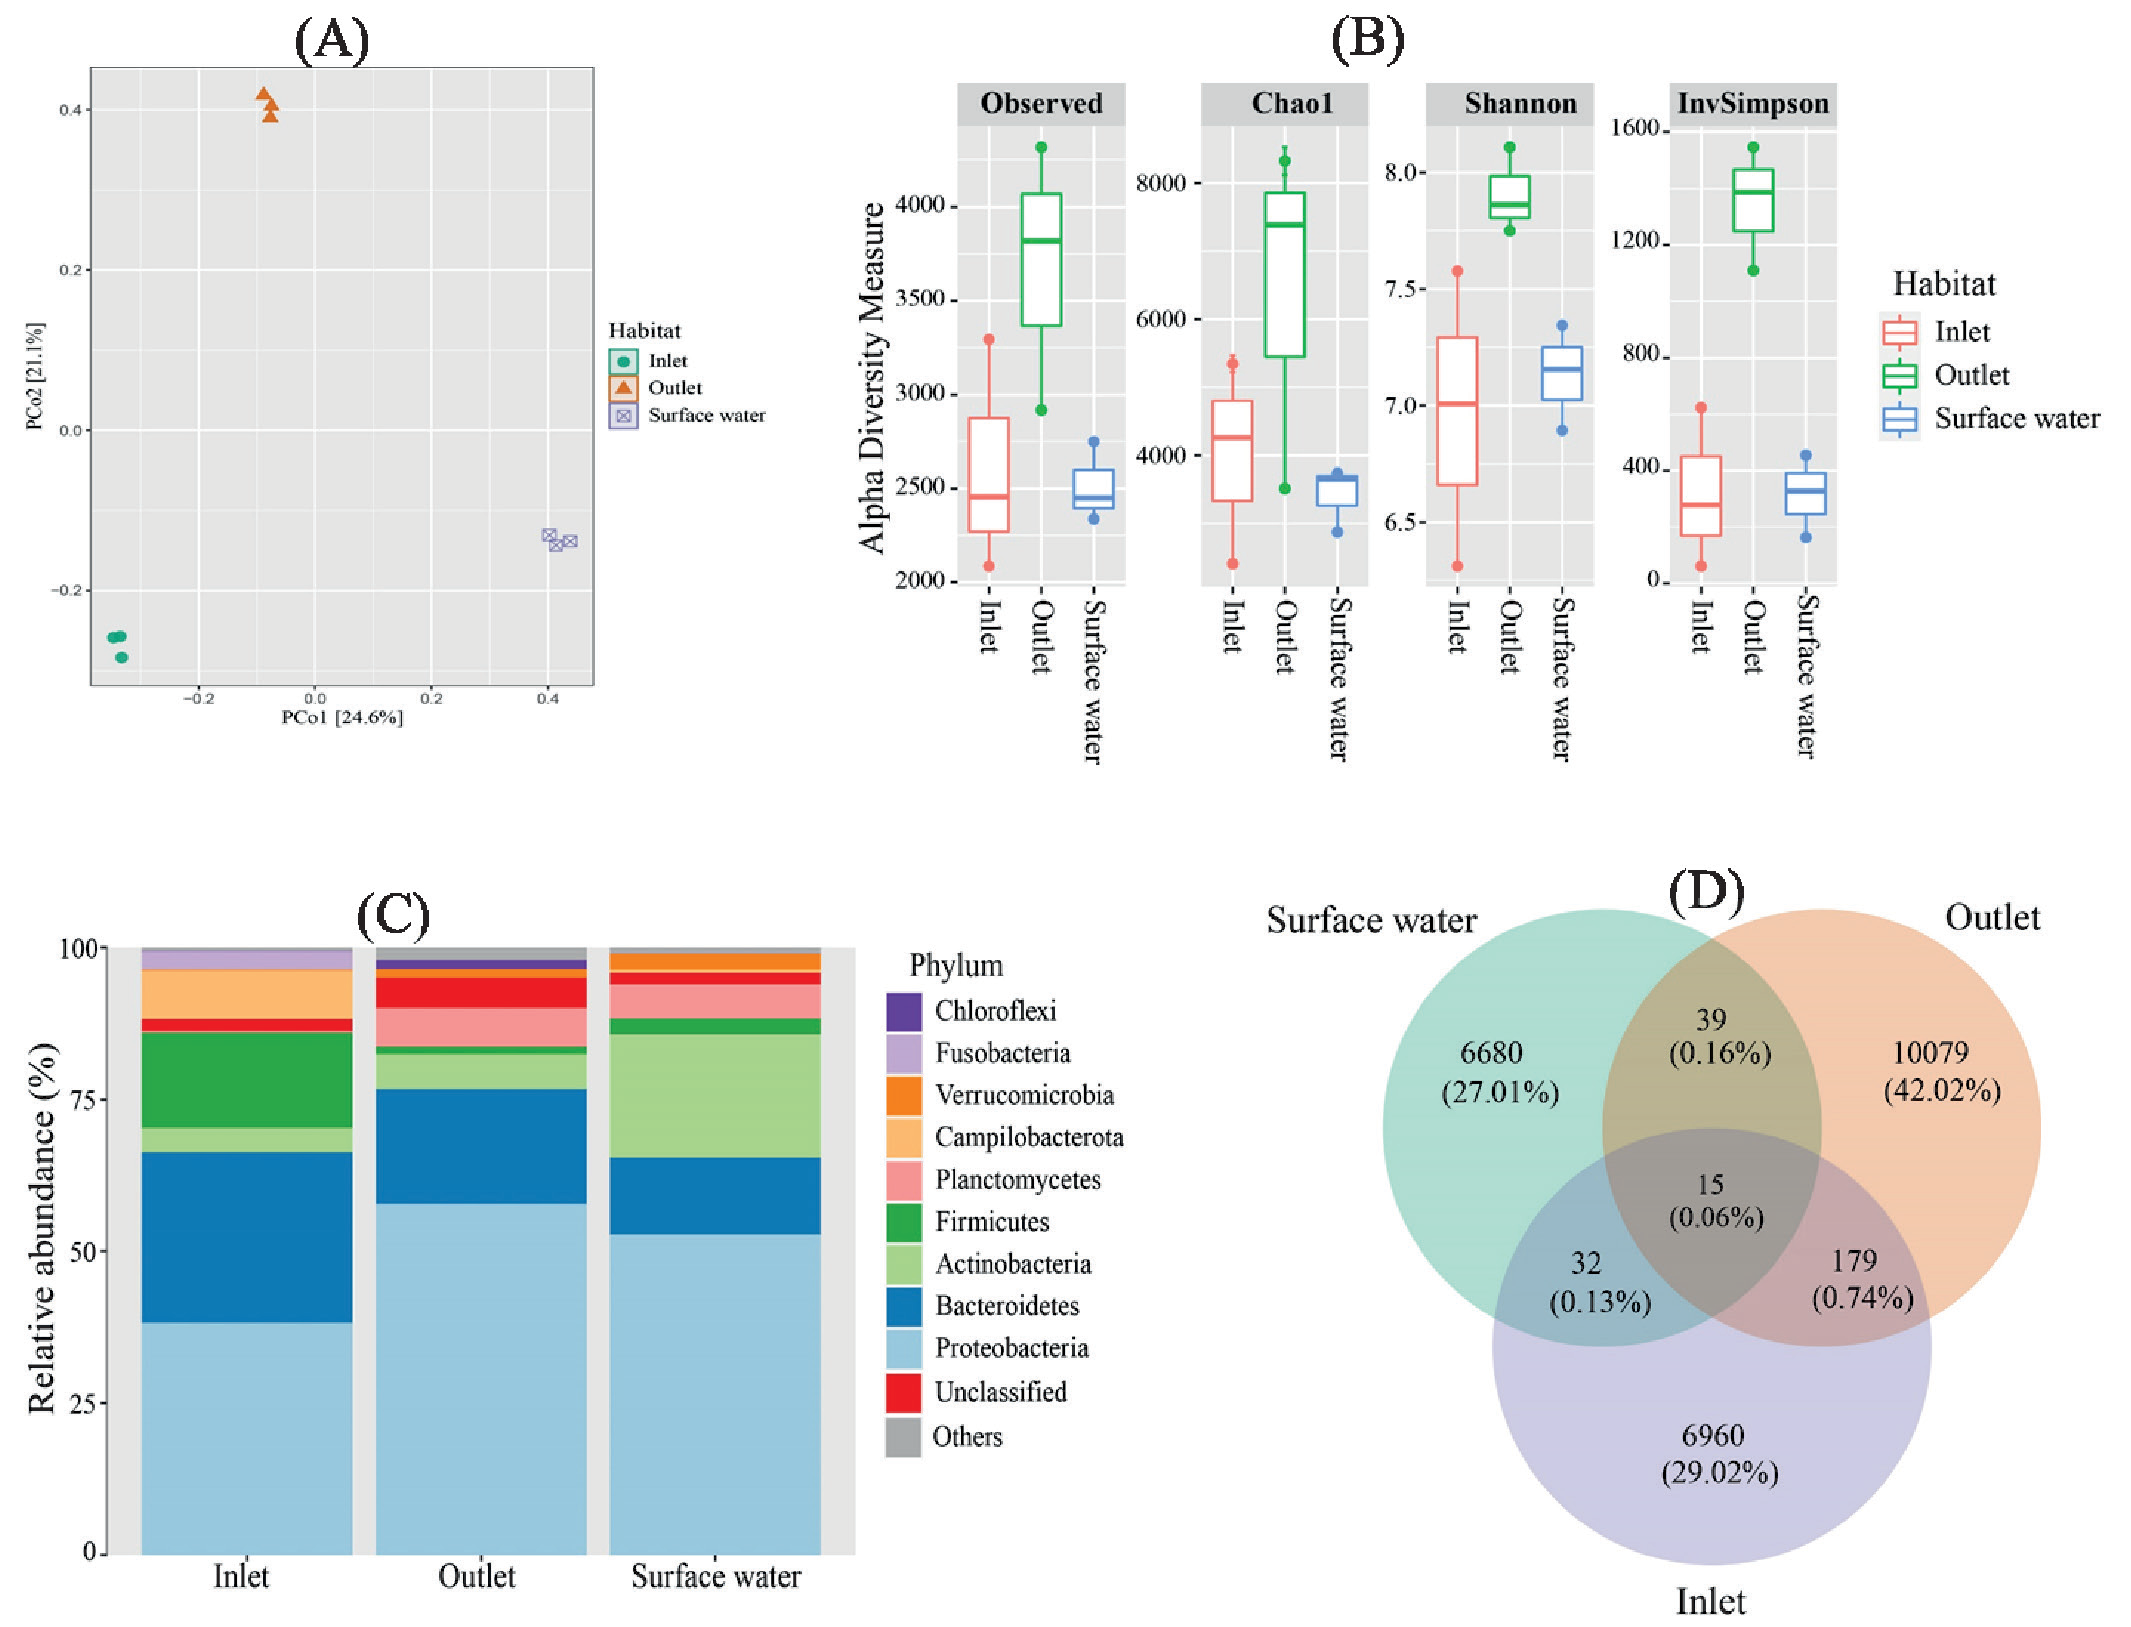
An environmental comparison study was conducted to examine the differences in microbial communities between three settings: the entrance and outlet of the Zenin WWTP, as well as an urban site at the Nile River. The PCoA plot showed that the microbial population structure varied between the three settings as depicted in (Fig. 1a). Prokaryotic populations dramatically varied according to habitat. The observed patterns were statistically validated by PERMANOVA and ANOSIM results (P<0.01). Cluster Analysis further corroborated these findings, as depicted in Fig. S1. The Venn diagram in Fig. 1d), illustrating shared percentages of 0.06% (n = 15) among the three tested habitats, further reinforces these results. While, 0.16% (n = 39) was the shared percentage between surface water and outlet stage, 0.74% (n = 179) was the shared percentage between inlet and outlet stages and 0.13% (n = 32) was the shared percentage between inlet and surface water. The habitats contained a large unique species as follows, outlet (n = 10079), inlet (n = 6960), and surface water (n = 6680). It is important to note that the microbial communities in the surface water of the river can vary due to multiple factors, including natural processes, land use patterns, and seasonal variations [28, 29]. However, the distinct differences observed between the inlet and outlet of WWTP and the river water indicate the substantial influence of wastewater treatment processes on the microbial composition [30-32]. The wastewater treatment plant’s outlet showed higher microbial diversity than inlet as well as the surface water from the Nile River (Fig. 1b). The higher diversity of microbial communities in the outlet of the wastewater treatment plant indicates that the treatment process effectively removes pollutants and facilitates the growth of diverse microbial communities [33, 34]. Previous studies have reported similar findings, supporting the idea that wastewater treatment processes can enhance microbial diversity in effluent. For instance, a study by Płaza et al. [33] investigated the microbial communities in WWTP and found that the treated effluent displayed different microbial diversity compared to the influent. The higher microbial diversity observed in the outlet of the wastewater treatment plant has significant implications for ecosystem health. Diverse microbial communities play essential roles in various ecological processes, including nutrient cycling and pollutant degradation. The presence of diverse microbial populations in the treated effluent suggests that the treatment process not only removes pollutants but also supports the growth of beneficial microbial species capable of performing important ecosystem functions.
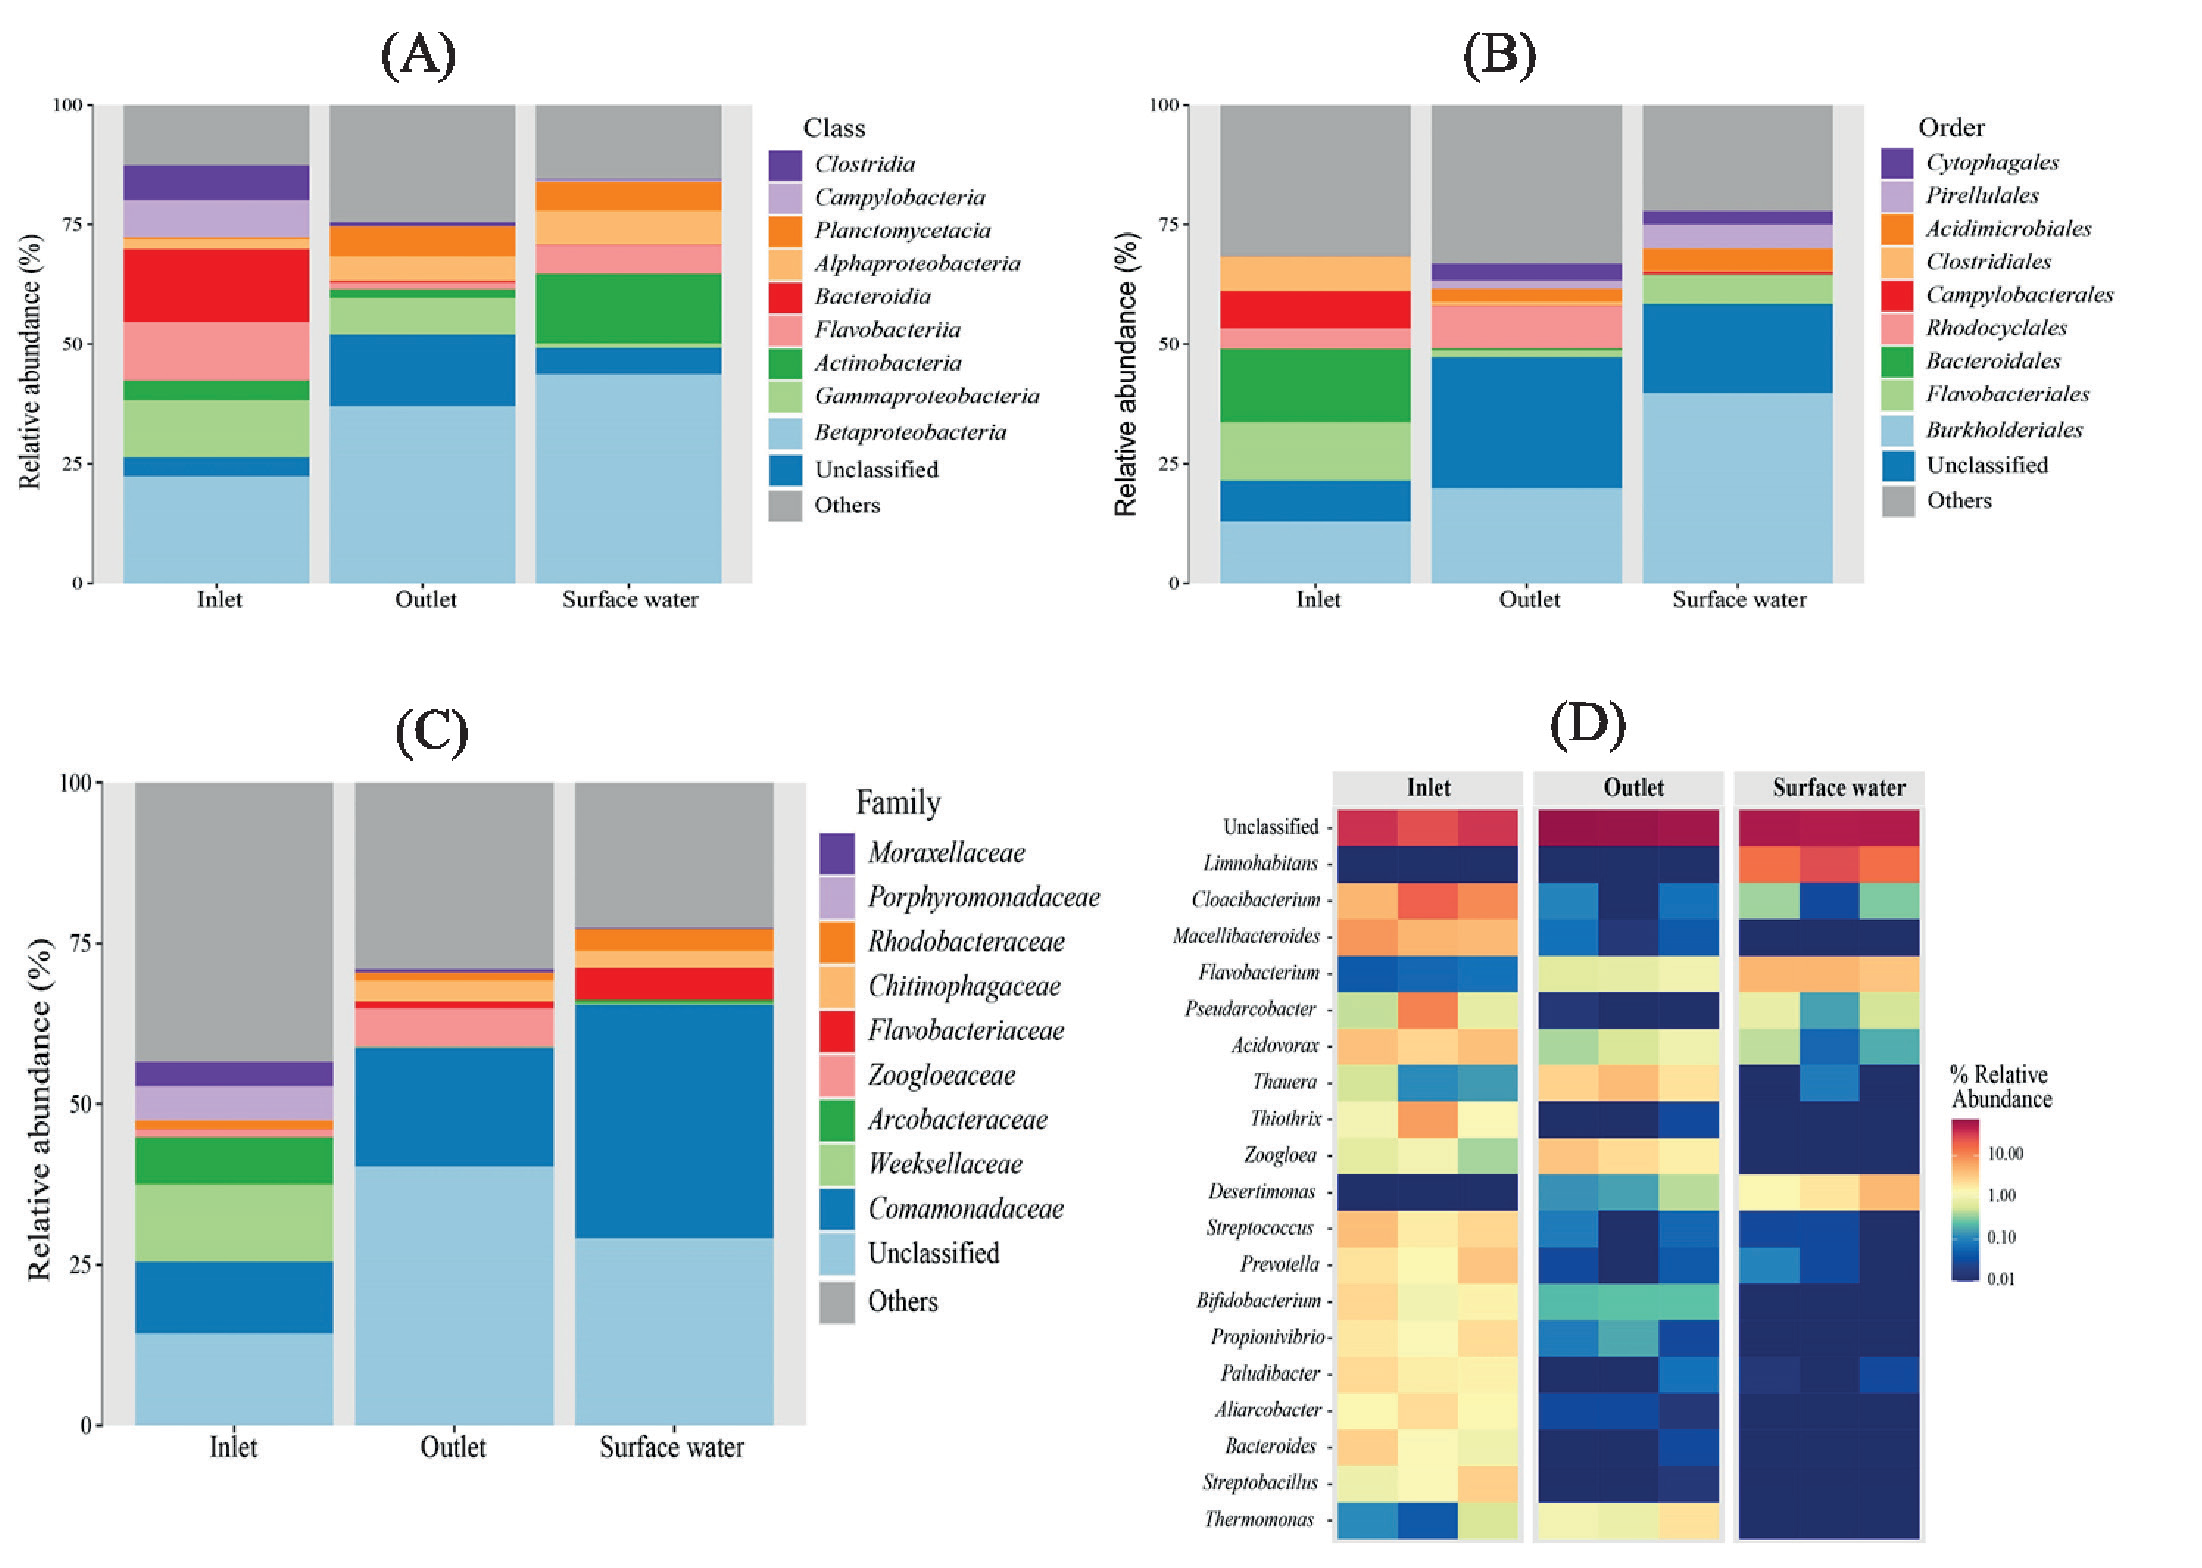
In the current study, the abundance of different phyla in different tested habitats were illustrated in Fig. 1c), where Proteobacteria slightly increased in outlet and surface water (>50%) compared to inlet (45%). This finding suggests that the treatment process employed in the wastewater treatment plant effectively removes pollutants and creates conditions that favor the growth and persistence of Proteobacteria. Proteobacteria are known to possess diverse metabolic capabilities and play crucial roles in various biogeochemical cycles, including nutrient cycling and pollutant degradation. According to existing literature, municipal WWTPs are predominantly populated by the phylum Proteobacteria. Additionally, other groups such as Bacteroidetes, Actinobacteria, and Firmicutes are also commonly found in these environments. However, the relative abundance of these groups can vary depending on multiple factors, including the type of sewage treatment plants, employed technology, influent composition, and hydraulic configuration [35-38]. Actinobacteria have the highest abundance (>20%) in surface water compared to the other habitats. Actinobacteria are well-known for their ability to produce bioactive compounds and play important roles in organic matter decomposition [39]. The higher abundance of Actinobacteria in the surface water suggests that this habitat provides favorable conditions for their growth, potentially due to the availability of specific resources or unique physicochemical parameters. Firmicutes and Campilobacteria decreased in outlet stage compared to their abundance in inlet phase. Additionally, Chloroflexi could be seen in the outlet stage only with a small percentage (2%). The decrease in the relative abundance of these phyla suggests that the treatment process in the wastewater treatment plant may have selective effects on certain microbial groups. The removal of pollutants and changes in environmental conditions during the treatment process could contribute to the decrease in the abundance of Firmicutes [40]. Betaproteobacteria was the most common class in all habitats (i.e., inlet, outlet and surface water), where it typically showed the highest abundance (48%) in surface water (Fig. S2). Betaproteobacteria are known to encompass various ecologically important groups, including nitrifiers, denitrifiers, and iron oxidizers [41, 42]. Their prevalence in all tested habitats indicates their adaptability and ecological importance in these habitats. Bacteroidia class has a relative abundance of 12% of inlet followed by Gammaproteobacteria (10%), Falvobacteria (9%) and Camplyobacteria (5%) and these percentages were significantly reduced in the outlet level. However, it could be noticed that the relative abundance of Betaproteobacteria and Planctomycetacia slightly elevated in the outlet compared to the inlet stage. Besides, the relative abundance of Falvobacteria was highest in the inlet compared to outlet and surface water (Fig. S2).
Fig. 1.
a) PCoA based on Bray-Curtis similarity for mapping prokaryotes in different habitat. b) Alpha diversity indices for prokaryotes. c) Bar plot showing the relative abundance of the prokaryotes phyla in different habitats. d) Venn diagram showed the unique and shared prokaryotic OTUs.
In this investigation, the order of Burkholderiales was the most abundant order (40%) in surface water followed by Flavobacteriales, Pirellulales (5%), and Acidimicrobiales (4%). It could be noticed that Burkholderiales, Bacteriodales, Campylobacrerales, and Rhodocyclals were the most abundant orders in the outlet samples. While, Burkholderiales and Bacteriodales were the abundant in inlet stage (Fig. S2). Comamonadacea was the most abundant family (25%) in the surface water group in this work. While, Weeksekkaceae, Arcobacteraceae, Porphyromonadaceae and Moraxellaceae families dramatically decreased (P≤0.05) in the outlet stage of the tested treatment plant compared to the inlet stage. Comamonadaceae, Zoogloeaceae and Chitinophagaceae families were significantly elevated in the outlet relative to inlet stage (Fig. S2). A heatmap showed the predominance of the top 20 prokaryotic genera in the analyzed specimens of three habitats, where Limnohabitans, Flavobacterium, and Desertimonas were the most common genera in surface water. While, Cloacibacteriumm, Macellibacteroides, and Acidovorax were the most dominant genera in inlet of the WWTP. However, a two genera including Thauera (1-3%) and Thermomonas were dominant in the outlet stage (Fig. S2). It is well known that different habitats exhibited distinct microbial communities compositions [43-45].
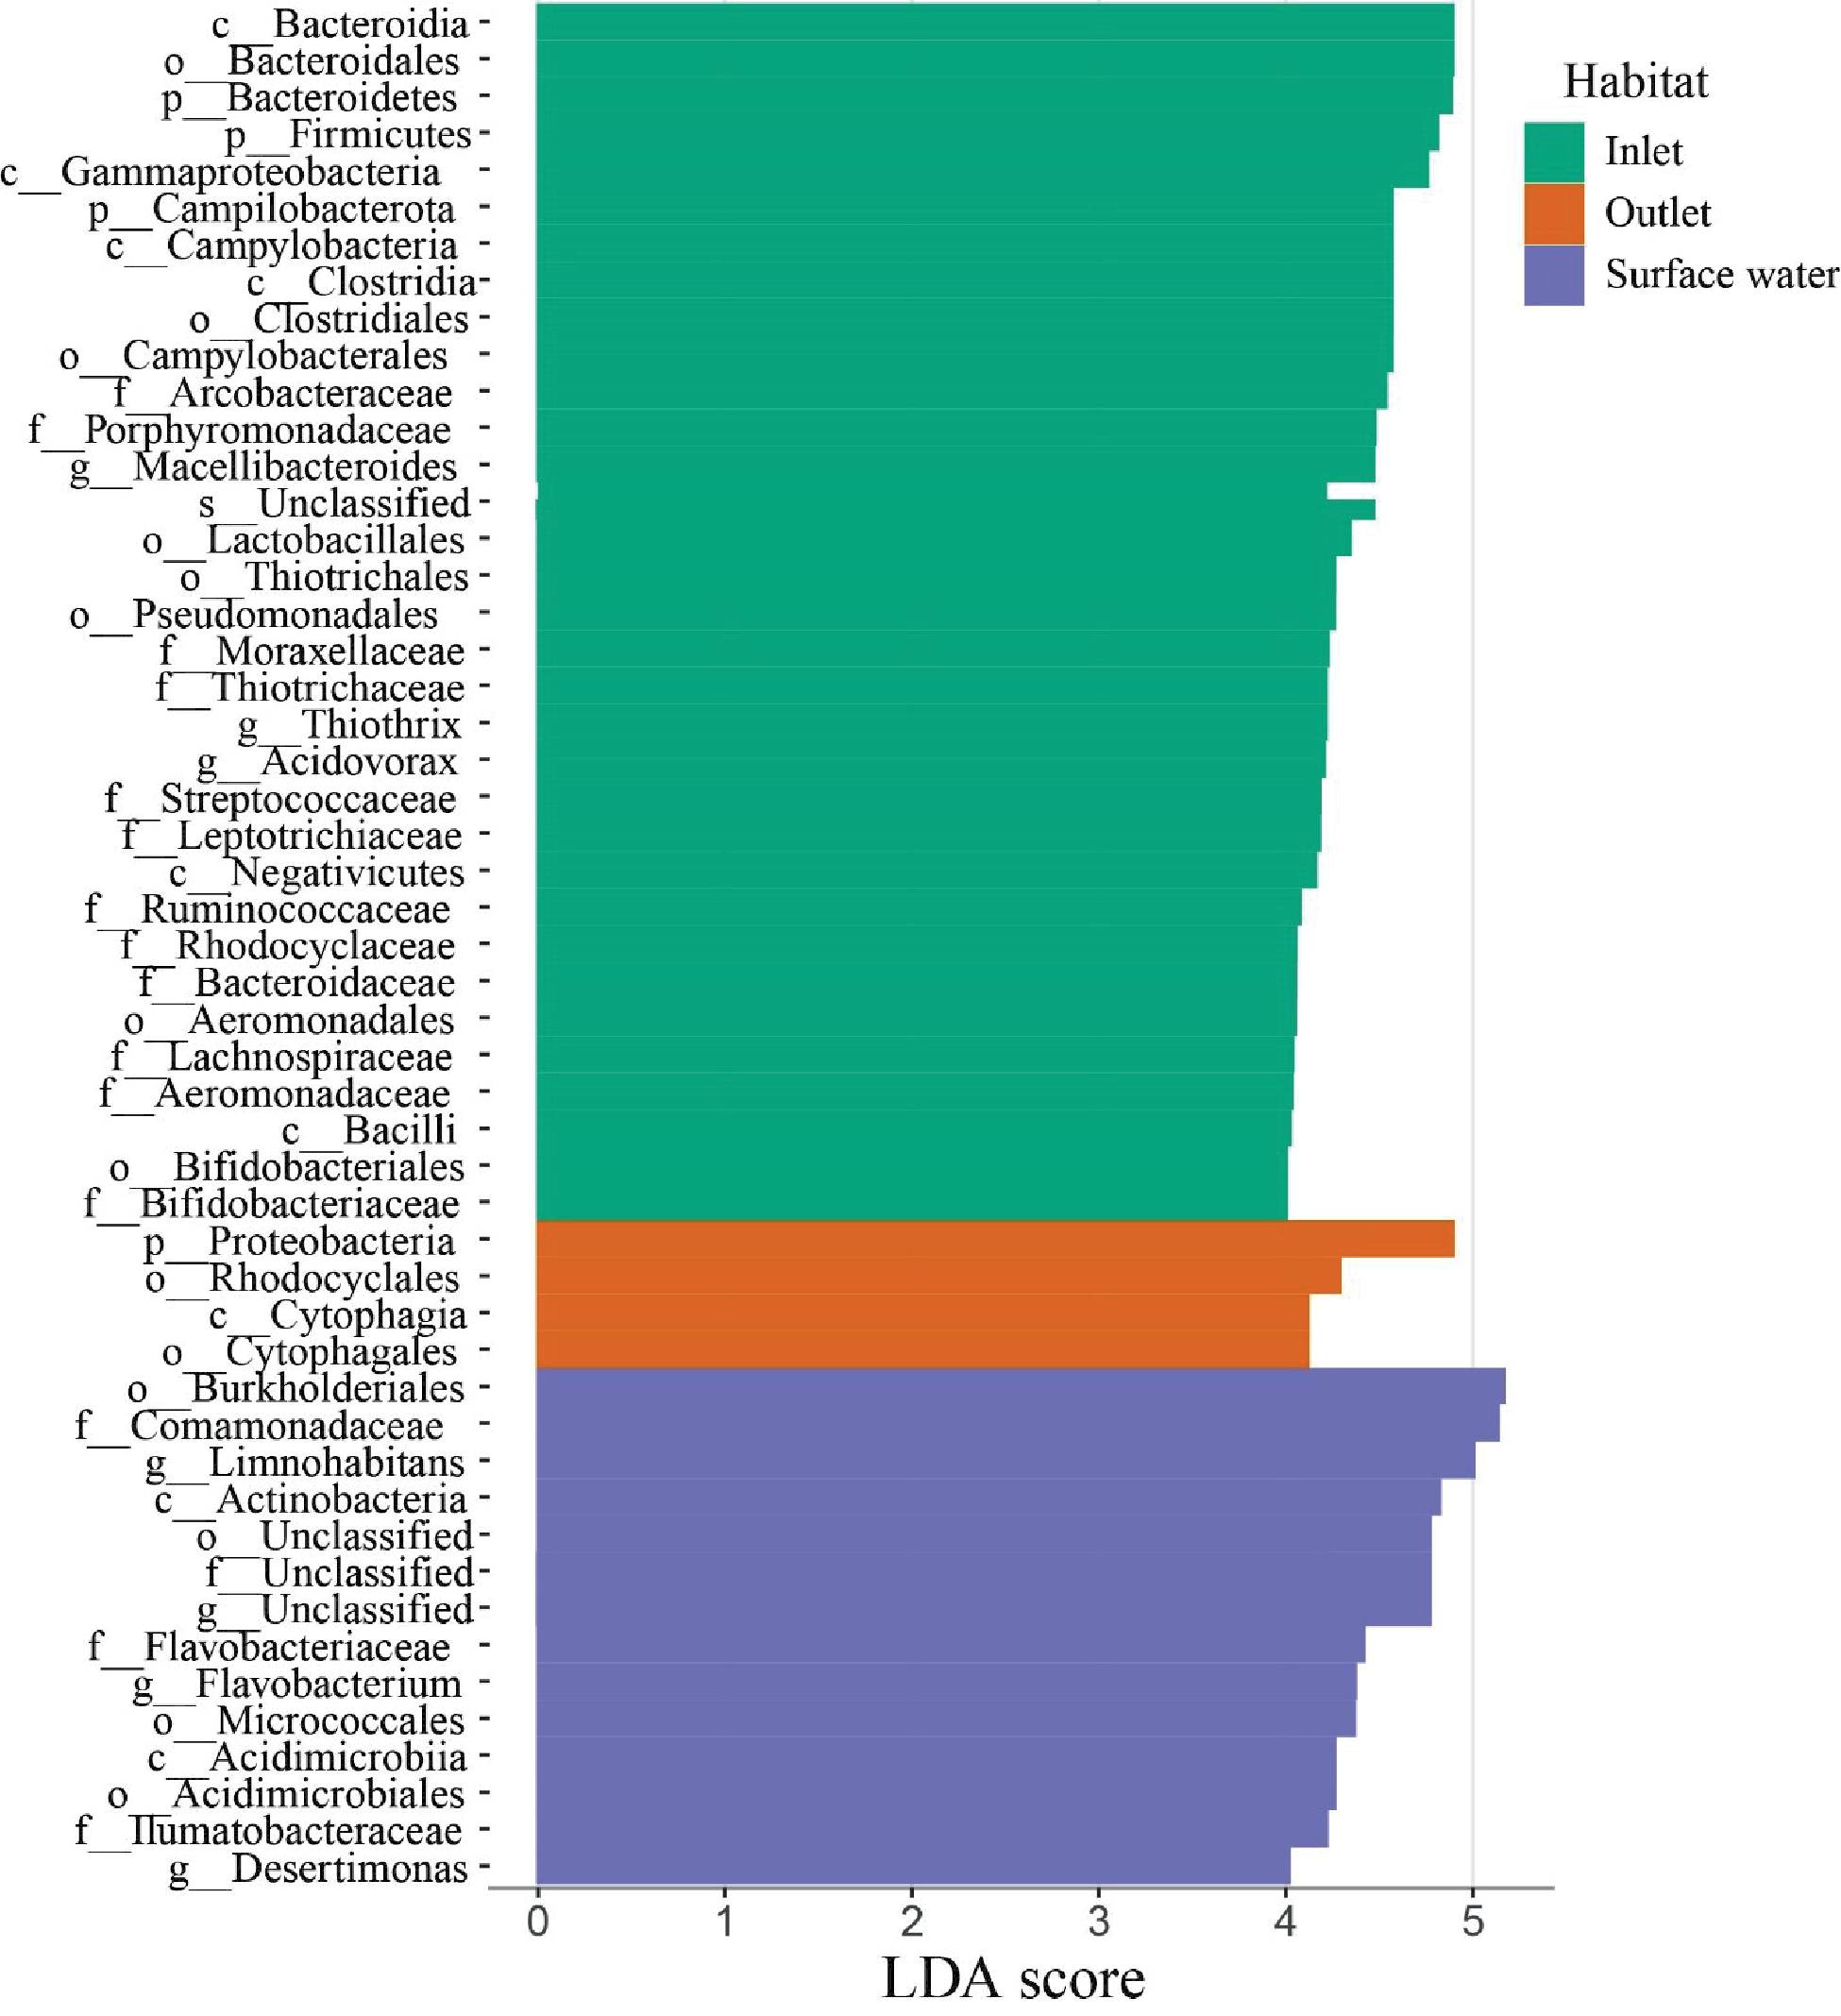
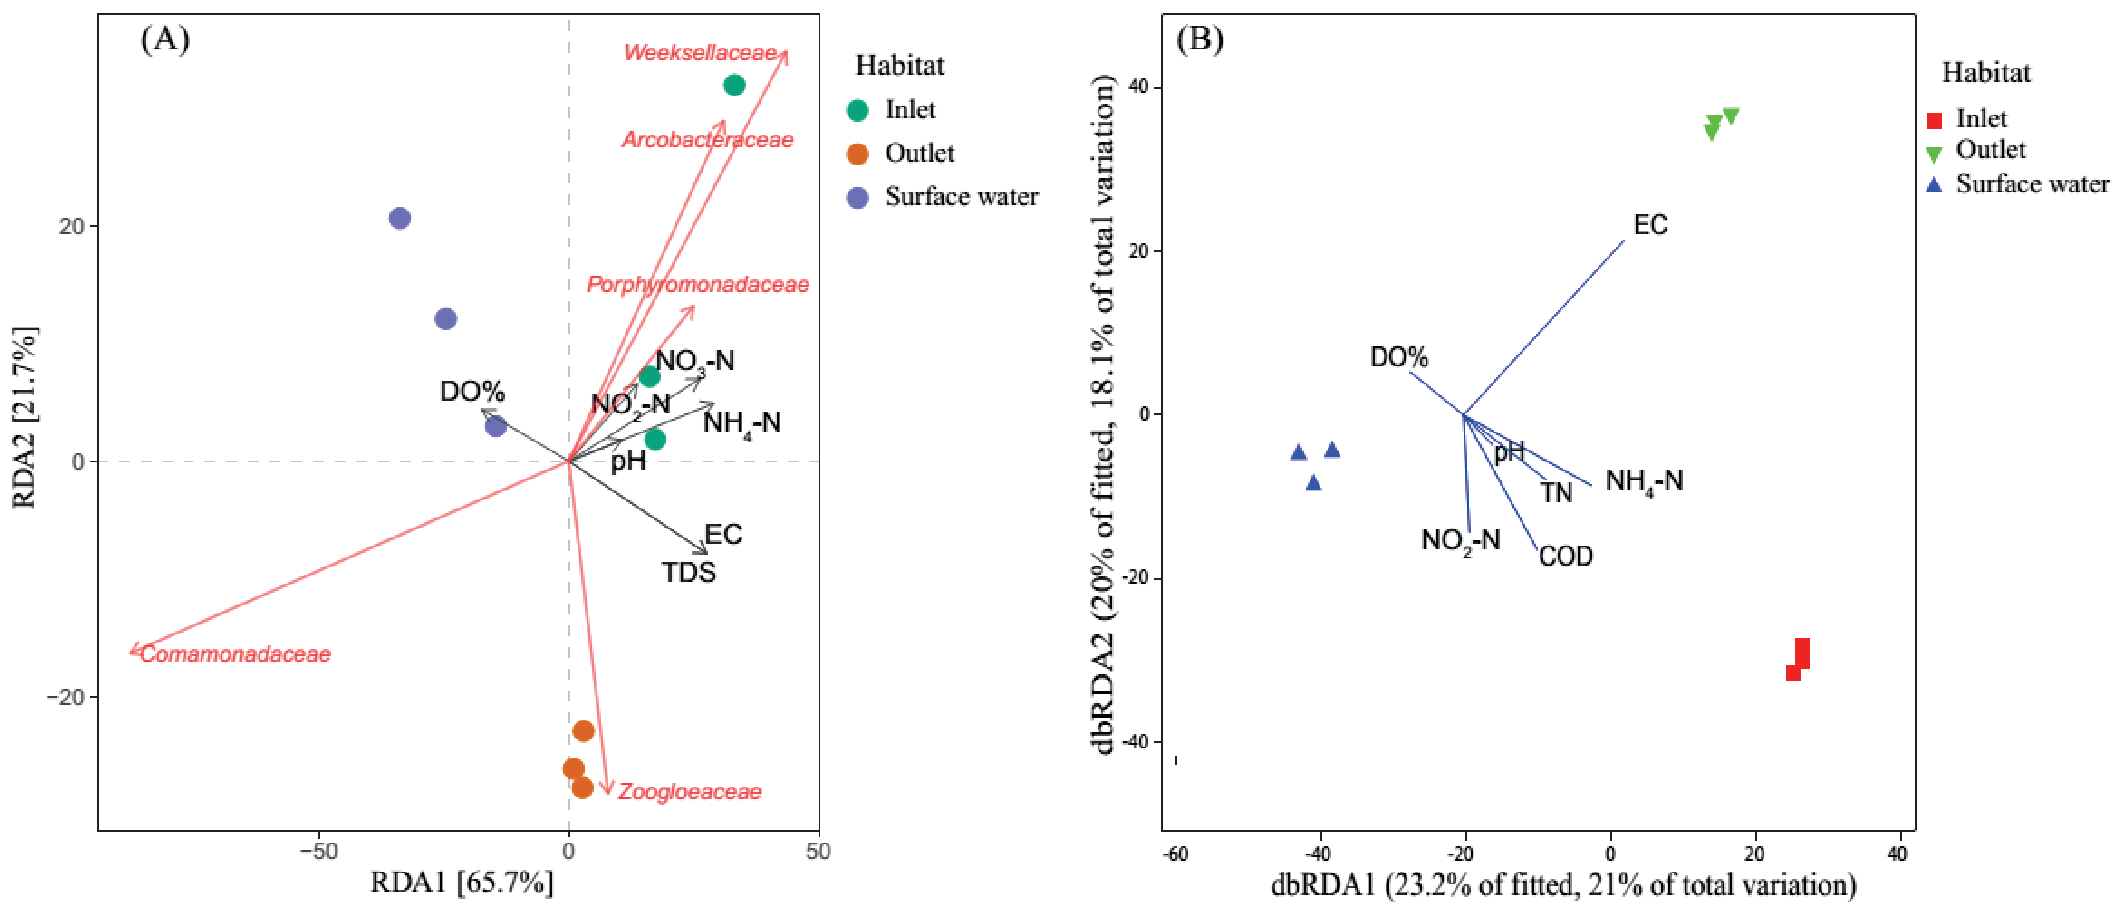
The distance-based linear model (DistLM) indicated that EC and NH4-N were the most significant variables in explaining the variation in microbial communities. The results also showed that pH, DO%, COD, NO2-N, and TN had lesser impacts on the microbial communities (Table 1). Kaestli et al., [46] and Kim et al., [47] reported that EC explained a large proportion of microbial communities variations in different environments. In this study, the influence of environmental factors on prokaryotic families across three groups of samples (inlet, outlet, and surface water) was elucidated using RDA, as shown in Fig. 2. The RDA axes, RDA1 and RDA2, accounted for 65.7% and 21.7% of the total community variation, respectively. It could be noticed that Prophyromonadaceae was correlated positively with the nutrients (e.g., NO2-N). The dbRDA plot showed that the three habitats clustered separately, indicating dissimilarity in the microbial communities between these habitats. The dbRDA environmental vectors revealed a strong association between the outlet microbial communities and EC. Conversely, the inlet microbial communities exhibited a strong association with organic pollution factors (COD) as well as inorganic pollution factors (NO2-N and NH4-N) (Fig. 2). The analysis of environmental parameters and microbial structure through RDA confirms the impact of TN, NH4-N, and NO3-N pollution on microbial communities [48]. In the present study, Linear discrimination analysis (LDA) effect size (LEfSe) is particularly useful in microbiome research as it allows for the identification of microbial biomarkers that are indicative of specific conditions. Bacteroidia and Bacteroidales was enriched in polluted condition as in the inlet of WWTP, when the condition become better, Protobacteria was the most abundant in the outlet of WWTP. While the LDA score for Burkholderiales and Comamonadaceae was the highest in surface water (Fig. 3).
Table 1.
Sequential tests showed the cumulative proportion of the variation of the microbial communities explained using forward selection by fitting the variables sequentially with 9999 permutations.
| Variable | Pseudo-F | P | Prop. | Cumul. |
|---|---|---|---|---|
| +EC | 1.84 | 0.010 | 0.21 | 0.208 |
| +NH4-N | 1.76 | 0.001 | 0.18 | 0.388 |
| +pH | 1.06 | 0.507 | 0.11 | 0.495 |
| +DO% | 1.02 | 0.510 | 0.10 | 0.598 |
| +COD | 1.03 | 0.481 | 0.10 | 0.700 |
| +NO2-N | 1.02 | 0.467 | 0.10 | 0.802 |
| +TN | 1.04 | 0.463 | 0.10 | 0.901 |
Conclusions
Employing next-generation sequencing technology facilitated a deeper and more comprehensive insight into the structure of microbial communities and the influence of environmental factors Moreover, the study highlights the importance of environmental factors such as EC and NH4-N in shaping the microbial community structure. LDA indicated that Bacteroidia and Bacteroidales were enriched in polluted inlet conditions of WWTP, while Protobacteria dominated the outlet. Burkholderiales and Comamonadaceae had the highest LDA score in surface water. Overall, this study has contributed to the understanding of the microbial ecology of wastewater treatment plants and river in Egypt and provides valuable information for the maintenance of ecosystem function.