

Introduction
The root-knot nematodes (RKN) disease induced by Meloidogyne spp. was widespread and caused a significant economic impact on a global scale [1]. Among the numerous pathogens associated with RKN, Meloidogyne incognita is considered to be the most severe pathogeny. It could infect over 3000 plant species, including food crops, cash crops, vegetables, fruit trees, ornamental plants, and weeds [2]. The second-stage juveniles (J2s) of M. incognita predominantly initiate infection in plant roots. This phenomenon results in the impairment of the plant's nutrient assimilation capabilities, thereby impeding the growth and development of the aboveground parts, ultimately leading to stunted growth, premature senescence, and potentially complete desiccation and plant death.
The rhizosphere soil of plants is the main site for J2 activity and infection. The rhizosphere, which serves as the interface between plants and soil, is characterized by dense microbial communities and metabolites [3]. Plants utilize their root system to acquire nutrients and energy from the soil, facilitating their metabolic needs for their growth and development. On the other hand, the roots and exudates may attract a wide range of microorganisms in the rhizosphere by means of nutrient and mucilage secretion, among other aspects [4]. The root exudates of plants infested by M. incognita may be altered in composition, resulting in the attraction of a greater number of advantageous microbes. Scientific investigations have revealed that a range of compounds, such as coumarins, triterpenes, flavonoids, benzoxazinoids, phytohormones, and others, can stimulate or inhibit the proliferation of specific microbes that are in close proximity to the host plants [5-7]. Finkel et al. have demonstrated that Variovorax bacteria can regulate Arabidopsis root growth by influencing the levels of plant hormones auxin and ethylene [8]. Plenty of evidence is also present for the ability of plants to regulate their microbiome by the exudation of various metabolites, pointing to potentially reciprocal relationships between the microbiome and metabolome of the host plant [9]. Therefore, it is possible to propose isolating chemical compounds and microorganisms with biocontrol effects from rhizosphere soil.
Exploring new biocontrol factors from the rhizosphere micro-environment of plants is currently a hot topic in RKN biocontrol, as demonstrated by the extensive current research efforts [10, 11]. The microorganisms in the rhizosphere's soil directly or indirectly influence plant metabolism [9] and are a potential source for developing novel biocontrol agents against M. incognita. Research has shown that differences in microbial communities' composition and the abundance of functional genes and metabolites have been found in soils with high and low M. incognita infestation levels [12]. When M. incognita J2s infected tobacco, the relative abundance of certain rhizosphere bacterial genera and genes related to metabolic and signal transduction functions significantly increased [13]. Therefore, it is very feasible to screen biocontrol factors with potential by utilizing plant rhizosphere metabolomics and microbial diversity.
It can be observed that the presence of M. incognita in plants may alter the metabolite profiles and microbial community structures within the rhizosphere soil. Accordingly, the objective of this study was to identify the metabolites and microorganisms with potential control for RKN based on examining the alterations in the rhizosphere soil of tomato plants employing metabolomic profiling and high-throughput sequencing. The results will establish a theoretical basis for accelerating the implementation of innovative biocontrol practices and pesticides.
Materials and Methods
Experimental Materials
The initial M. incognita population was obtained from the Academy of Agriculture and Forestry Sciences of Beijing, China, and subsequently propagated using living tomato plants in a laboratory incubator. The Qinshu Shanghai 908 tomato cultivar, which was procured from Xi'an Qinshu Agriculture Company Limited, was used as the plant material in this study. The experiment was conducted in 2021 at the Agricultural Environmental Ecology Research Laboratory of Hebei Innovation Center for Bioengineering and Biotechnology of Hebei University, China (38°87' N,115°51' E).
Rhizosphere Soil Samples
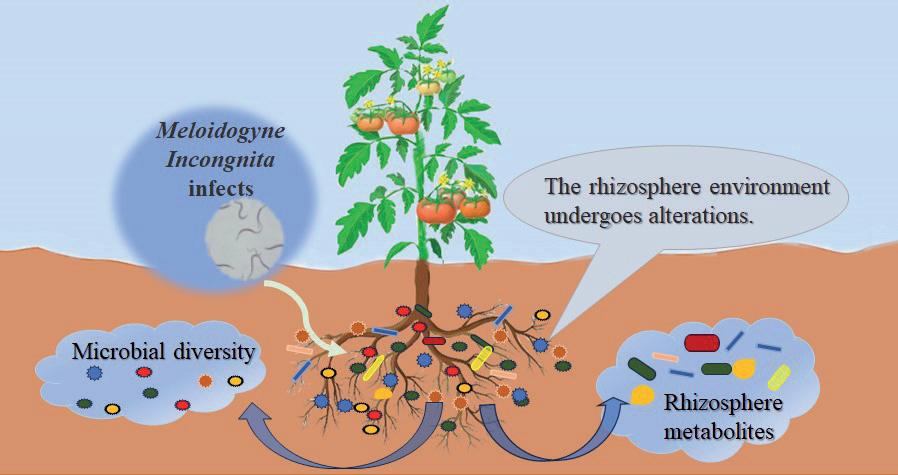
Four tomato seeds were sown in each of the plastic pots, with dimensions of 11 cm in height and diameter, filled with 250 g of soil obtained from the Baoding urban area of Hebei Province, China. The soil contained 52.90 ± 0.22 mg/kg of total nitrogen, 16.44 ± 0.11 mg/ kg of available phosphorus, 132.55 ± 1.29 mg/kg of total potassium, and 0.22 ± 0.01% organic matter. The tomato plants were grown in a controlled environment chamber specifically designed for climate simulation through artificial intelligence. Inoculation with M. incognita was performed once the tomato seedlings had reached the 3-4 leaf stage. To introduce the nematodes, small holes were carefully created in the soil near the roots using a glass rod, and a suspension containing 2000 M. incognita J2s was then infused using a pipette. After a period of 35 days, soil samples (Nematode-infested Soil, NS) were collected from the rhizosphere-surrounding area. Rhizosphere soil of tomato plants without the presence of nematodes was also collected to be used as the control (Healthy Soil, HS) (Fig. 1). Three replicates were obtained for each soil sample. The samples were stored in a -80℃ freezer for DNA and metabolite extraction (performed one week after sampling) and were sent to Shanghai Majorbio Bio-pharm Technology Co., Ltd. (China) for metabolomics and high-throughput DNA sequencing.
Metabolomics Analysis
Soil samples weighing 50 mg were accurately measured and subjected to metabolite extraction using a methanol: water (4:1, v/v) solution at a volume of 400 µL. The resulting soil suspension was allowed to settle at a temperature of -20 ℃ and subsequently processed using a high-throughput tissue grinder, Wonbio- 96c (Shanghai Wanbo Biotechnology Co., Ltd), at a frequency of 50 Hz for 6 minutes. This was followed by vortexing for 30 seconds and ultrasonication at a frequency of 40 kHz for 30 minutes at a temperature of 5 ℃. To precipitate proteins, the samples were placed at -20 ℃ for 30 minutes. Subsequently, centrifugation was performed at 13000 g and 4 ℃ for 15 minutes, and the resulting supernatant was carefully transferred to sample vials for LC-MS/MS analysis. After UPLC-TOF/MS analyses, the raw data were imported into Progenesis QI 2.3 (Nonlinear Dynamics, Waters, USA) for peak detection and alignment. The mass spectra features were identified by querying the accurate mass, MS/MS fragment spectra, and isotope ratio difference information in reliable biochemical databases such as the Human Metabolome Database (HMDB) (http:// www.hmdb.ca/) and Metlin Database (https://metlin. scripps.edu/). Principle Component Analysis (PCA) was employed to comprehensively characterize the metabolic data, enabling visualization of treatment clustering, trends, and outliers. Orthogonal partial least squares discriminate analysis (OPLS-DA) was utilized for statistical analysis to identify global metabolic alterations between the different treatment groups. Univariate analysis (T-test) was employed to identify differential metabolites between the groups.
High-Throughput DNA Sequencing
The total DNA of the microbial communities in the soil samples was extracted using a DNA extraction kit (E.Z.N.A ® Soil DNA Kit). The purity, concentration, and integrity of the extracted DNA were assessed using a microspectrophotometer and 1% agarose gel electrophoresis (voltage 5 V/cm, time 20 min). Primers 338F/806R and ITS1F/ITS2R (Table 1) were used to amplify the V3 to V4 variable regions of the bacterial 16S rRNA gene and the ITS1 region in fungi, respectively. The 16S rRNA gene PCR amplification was performed as follows: 95 ℃ for 3 min, 27 cycles (95 ℃ denaturation for 30 s, 55 ℃ annealing for 30 s, 72 ℃ extension for 30 s), final extension at 72 ℃ for 10 min, and a hold step at 4 ℃ (PCR instrument: ABI GeneAmp ® 9700 type). The PCR mixture contained 5 × TransStart FastPfu buffer 4 μL, 2.5 mM dNTPs 2 μL, forward primer (5 μM) 0.8 μL, reverse primer (5 μM) 0.8 μL, TransStart FastPfu DNA Polymerase 0.4 μL, template DNA 10 ng, and ddH2O up to 20 μL. PCR reactions were performed in triplicate. The PCR product was extracted from 2% agarose gel and purified using the AxyPrep DNA Gel Extraction Kit (Axygen Biosciences, Union City, CA, USA) according to the manufacturer's instructions and quantified using a Quantus™ Fluorometer (Promega, USA).
Table 1.
Primers used for PCR amplification.
| Name | Forward primer sequence (5’-3') | Reverse primer sequence (5’-3') |
|---|---|---|
| Bacterial 16s region | ACTCCTACGGGAGGCAGCAG | GGACTACHVGGGTWTCTAAT |
| Fungal ITS1 region | CTTGGTCATTTAGAGGAAGTAA | GCTGCGTTCTTCATCGATGC |
The original amplicon sequences were processed and filtered with Fastp software (version 0.19.6) and Flash software (version 1.2.11) to obtain the final high-quality sequences. Duplicate sequences were removed using Uparse software (version 7.0.1090). The identified non-repetitive sequences were assigned to operational taxonomic units (OTUs) based on 97% similarity. Chimeras were removed, and the sequence with the highest abundance in each OTU was selected as a representative sequence of that OTU for subsequent bioinformatics analysis.
Microbial Community and Differential Abundance Analysis
OTUs were normalized based on the minimum sum count across the given OTU. The taxonomic annotation of each sequence was performed by alignment against the Silva 16S rRNA database using the RDP classifier software (version 2.11), with an alignment threshold of 70% to identify the microbial community composition of each sample at the phylum and genus levels, respectively. The alpha diversity metrics Chao index, which reflects community richness, and the Shannon index, which reflects community diversity, were calculated using Mothur software (version 1.30.2). Beta diversity analysis was performed using R language (version 3.3.1). Bray Curtis distances were calculated for the distance matrix, and PCoA was used to visually display the results. The stats package in the R language and the SciPy package in Python were used for the analysis of microbial community composition and the different abundances of different taxonomic groups.
Data Analysis
The SciPy (Python) software (Version 1.0.0) was employed to examine the association between differential metabolites and microbial diversity in soil samples. To compare the mean values between groups, T-tests and one-way analysis of variance were conducted using the SPSS software, and Tukey's HSD method was employed for statistical significance testing at a significant level of 0.05 [14].
Results
Differentially Accumulated Metabolites of Rhizospheric Soils
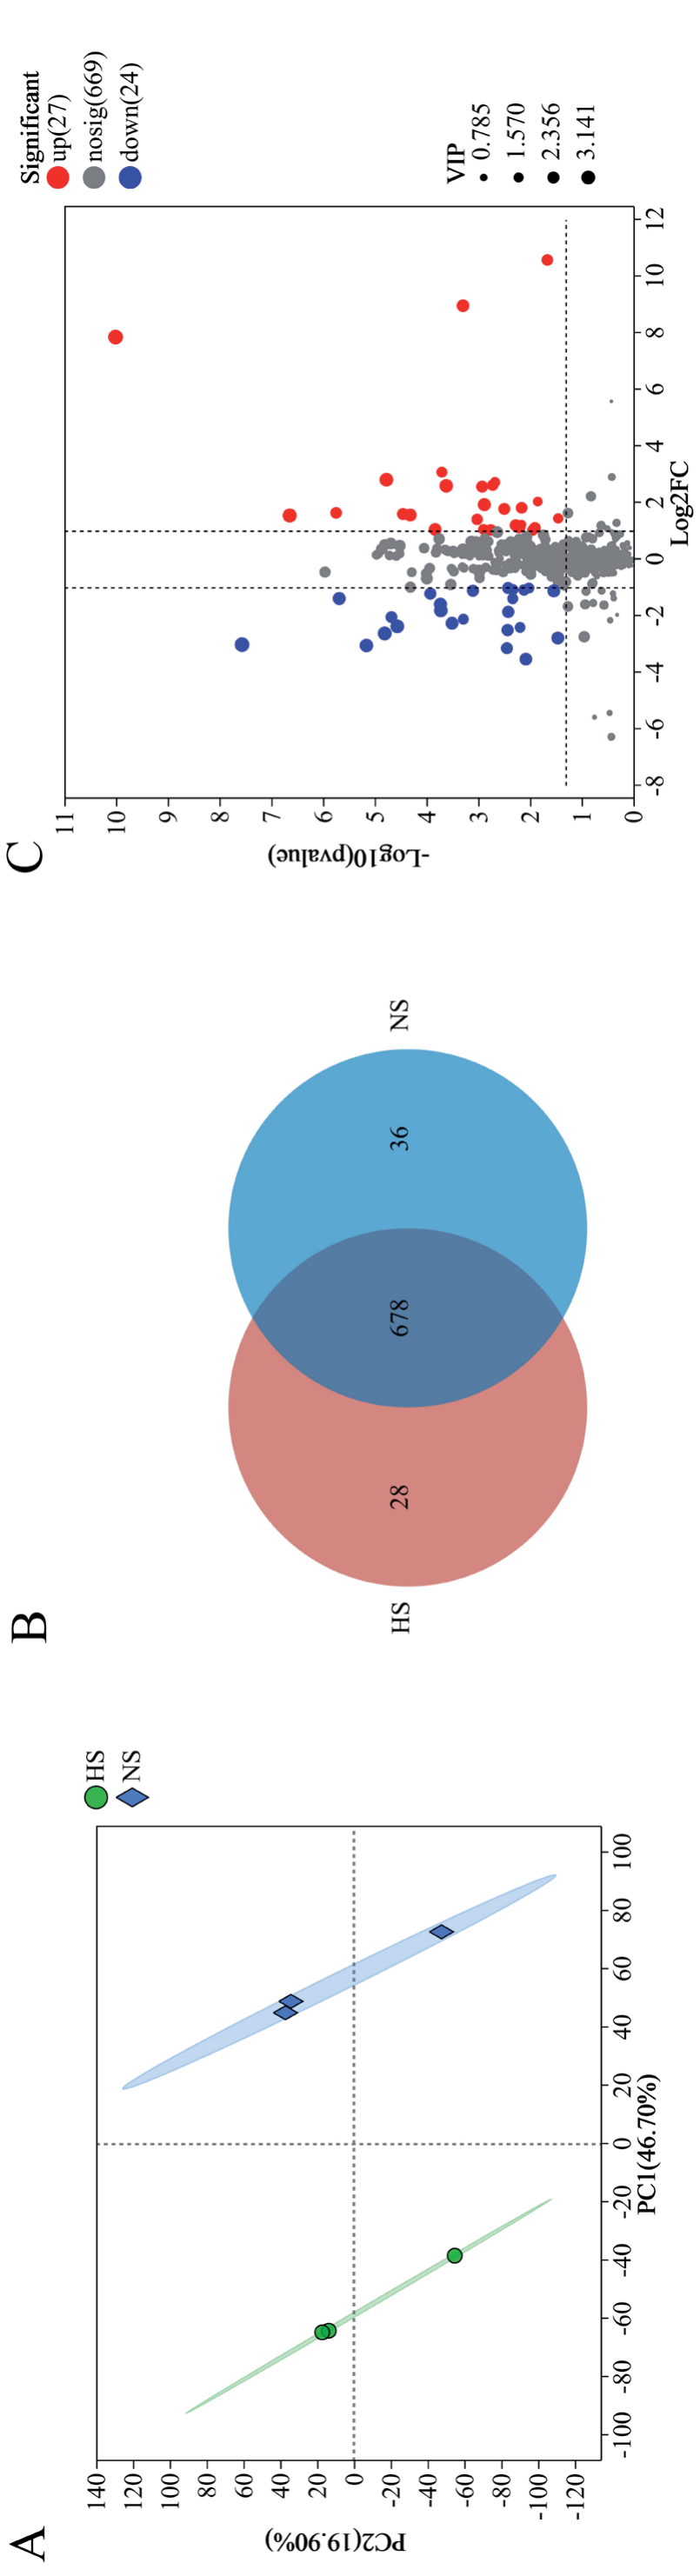
The unsupervised principal component analysis (PCA) was employed to elucidate the metabolite differences between samples, with PC1 and PC2 accounting for 46.70% and 19.90% of the variance, respectively. A significant difference was observed in the total metabolite composition of the individual samples from the two groups based on their distribution across the PC axes. Notably, the individual samples from each treatment group, corresponding to the metabolite profiles from the rhizospheric soils before and after M. incognita infestation, were clustered (Fig. 2A). Venn analysis of the metabolites in healthy and infested soil, which numbered 706 and 714, respectively, revealed that 678 metabolites were shared between the two conditions. Additionally, 28 metabolites were unique to healthy soil, while 36 were exclusive to infested soil (Fig. 2B). A volcano plot was generated based on T-tests, where significant differentially accumulated metabolites were defined as having a VIP score > 1, a P-value < 0.05, and 0.5<fold change<2 (Fig. 2C). The volcano plot analysis revealed that 51 metabolites exhibited significant differential accumulation, while 27 metabolites were upregulated (Table 2) and 24 metabolites were downregulated (Table 3).
Differences in Microbial Community Composition
Following the optimization of the original bacterial sequences, a total of 296,094 sequences were obtained, with an average sequence length of 417.24 base pairs (bp). A total of 29,560 OUTs were obtained through cluster analysis, with species annotated to 39 phyla, 102 orders, 2,480 families, 688 genera, and 1,299 species. The original fungal sequence was optimized to yield a total of 356,980 sequences, with an average sequence length of 260.90 bp. Cluster analysis yielded 518 OUTs. The species were annotated to 7 phyla, 26 orders, 62 families, 208 genera, and 329 species.
Regarding the alpha diversity analysis, as shown in Table 4, the Shannon index of the bacterial community in healthy soil was comparatively greater than in infested soil. Conversely, the Shannon index of the fungal community of the healthy soil was comparatively lower than that of the infested soil. These results suggest variability in the richness and diversity of microorganisms between healthy and nematode-infested soils, although none of the differences were statistically significant.
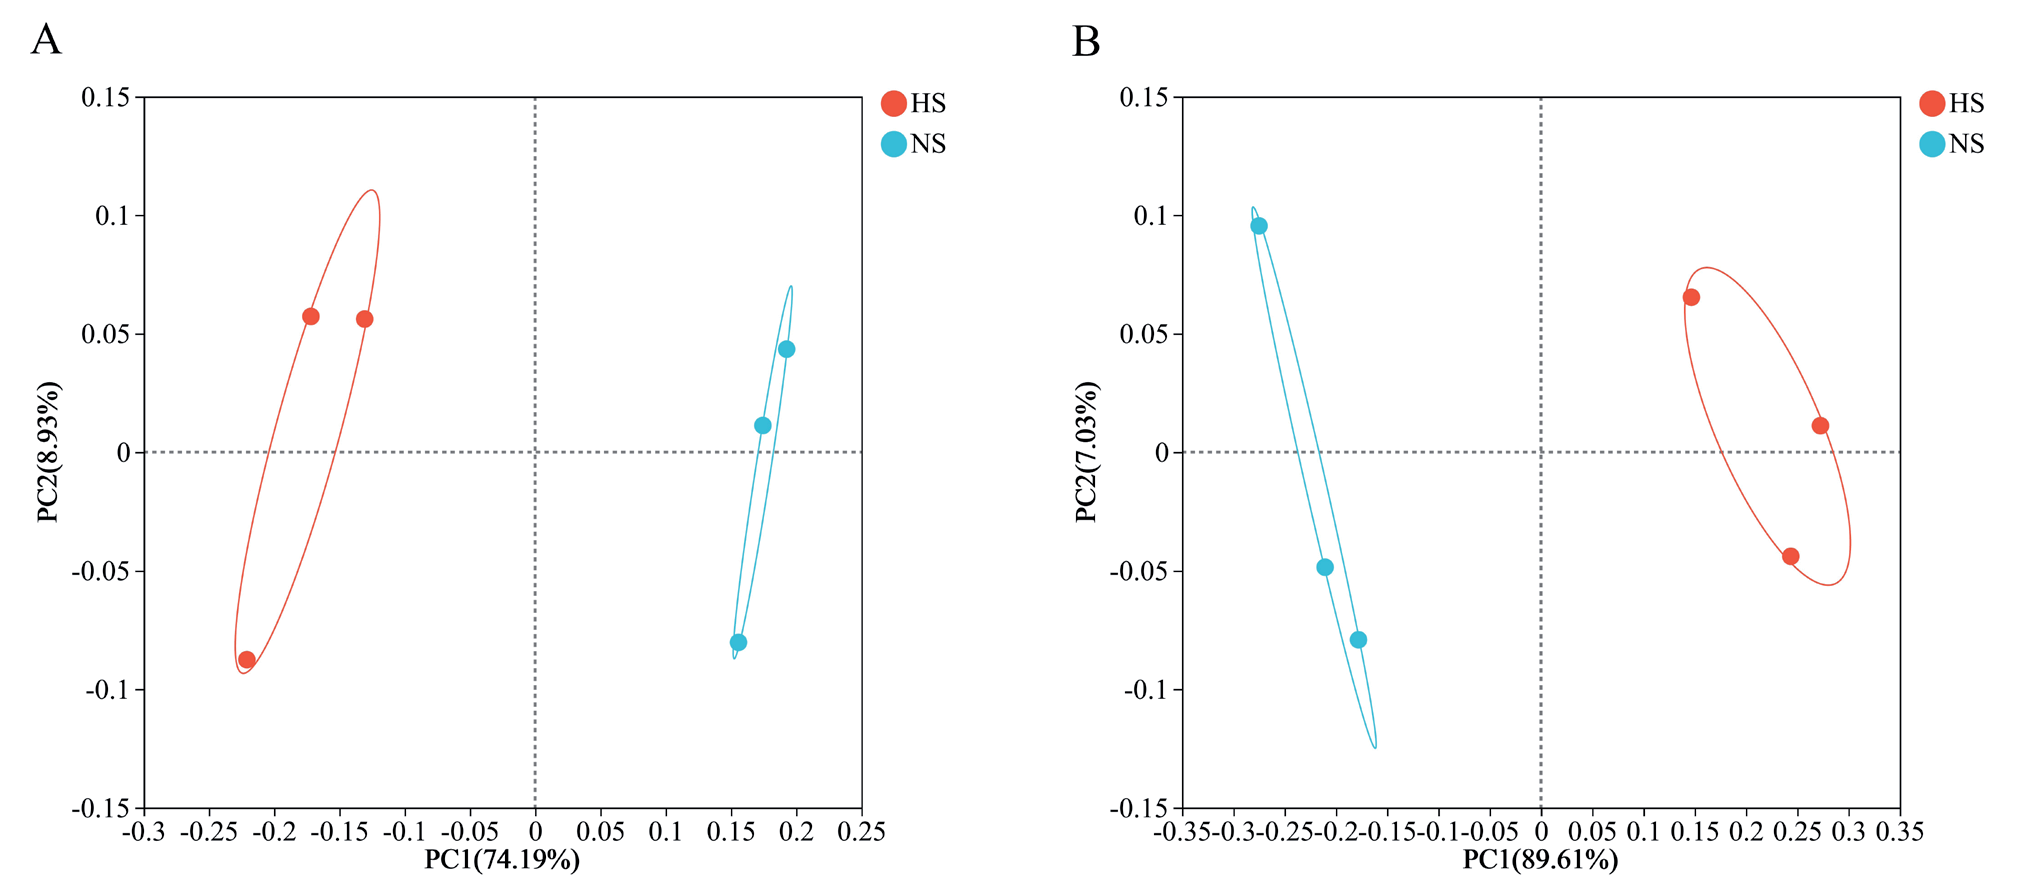
PCA revealed a clear distinction in the composition of bacterial and fungal communities in the soil samples of each treatment group, with the differences between the treatments being significantly greater than those within the soil samples from the same treatment (Fig. 3A-B). These results indicated that the beta diversity of the bacterial and fungal communities was significantly different between soils infested with M. incognita and healthy soils.
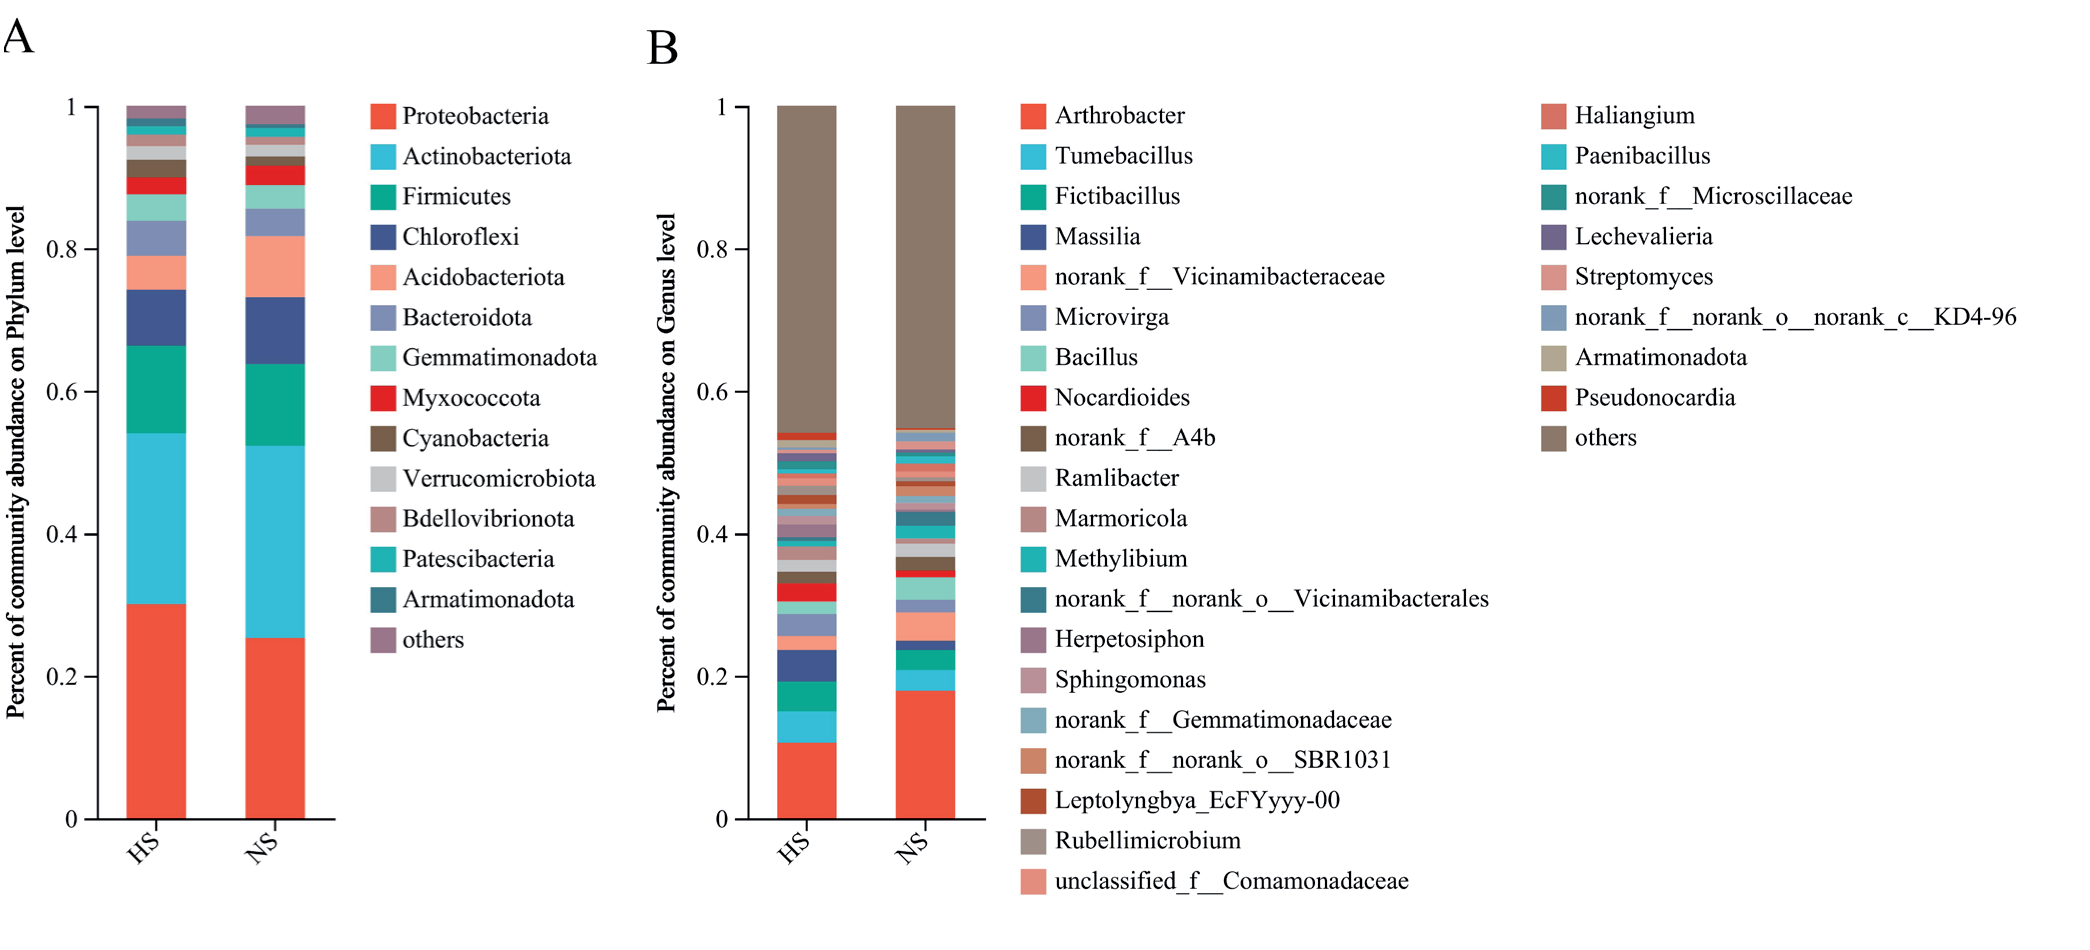
Assessing the differences in bacterial community structure and composition between healthy and infested soils at the phylum level, using a relative abundance greater than 1% as the threshold, revealed that the bacterial microbial community composition in the two soil treatment groups was similar (Fig. 4A), both containing 13 bacterial phyla. Among them, Proteobacteria, Actinobacteriota, Firmicutes, Chioroidota, and Acidobacteriota were the dominant bacterial phyla in the samples of each treatment group. Proteobacteria had a relative abundance of 30.07% in the healthy soils, which was higher compared to the 25.31% observed in the infested soils. Actinobacteriota had a relative abundance of 25.31% in infested soils, higher than 23.98% in healthy soils. Firmicutes had a relative abundance of 12.30% and 11.47% in healthy and infested soils, respectively. At the genus level, with a threshold of relative abundance not less than 1%, 28 bacterial genera were identified in both treatment groups (Fig. 4B). Among them, Arthrobacter, Tumebacter, and Fictibacillus were the dominant bacterial genera in both treatment groups. Arthrobacter had a relative abundance of 17.83% in the infested soils, higher compared to the 10.68% in the healthy soils. Tumebacter had a relative abundance of 4.42% in the healthy soils, higher compared to the 2.98% in the infested soils. Fictibacillus had a relative abundance of 4.22% and 2.79% in healthy and infested soils, respectively.
Using a relative abundance greater than 1% as the threshold, twenty-five bacterial genera exhibited significant differences in relative abundance (p < 0.05) between the two soil treatment groups (Fig. 5). The relative abundances of 11 bacterial genera were higher in NS than that in HS, as follows:
Arthrobacter, norank_ f __Vicinamibacteraceae, Bacillus, Methylibium, norank_ f __norank_o__ Vicinamibacterales, norank_ f __norank_o__ SBR1031, Haliangium, Paenibacillus, Streptomyces, and norank_ f __norank_o__norank_c_KD4-96,
norank_ f __Anaerolineaceae. The relative abundance of these 11 bacterial genera was 1.67, 2.10, 1.80, 2.39, 3.59, 2.22, 1.71, 1.55, 2.46, 3.51, and 3.12 times higher in NS than that in HS, respectively. A total of 14 bacterial genera exhibited a higher relative abundance in HS compared to NS, as follows: Tumebacillus, Fictibacillus, Massilia, Microvirga, Nocardioides, Marmoricola, Herpetosiphon, Sphingomonas, Rubellimicrobium, unclassified_ f __Comamonadaceae, norank_ f __ Microscillaceae, Lechevalieria, norank_ f __ norank_o__norank_c__norank_p__Armatimonadota, Pseudonocardia. The relative abundance of these 14 bacterial genera in HS was 1.53, 1.51, 3.41, 1.80, 2.79, 2.69, 5.19, 1.33, 2.27, 1.37, 1.93, 3.07, 2.37, and 4.66 times higher than that of NS, respectively.
Table 2.
The significantly upregulated metabolites in rhizosphere soil after M. incognita infestation.
[i] Note: VIP indicates the VIP value of each metabolite in the OPLS-DA model between the two treatment groups. FC indicates the fold change of each metabolite concentration between the two groups. P_ value represents the significance test result of the difference between the two groups for that metabolite. The same applies in the table below.
Regarding the differences in the fungal community composition between healthy and infested soils at the phylum level, using a relative abundance greater than 1% as the threshold, a high similarity in the fungal microbial community between soil treatment groups was observed (Fig. 6A). Three fungal phyla, Ascomycota, unclassified_ k_ fungi, and Mortierellomycota, exhibited the highest relative abundance in both groups. Among them, Ascomycota was the most dominant fungal phylum in both samples, accounting for 98.03% and 90.75% of the total fungal community in healthy and infested soils, respectively. At the genus level, using a threshold of relative abundance greater than 1%, 14 fungal genera were identified in both soil treatment groups (Fig. 6B). Iodophanus was the dominant fungal genera in the samples of each treatment group. It had a relative abundance of 56.92% in the healthy soils, higher compared to the 23.61% in the infested soils.
Table 3.
The significantly downregulated metabolites in rhizosphere soil after M. incognita infestation.
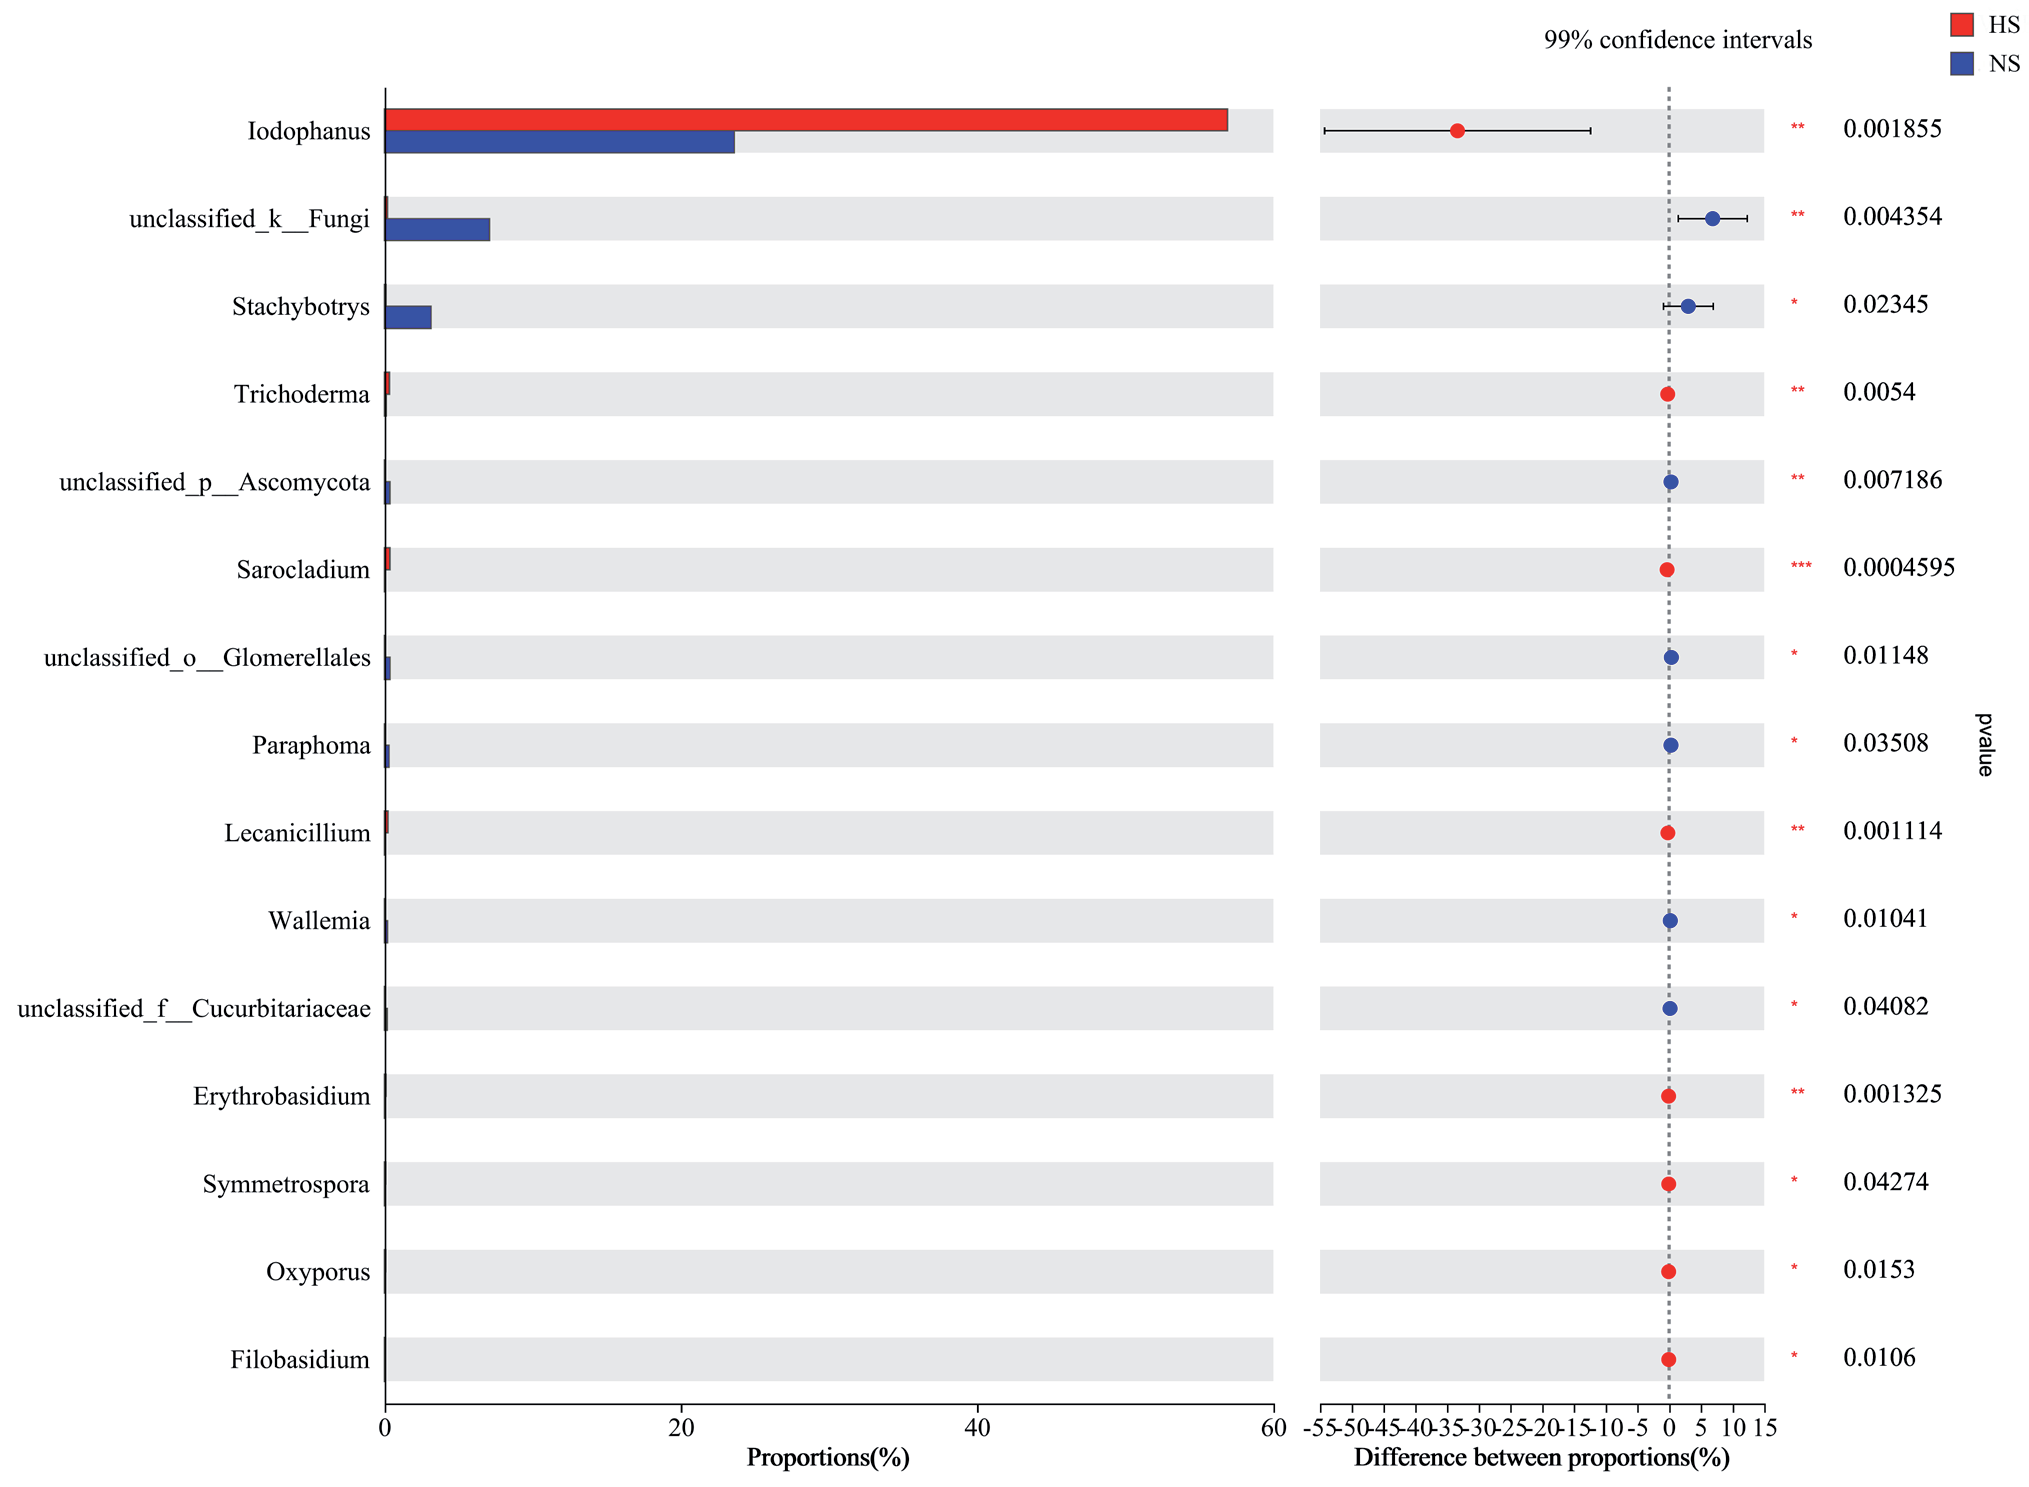
Using a relative abundance greater than 1% as the threshold, three fungal genera exhibited statistically significant differences (P < 0.05) (Fig. 7). Among them, there were two fungal genera; the relative abundance in NS was significantly higher than in HS, as follows: unclassified_k__Fungi and Stachybotrys. The relative abundance of these two fungal genera was 35.59 and 32.64 times higher in NS than that in HS, respectively. There was one fungal genus, Iodophanus, with a relative abundance of 56.92% in HS, significantly higher than in NS.
Correlation Analysis
In the correlation heat map between the 51 differential metabolites (27 upregulated and 24 downregulated) and the 28 differential microbial genera (25 bacterial genera and three fungal genera) in the soils before and after infestation with M. incognita (Fig. 8), a total of 182 significant pairwise correlations were identified (P < 0.01, |r| > 0.9). These correlations included 49 metabolites and all microbial genera included in the analysis. Among them, five metabolites were associated with more microorganisms; they were eremopetasinolone A, fusarochromanone, auberganol, 5-fluorouridine monophosphate, and (2S, 2'S)- pyrosacharpine, respectively. Nine microbial genera are associated with more metabolites; they were norank_ f __ norank_o__norank_c__norank_p__Armatimonadota, Methylibium, Paenibacillus, Iodophanus, Haliangium Bacillus norank_ f _Microscillaceae, Streptomyces, and Marmoricola, respectively.
Table 4.
Alpha diversity of the bacterial and fungal communities in healthy soils (HS) and infested soils (NS).
| Soil sample | Bacteria | Fungi |
|---|---|---|
| Shannon index | Shannon index | |
| HS | 5.73±0.08a | 1.9±0.08a |
| NS | 5.66±0.05a | 2.46±0.27a |
Discussion
The current study aimed to discern the changes in the metabolite profiles among diverse soil samples attributed to M. incognita infestation in plants. Subsequently, following a correlation analysis between microbial genera and soil metabolites, we identified five metabolites that were significantly correlated with the abundance of specific microbial genera. Including 5-fluorouridine monophosphate, fusarochromanone, eremopetasinolone A, auberganol, and (2S, 2'S)- pyrosaccharopine. According to Tables 2 and 3, except for auberganol, all metabolites were significantly upregulated after infection with M. incognita. The compound 5-fluorouridine has been identified as a specific antivirulence agent, which can disrupt bacterial RNA metabolism and inhibit the synthesis of pyridoxine, a key toxin, thereby reducing Pseudomonas aeruginosa virulence in Caenorhabditis elegans [15]. Additionally, 5-fluorouridine monophosphate may have potential therapeutic properties and application potential for treating Candida albicans infections [16]. Fusarochromanone is a poisonous metabolite generated by Fusarium equiseti, a fungus commonly found in the decomposing tissues of cereal plants in northern latitudes. As shown in previous studies, fusarochromanone serves as an adjuvant molecule, effectively impeding melanoma growth and recurrence by increasing tumor cell apoptosis and reducing tumor growth in vivo [17]. However, the potential impact of fusarochromanone on M. incognita fitness and metabolism remains unclear and warrants further investigation. Notably, the properties and effects of eremopetasinolone A, auberganol, and (2S, 2'S)- pyrosaccharopine are not yet investigated.
Fig. 5.
Bacterial genera with significantly different abundance between healthy soils (HS) and infested soils (NS). Note: The x-axis represents the average relative abundance of a species in different treatment groups, and the y-axis represents the different groups. *P ≤ .05. **P ≤ .01. ***P ≤ .001.
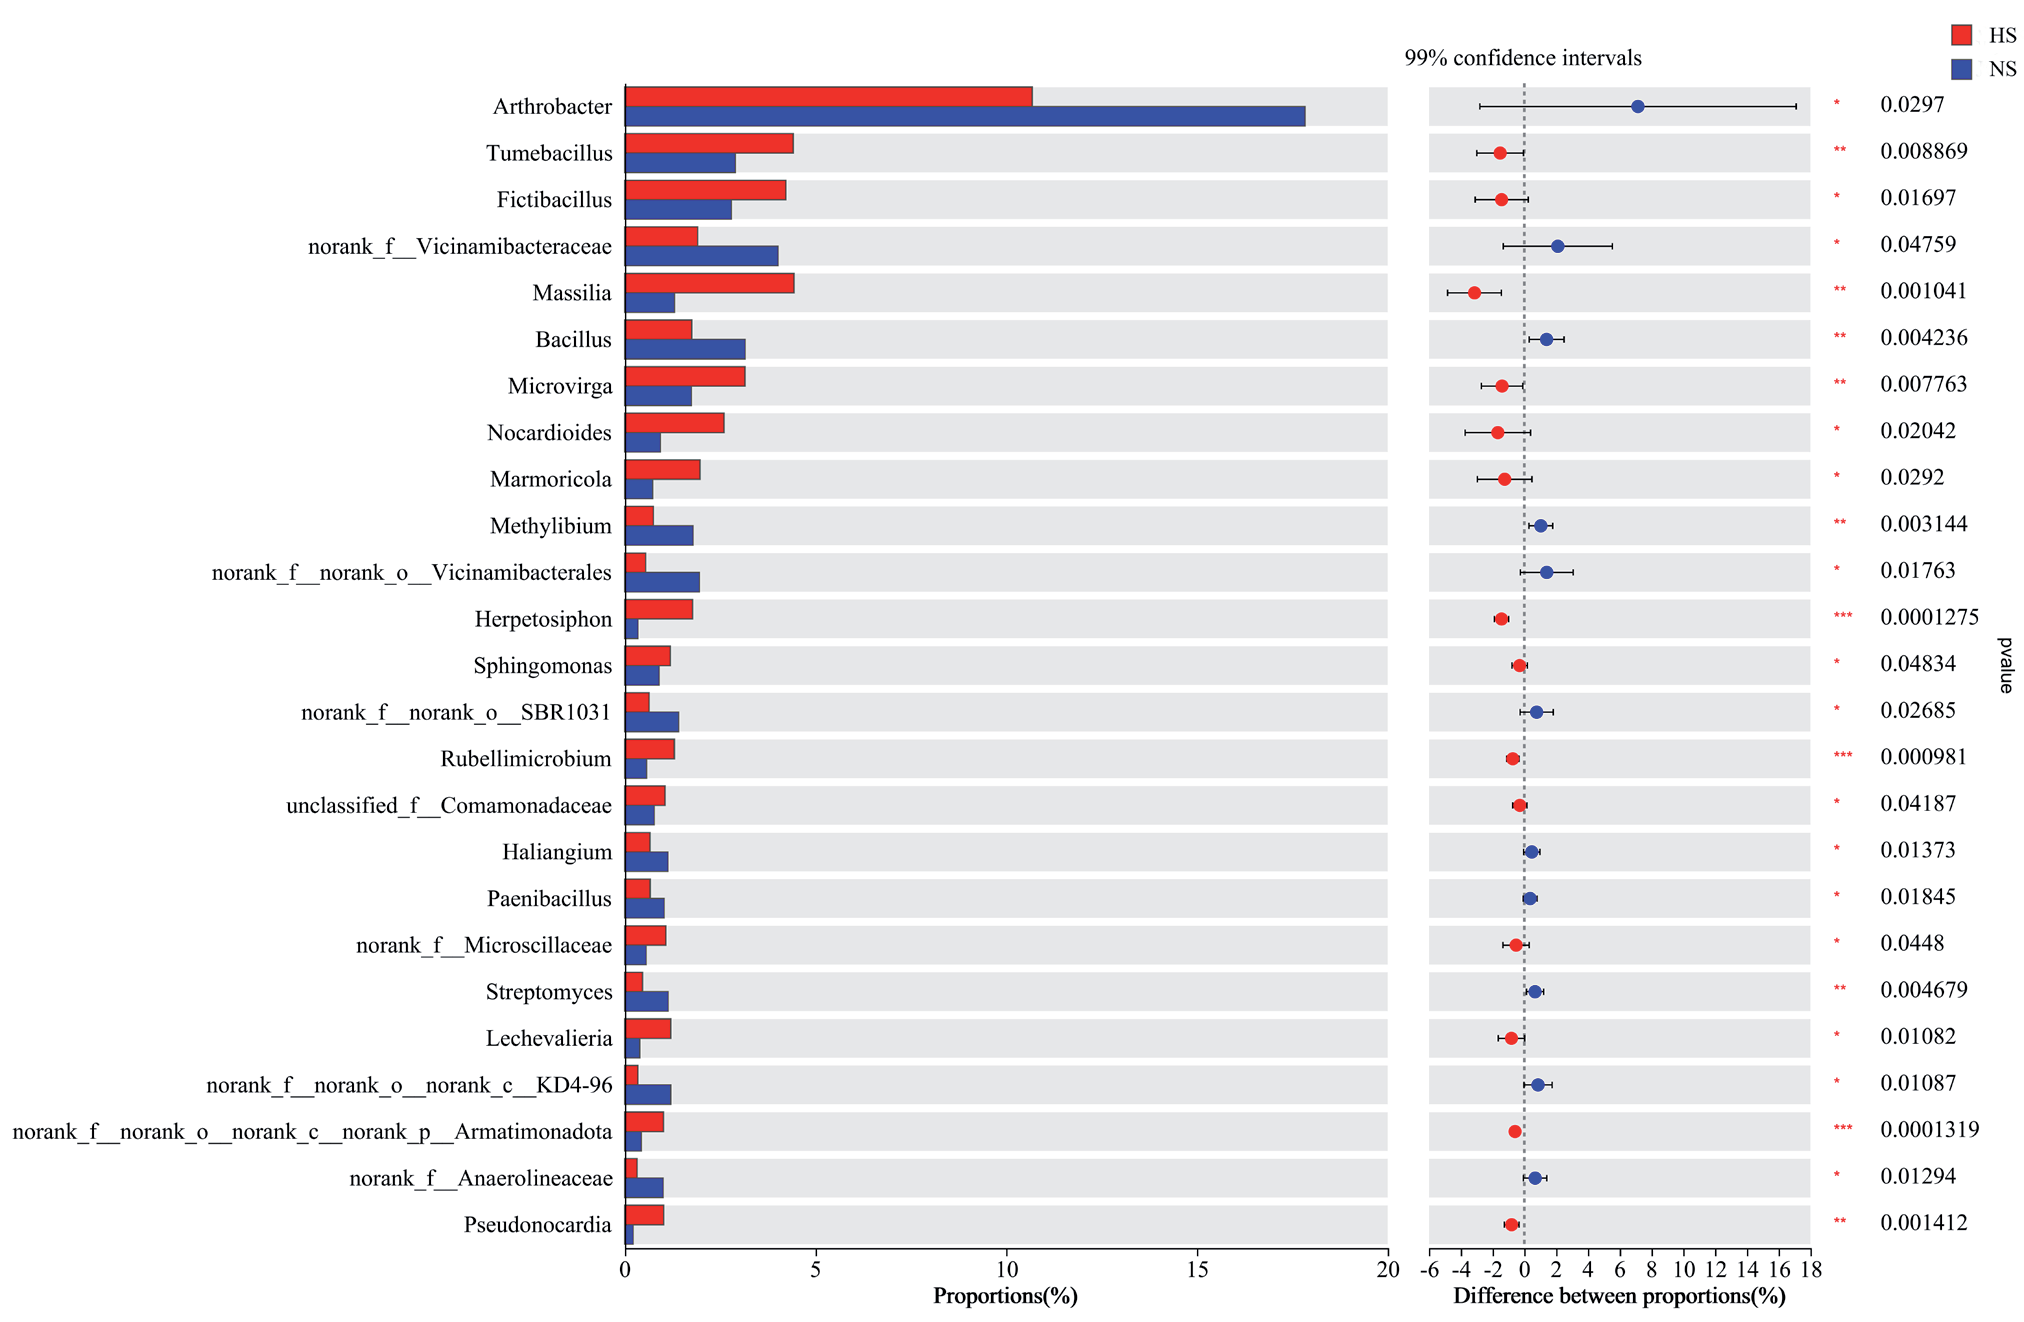
Fig. 6.
Fungal community composition in healthy soils (HS) and infested soils (NS). (A) Phylum. (B) Genus.
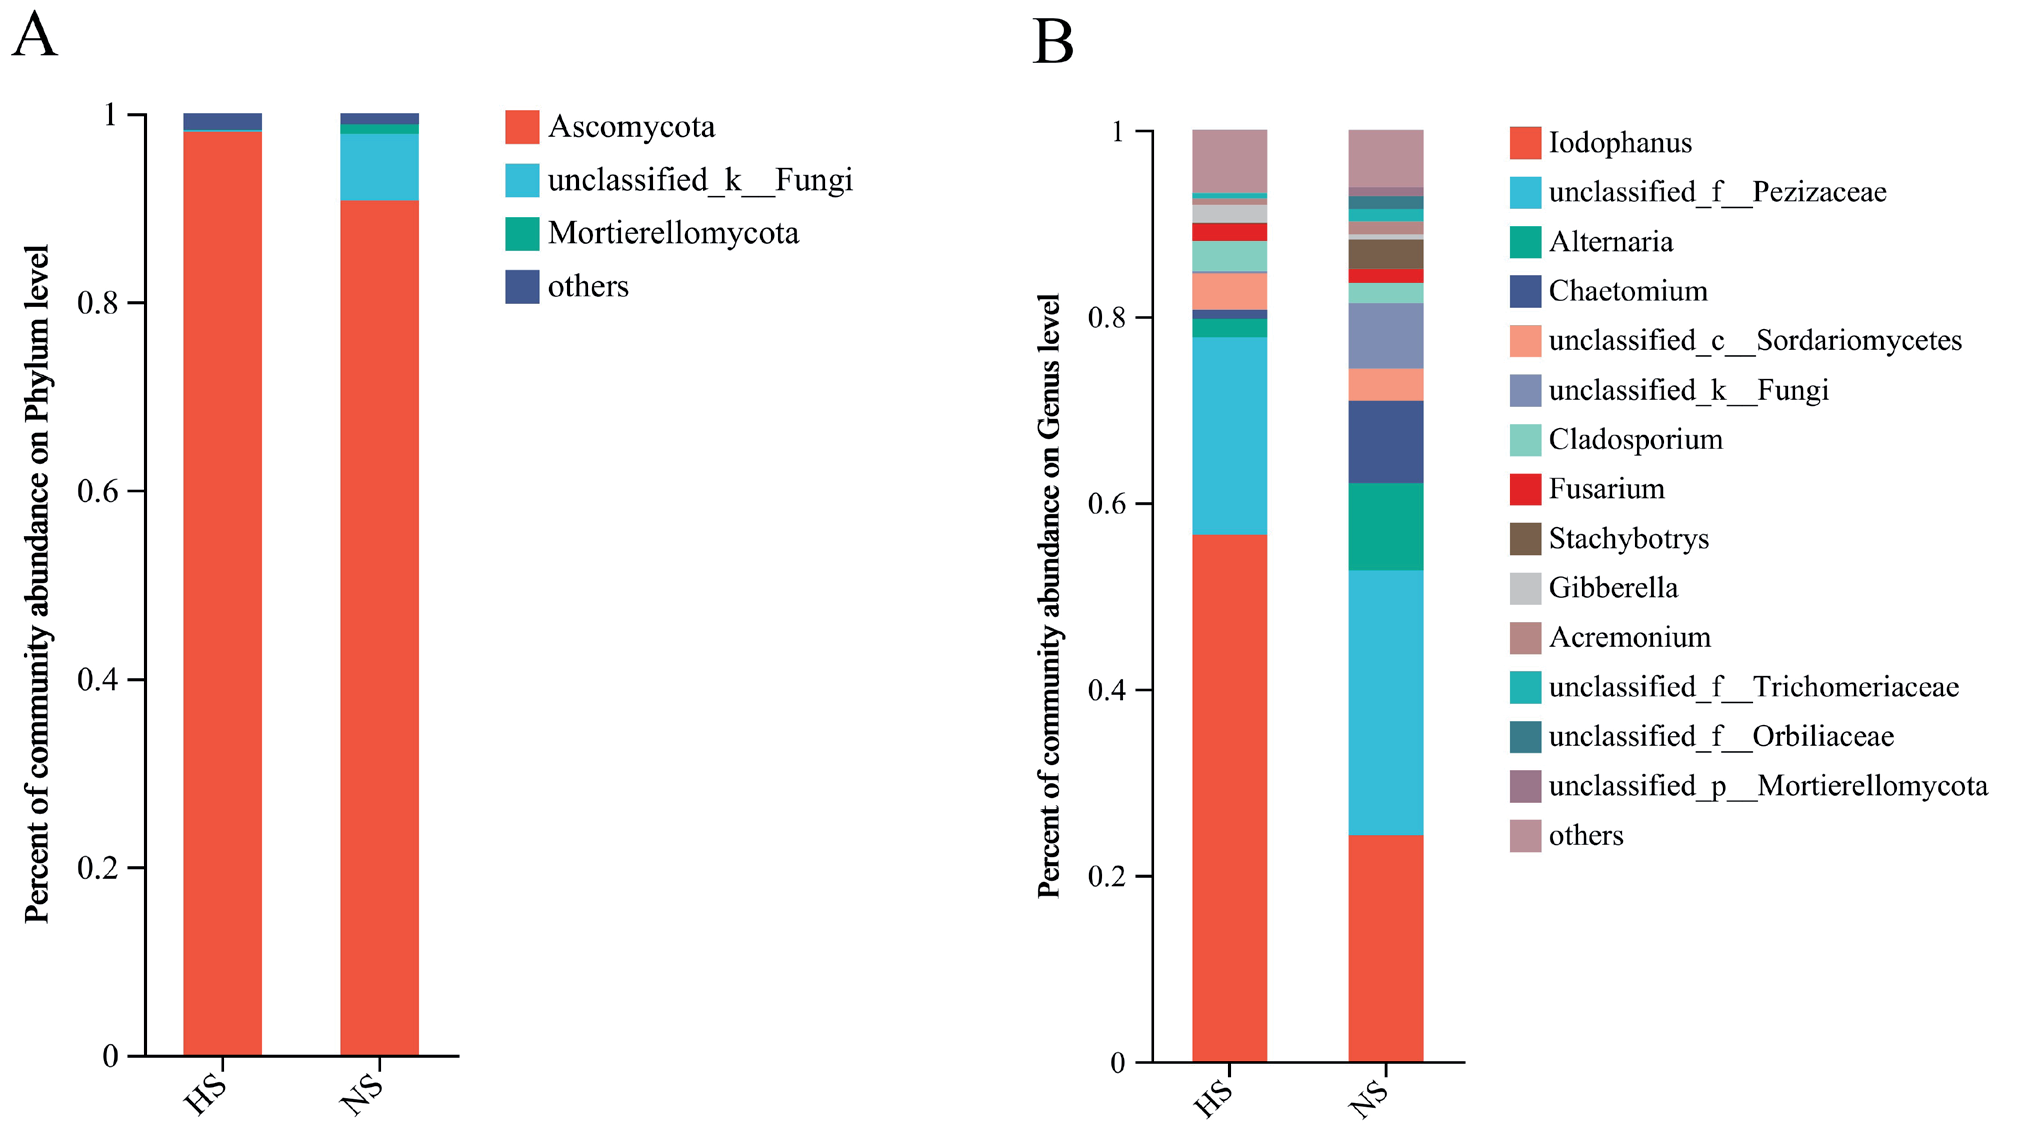
Fig. 7.
Fungal genera with significantly different abundance between healthy soils (HS) and infested soils (NS). Note: Same as Fig. 5.
It is crucial to acknowledge that rhizosphere soil metabolites are not solely derived from rhizosphere microorganisms. The role of root secretions in metabolite production should not be underestimated. According to reports, the root exudates of rosemary and marjoram can inhibit the survival, development, and reproduction of nematodes [18]. Therefore, metabolites with a weak correlation with changes in rhizosphere soil microbial communities in this study may be produced in plants. However, further exploration is needed to determine whether these metabolites have biocontrol potential.
The microbiome of the plant rhizosphere directly influences plant health and resistance to pathogens [19]. Following exposure to aboveground or belowground stressors, significant alterations in root metabolic profiles and gene expression occur in conjunction with changes to the rhizosphere microbial community [20]. In this study, we explore the differences in microbial community structures of the rhizosphere soil around tomato roots before and after M. incognita infestation. There were 51 microbial genera with significant differences. Among them, the relative abundance of 13 microbial genera was significantly higher in NS than in HS. Previous research has established Bacillus as a well-established microorganism that can effectively inhibit the activity of M. incognita. Bacillus, a genus of bacteria that resides extracellularly between the rhizosphere and root cortex cells, has been classified as a rhizosphere bacterium that promotes plant growth and primarily protects against M. incognita through the secretion of hydrolases and secondary metabolites [21]. In a previous study, Bacillus strain R2, isolated from tomato, was highly effective against M. incognita, with a biocontrol lethality rate of 90.94%. Additionally, a nematocidal compound identified as styrene was isolated [22]. Notable antibiotics produced by Streptomyces include avermectin, meliloticin, and nicomycin, among others. According to reports, natural fermentation products contain eight avermectins with structurally similar properties and a common mother nucleus, each exhibiting insecticidal activity. The widely used ivermectins are highly effective antibiotics developed through structural modification of avermectins [23]. Stachybotrys exhibited nematicidal activity, up to 35% [24]. A significantly higher relative abundance of Methylibiumv has been reported in soil from artificially disturbed areas compared to naturally undisturbed areas [25]. It has been reported that Haliangium is isolated from coastal areas and confirmed to have salt tolerance properties [26]. Paenibacillus has been proven to have radiation resistance [27]. However, we observed that the relative abundance of 15 microbial genera was significantly higher in HS than in NS. Fibribacillus phophorivorans G25-29 has been proven to have nematicidal ability and can inhibit its free activity [28]. Two strains of bacteria with nematicidal ability have been isolated from mangrove sediments and identified as Pseudonocardia [29]. Marmoricola is an actinomycete found in oceans and ash, among other environments [30, 31].
Fig. 8.
Correlation Heatmap between the differentially accumulated metabolites and the differentially abundant microorganisms. Note: The right side of the plot indicates the metabolite name, and the bottom indicates the microbial genera. Each lattice in the plot represents the correlation between two attributes (metabolite and microbial genus), and different colors represent the magnitude of the correlation coefficient between attributes. *P ≤ .05. **P ≤ .01. ***P ≤ .001.
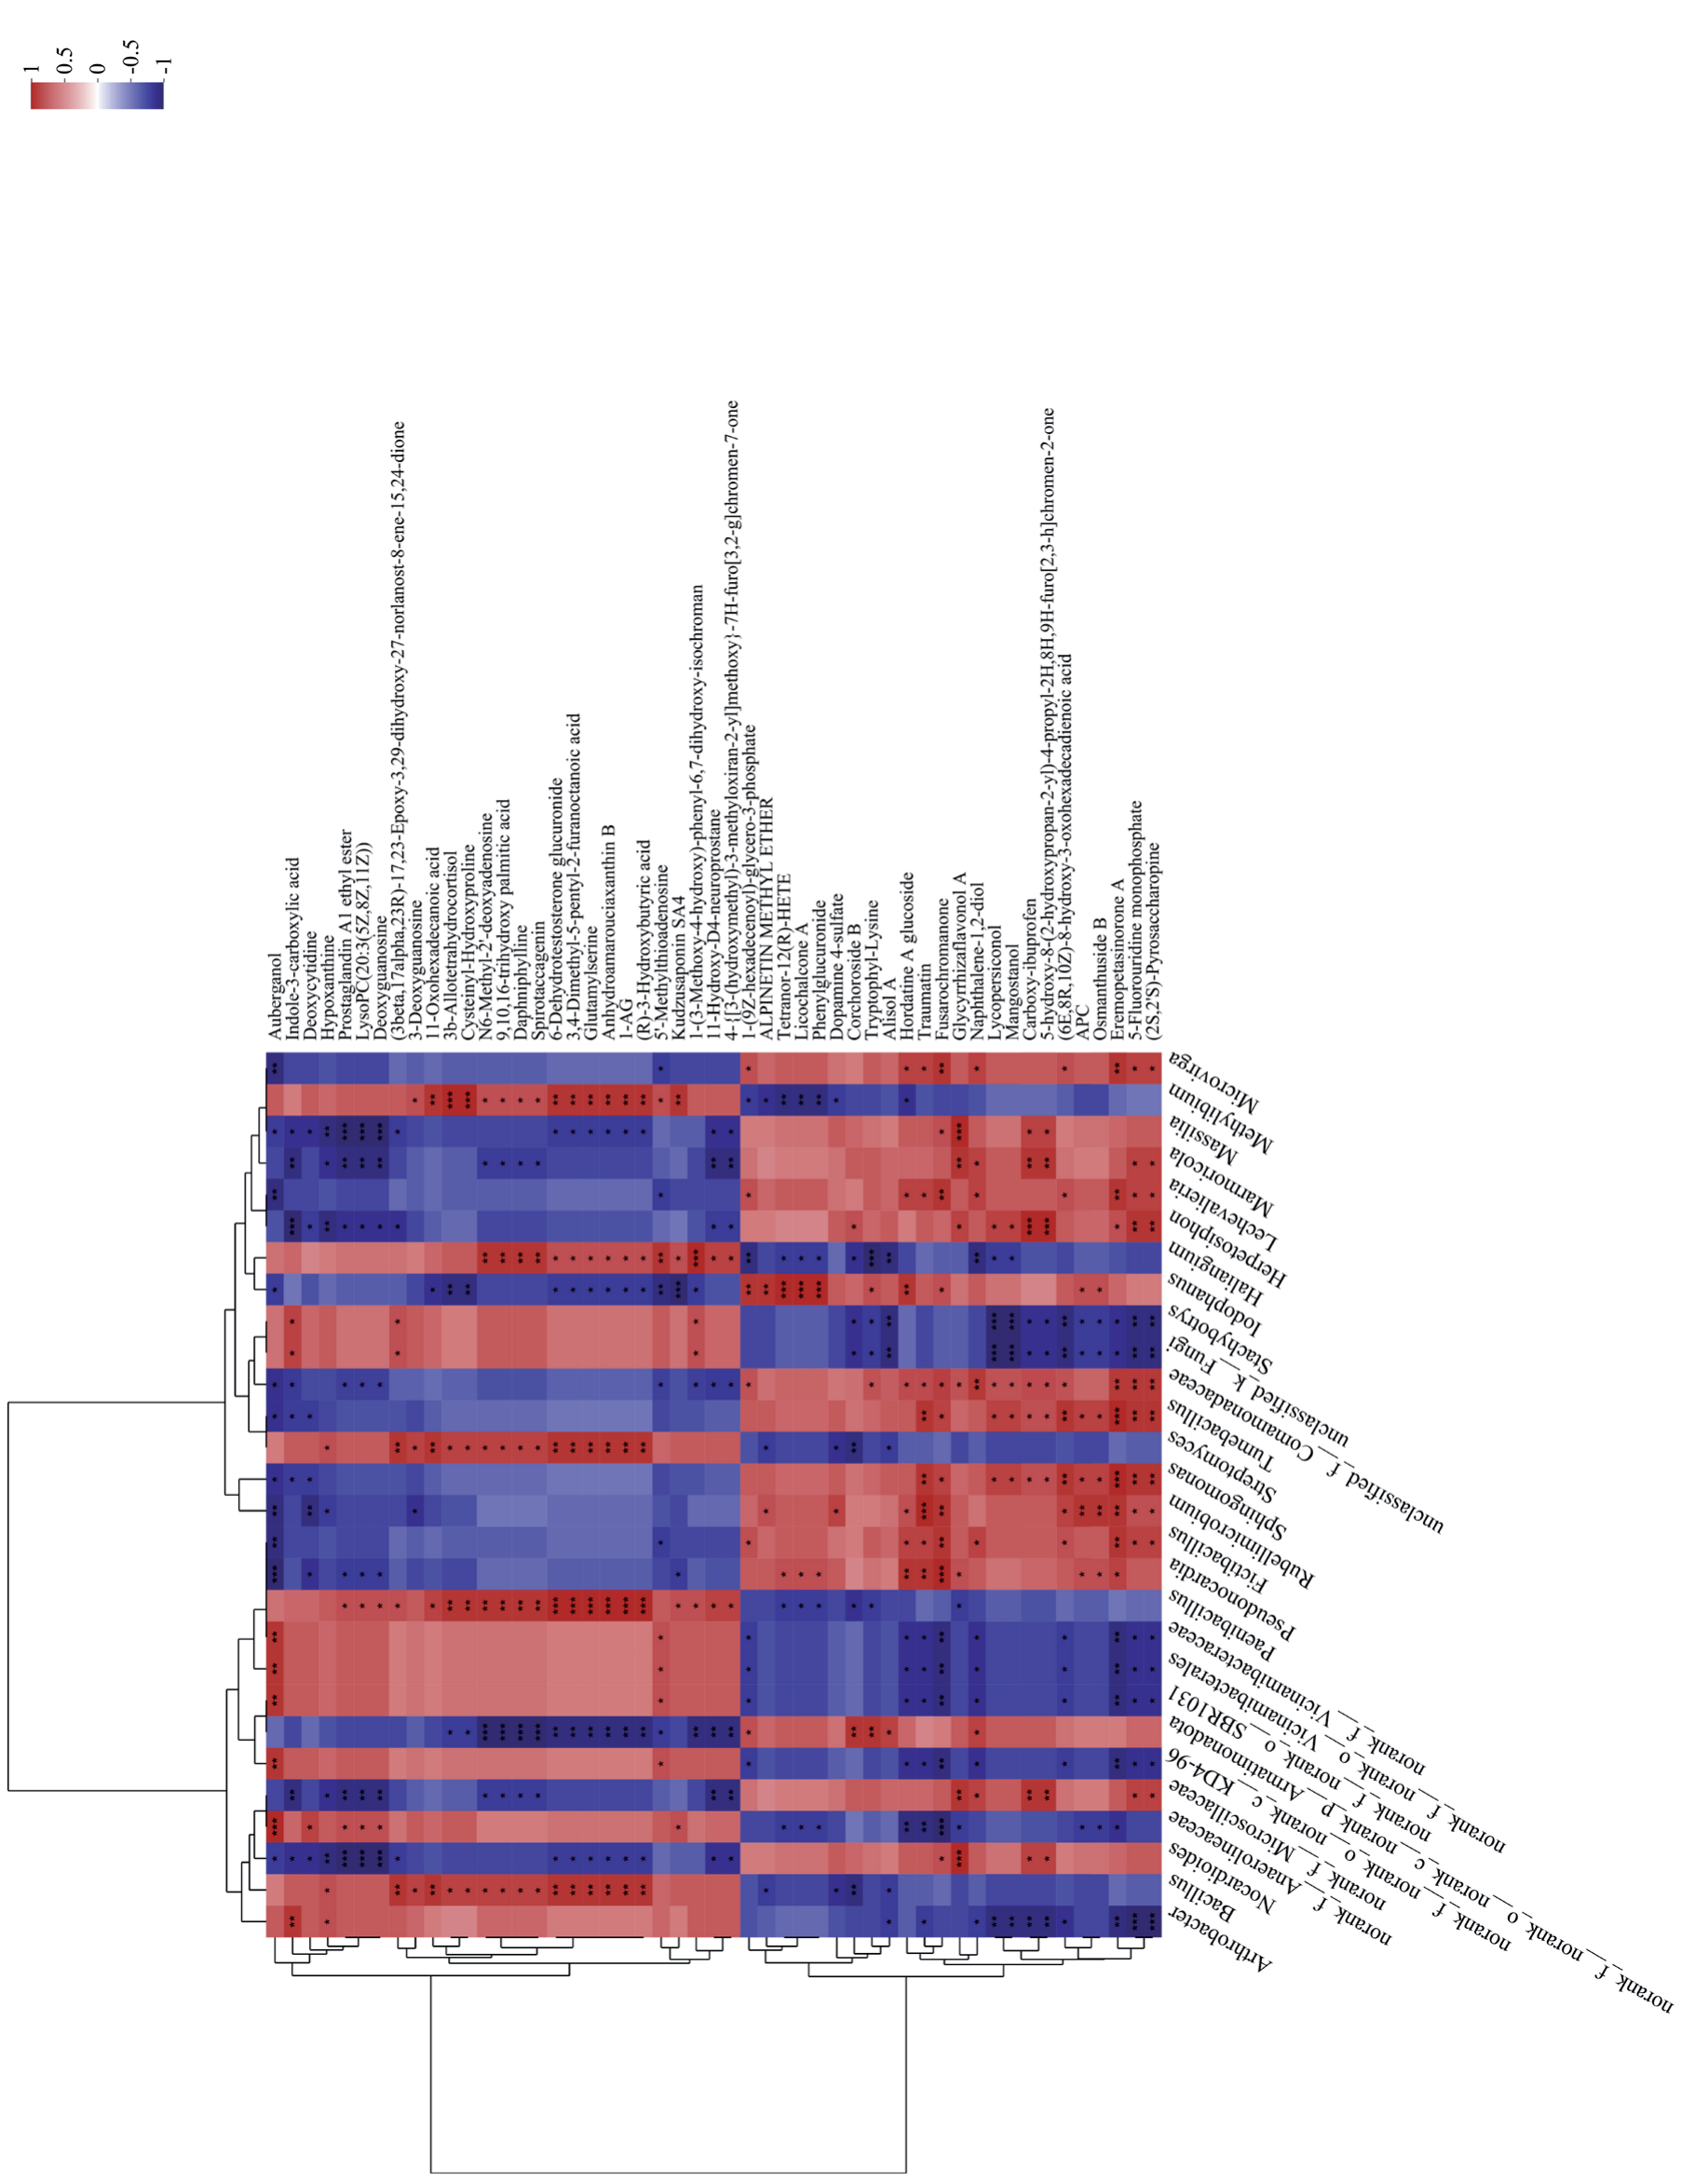
Furthermore, nine microbial genera were identified as significantly associated with specific microorganisms through association analysis. Based on existing research results, among these nine microbial genera, Bacillus and Streptomyces have nematicidal activity. While Paenibacillus, Methlibium, Halangium, and Marmoricola were shown to be sensitive to abiotic stress conditions. However, no studies on Iodophanus and two other undefined bacterial genera have been conducted so far.
Conclusion
The presence of M. incognita in the rhizosphere soil of tomato plants has been observed to significantly impact the soil metabolite levels and microbial diversity. Through correlation analysis, a total of five metabolites and nine microbial genera were identified as potential candidates for the biocontrol of M. incognita. These findings can provide a better understanding of the environmental changes in the rhizosphere soil of tomatoes infected with Meloidogyne incognita and establish a theoretical basis for discovering novel biocontrol agents.

