

Introduction
Faced with climate change, energy, agriculture, tourism, and pollution issues, the EU is determined to address them in such a way as to provide its citizens with the highest environmental standards. Objectives and goals to be achieved within 2020-2050 are set and prioritized by research programs, legislation, and funding [1]. Pollution is a global problem that affects human health, biodiversity, and the economy. It is a problem that is undoubtedly growing, creating a cascade of consequences that require a completely different approach with the same outcome: a clean, healthy, and sustainable environment. In recent years, many types of pollutants have been uncontrolled and released into the environment. Hydrocarbons, heavy metals, and pesticides are organic and inorganic contaminants with the highest toxic effects.
Ecology can be seen as the study of nature’s household viewed through abiotic and biotic perspectives, researched via the metabolism of living organisms, and understood as a weapon against domicide [2]. The reasoning behind the classical definition of ecology is an accurate understanding of humanity’s demands on the biosphere. Thus, frequently exposed to a mixture of xenobiotics such as heavy metals, pesticides, and other toxic substances, bioaccumulation and biosorption can be only evolutionary steps [3]. In line with what was previously said, natural techniques consider the use of biological remediation methods for the contaminated environment as essential. The biological methods available for soil remediation are phyto- and bioremediation, considered the safest and, thus, ecologically acceptable [4, 5].
Phytoremediation is a natural process in which plants uptake and transform a variety of pollutants. This method also involves rhizosphere stabilization, boosting microbial growth in plants’ root zone [6- 8]. Although applicable in water, phycoremediation is categorized as phytoremediation and bioremediation [9], in which algae and microalgae naturally remove nitrogen, phosphorus, sulfur, and minerals from water. A simple explanation would be that algae feed on these pollutants in water [10]. Also, using algae and microalgae to remove phenolic compounds from household wastewater was useful in treating communal wastewater [11]. Sing et al. (2021) showed that filamentous algae, a genus Spirogyra, can be used for Cr(VI) removal and decolorization of methylene blue dye in wastewater [12]. Another method called mycoremediation, or fungal bioremediation, includes fungal microbes’ capability to effectively adapt and clean the environment [13]. The bioremediation method includes physical, chemical, and biological techniques. These include mechanisms categorized into in-situ and ex-situ methods in the polluted place or outside it. Certain organisms have the genetic potential to remove, degrade, metabolize, or immobilize a wide range of contaminants from soil and water. The biochemical ability to tolerate high concentrations of toxic organic and inorganic pollutants is related to specific genes involved in detoxification by enzymes, transport protein families, and efflux pump proteins due to adaptation to specific environmental conditions [7, 14]. Bioremediation as a technique presents a series of reactions in which the main role is played by enzymes, synthesized by the actors of bioremediation themselves, microorganisms. This process involves the participation of archaea, bacteria, algae, and fungi whose application showed successful, inexpensive, and safe methods [15].
Sustainable Development, Ecology/Environment, and Bioremediation Are Closely Intertwined Fields
Sustainable development aims to balance human needs with using natural resources (soil, air, water). Considering economic growth and social rights, 17 sustainability development goals are brought by the UN Agenda 2030 [16]. All sustainable goals are viewed through different sets of indicators; they include life quality and ecological/environmental sustainability. Sustainable development is supported by three pillars (environment, economics, and society), and goals are set for each of them [17]. Particularly, environmental sustainability is related to the capacity of the planet to give and stay preserved. Therefore, development must include remediation of polluted soil and water and/or proportionately using the resources of the planet Earth [18]. Therefore, the European Union set goals and objectives within the European Green Deal, focusing on zero pollution and research and innovation (R&I) action [18, 19]. One segment of this document, imperative for achieving environmental and ecological goals, presents the missions on soil, ocean, sea, and water remediation [20]. The EU supports science as a foundation for research and innovation, and this topic is detailed and elaborated under Cluster 6, which tackles the loss of biodiversity, healthy soil and water shrinkage, pollution problems, and environmental deterioration [21]. Specifically, the Horizon Europe program through Cluster 6 boosts sound solutions by implementing a circular economy and bioremediation to preserve, prevent, and remediate the environment. R&I in this cluster aims to prioritize the selected headline indicators built on the European Green Deal and the 8th Environment Action Program (2022-2030) [21].
Within Cluster 6, some projects prioritize molecular aspects of bioremediation. The Biosysmo project (BIOremediation systems exploiting SYnergieS for improved removal of mixed pollutants) brings novelty to bioremediation applications. Bioinformatics methods and genetic material manipulation techniques were applied to create a synergetic biosystem. There was also a special focus on secondary toxic substances and their neutralization. There are several publications funded by this project [22]. Dvořák et al. (2023) analyzed the potential of Pseudomonas putida mt-2 to biodegrade m-xylen (aromatics) under different water conditions, such as water potential. The research focused on the expression of the biodegradative xyl genes that encode for the XylR A domain. When modifying water potential, matric stress tolerance was shaped by the expression of xyl genes (the TOL plasmid pWW0). It modified the A domain of XylR (the structure of the binding pocket and entry tunnel). Water-dependent transcription changes led to low fitness of genotypes but adapted strains to new conditions [23]. Avendaño et al. (2023) revealed the Pseudomonas putida KT2440 catabolic pathway in the selenite to selenium nanoparticles transformation. The transformation involves sucA, D2HGDH, and PP_3148 genes that encode enzymes that bond the 2-ketoglutarate and enzymes of oxidative stress GSH by glutathione. An important role is the sulfite reductase encoded by cysG, sqr, pdo2, sqrR, ssuEF, and sfnCE sulfur metabolism genes. Certain genes revealed in P. putida (WT strain), such as sqrR, pdo2, and sqr, haven’t been found in other bacterial strains that metabolize selenium. Furthermore, these genes are of great interest because switching them off increases selenium nanoparticle production [24]. The GREENER project aims to provide natural, low-cost, and safe solutions by introducing bioremediation and bio-electrochemical technologies. To successfully remove metals and metalloids, PAHs, TPHs, antibiotics, pesticides, etc., a combination of bioremediation methods and risk assessment will be applied. [25]. The results of this project are presented in several studies. Martínez-Cuesta et al. (2023) explored aliphatics and aromatics breakdown rates in polluted soils from Ireland using more than one bioremediation technique (Ecopiling). During the process, DSeq2 software was used for microorganism identification and appearance in the given remediation period. It was notable that at the beginning of the process, there were more TPH degraders (Alcaligenaceae or Pseudomonadaceae, Gammaproteobacteria); afterward, bacterial populations inhabit less polluted soils such as Actinobacteria, Saccharimonadia, Acidobacteriota, and some Actinobacteriota or Firmicutes were noticed. At the end of bioremediation, about 90% of TPHs in polluted soil were degraded [26]. Similarly, Verdel et al. (2021) explored the degradation rate of organic additives used in paper mills. Consortia of native and engineered microorganisms were used. The four bacteria in the process of bioaugmentation, Xanthomonadales bacterium and Sphingomonas sp., and Cellulosimicrobium and Aeromonas sp., showed the biggest potential [27]. Garrido-Sanz et al. (2020) showed that within the Rhodococcus genus, the majority can break down aromatic and aliphatic compounds; however, only some strains, like Rhodococcus sp. WAY2 can degrade short-chain n-alkanes [28]. Further investigation of Rhodococcus sp. WAY2 showed independent clusters of bph, etb, and nah genes involved in the biodegradation pathway of aromatic compounds. The hydrogenase enzyme system coded by these clusters enables Rhodococcus sp. WAY2 to metabolize biphenyl, naphthalene, and xylene. The ability to use short and long chains of n-alkanes is due to the presence of chromosomal alkB and ladA genes [29]. Another project funded by the EU, MIBIREM, brings solid solutions regarding bioremediation. This project aims to create a toolbox to gather molecular methods and bioinformatics tools. Microorganisms will then be applied in cyanides, hexachlorocyclohexane (HCH), and petroleum hydrocarbons (PHC) polluted cities within Europe. The expected outcome is the creation and application of the MIBIREM TOOLBOX, which includes identified bioremediation and their genetic information, as well as IT modeling and forecasting tools, resources, and protocols [30]. The objective mentioned in previous documents is achieving the SDGs regarding the environmental pillar ˝planet˝ by using biocenosis (decomposers) in pollution removal.
The introduction of organic and inorganic pollutants into an ecosystem causes adverse impacts on health, biota, and environmental change. The remediation process includes pollution elimination to preserve the ecosystem’s health [31]. An ecological approach includes decomposers, as they can break down pollutants into harmless byproducts, therefore restoring ecosystems. Biological processes include bioremediation and biotechnology. Apart from biosensors and their wide use in bioremediation, genetic engineering/genetic modification or manipulation still keeps the top position by using living organisms as a tool in technology [32].
Bioremediation is considered economical, as a natural way of eradicating toxins and restoring polluted sites by choosing the right techniques. Following the principles of bioremediation, there are two main techniques, biostimulation and bioaugmentation, used to degrade harmful and toxic substances.
Bioaugmentation introduces specialized microorganisms to enhance pollutant degradation, and they act as biocontrol agents in contaminated soil, whereas biostimulation involves the transfer of certain microorganisms that act as biostimulants to indigenous bacterial populations [32-34]. The relationship between ecology and bioremediation lies in their shared goal of understanding and restoring environmental health. Ecological principles inform bioremediation strategies by providing insights into the complex interactions within ecosystems and the potential impacts of contaminants on organisms and their habitats. Biotic and abiotic factors that have anthropogenic factors influence every ecosystem and, thus, all processes within. Additionally, the speed and degree of breaking down pollutants depend on the physicochemical characteristics such as solubility, volatility, and bioavailability of pollutants, as well as abiotic factors such as pH, temperature, and moisture content, and biotic factors such as the microbial community and interactions with living and nonliving nature. Overall, integrating ecology and bioremediation is essential for developing effective, sustainable solutions to environmental pollution and ensuring the long-term health and resilience of ecosystems [35, 36]. Therefore, successful bioremediation depends on microbial ecology and metabolism. Thus, developments in molecular microbiology can optimize bioremediation processes in a targeted manner. Novel advances in biotechnology introduced their genetic background, namely enzymes and genes that enable the degradation or chemical change of harmful organic and inorganic pollutants. The promising technology of genetic engineering provides a myriad of possibilities when it comes to removing As, Cd, Cu, Fe, Hg, and Ni, as well as POPs such as PAHs and PCBs [37], by using genetically modified organisms (GMOs) [38, 39] / genetically engineered microorganisms (GEMs) in bioremediation [40].
Materials and Methods
The framework of this article is built around imperatives given by the European Council through important documents related to environmental protection, climate change, and clean energy. Also, under this topic and with funds provided through the European Green Deal, Horizon Europe, and EU partnership for R&I, researchers are given the benefit of the doubt to halt and remove pollution [19-21, 39]. In this document, the application of living organisms native to polluted areas brings green solutions and deals with issues related to agricultural soil, farming, and unhealthy soils. Also, the freshwater issue and polluted oceans and seas can be addressed by bioremediation. Under Horizon Europe, the EU research and innovation program (2021-27) aims to bring solutions and results by 2030 with 5 missions. This research focuses on living labs and lighthouses within “A Soil Deal for Europe: 100 Living Labs and Lighthouses to Lead the Transition towards Healthy Soils by 2030 and Restore our Oceans and Waters by 2030” [20]. The term living labs refers to on-site dealing with pollution, while lighthouses will promote and share good practices. Mission hotspots include soil pollution, restoration, clean oceans, and freshwater. With 60-70% of European soils polluted and 100 living labs and lighthouses across the EU, the estimations are 30-40% remediated soil by 2030 [41]. The mission to restore oceans and waters includes lighthouses set to be enforced across the EU seas and rivers [42]. One important document, EU Circular Talks, brings bioremediation as techniques that use living organisms such as bacteria, fungi, and algae to reuse and recycle valuable materials, such as heavy metals and organic residues, from hazardous waste fractions. Also, in this context, bioremediation reduces soil and water pollution, thus removing it [43]. This study aims to present substantive examples of the capacity of some microorganisms to remove toxic substances from the environment. Therefore, the study focuses on the determinants in native and engineered organisms. The article aims to provide biochemical data valuable in cleaning particular pollutants in vivo/ in vitro, showing scientific data available to researchers and stakeholders. The paper aims to clarify the potential of in situ biological techniques for their application to different inorganic or organic soil contaminants in the environment.
What Makes Microorganisms ValuableT in Bioremediation
What is behind the ability to clean the environment from different types of pollutants? How do they effortlessly and successfully do it? All living organisms are determined by their genetic material (genotype) and their environmental interaction (phenotype/phenotypic plasticity).
The adaptive capacity of microorganisms, based on their phenotypic plasticity, confers a fitness advantage in different environments. The genomic plasticity of biodegradation strains enables unhindered growth and development in the presence of very toxic pollutants [44]. The great success lies in the fact that microorganisms can transform pollutants into useful micronutrients. Therefore, the catabolic mechanism of these organisms is subject to many research articles, with genes and proteins/enzymes being target participants. Although proven to break down very toxic organic pollutants, some artificial organic chemicals, such as xenobiotics, are difficult to remove. Also, partial degradation can create more harmful compounds than the targeted ones. Engineered microorganisms have the genetic capacity to overcome these obstacles with efficient catabolic pathways [15, 45-47].
Genetic engineering has been widely used when it comes to removing mixtures of pollutants. Many bacteria can now grow and naturally remove both organic and inorganic pollutants. Targeted enzyme genes are cloned and expressed for successful bioremediation applications. For instance, the genetically modified strain Pb2-1 and the Rhizobium strain 10320D, Pseudomonas putida 06909, can degrade trichloroethylene (PCBs) but can remove/degrade HMs (Cd2+) by EC20 metal binding protein placed on the cell surface [46].
Finding appropriate bacterial strains that are able to remove mixtures of organic and inorganic substances in one place/polluted soil presents a quite difficult task [49]. Predominantly, inorganic pollutants such as NH3, NO3, and NH4ClO4 can be successfully transformed by microbes, while removing heavy metals is more demanding. This process involves a change in the valence states of metals. The catabolic pathway includes a genomic background. Genes that encode specific enzymes are located on chromosomes or belong to plasmids, transposons, or integrative-conjugative elements (extrachromosomal DNA). Molecular aspects of bioremediation via the catabolic pathway describe the strain’s physiology and metabolism, the species’ biodegradation potential, and homologous genes, directly indicating similarities and differences among different biodegradation strains [50]. Nowadays, omics technologies (genomics, transcriptomics, proteomics, and metabolomics) are the basis of successful remediation.
To be in a position to decide which microorganisms can apply or whether some should or need to be modified, microbe-based whole-cell systems and their enzymes are investigated by different biotechnological methods (direct DNA isolation, sequence-based techniques (Sanger sequencing (SS), capillary electrophoresis (CE), and next-generation sequencing (NGS), PCR methods) molecular fingerprint techniques (denaturing gradient gel electrophoresis (DGGE) and temperature gradient gel electrophoresis (TGGE)), fluorescent in situ hybridization (FISH), terminal-restriction fragment length polymorphism (TRFLP), single-strand conformation polymorphism (SSCP), etc. [51-53]. Several transport protein groups enable microorganisms to have the influx and efflux of different metal ions. This mechanism protects them from overtoxicity. Microorganism HM resistance is linked to encoded metal-binding peptides metallothioneins (MTs) and enzymatically produced phytochelatins (PCs). Also, there are various transporters belonging to Natural Resistance-Associated Macrophage Proteins (NRAMP), Zrt-Irt-like proteins (ZIP), Fe-Transporter (FTR), and Cu-transporter (CTR). The other group is responsible for HM exports from cells by the cation diffusion facilitator (CDF), P1B-type ATPases, FerroPortiN (FPN), and the families Ca (II)-sensitive cross-complementer1/ Vacuolar iron transporter1 (Ccc1/VIT1). Microalgae are well known for their successful removal of Zn. The most researched proteins are metallothioneins, which play a role in metal chelation. MTS has several classes with different roles; MTS class III is linked with successful Zn chelation. Although not that effective, MTs-class II is found in Chlorella, Aureococcus, Symbiodinium, Nannochloropsis, Thalassiosira, and Ostreococcus genera, also proven to be heavy metal-tolerant microorganisms [54]. Although MT-III is significantly more reactive than MTs-I/II, the class II MTs are polypeptides obtained from direct gene products and bind with Cu, Cd, Hg, and Zn [54]. Additionally, heavy metals can boost PAH biodegradation by enzyme-metal-substrate complexes. Since bioremediation of heavy metals is pollution concentration dependent. Being coenzymes, HMs stimulate a biological response in microorganisms towards PAH removal, even in a concentrated environment. Also, PAHs influence the microorganism’s membrane, changing the cell membrane permeability and the electrical potential of the cell membrane, etc., and enhancing HM intake [55]. When tackling PAH removal, enzyme-based metabolic pathways include genes encoding oxygenase, peroxidases, reductases, hydroxylases, monooxygenases, and dehydrogenases [56]. In PAHs, aromatic ring cleavage, side chain, and central aromatic processes participate in two types of oxygenases: cytochrome P450 monooxygenases (CYPs) and ring-hydroxylating dioxygenases (RHDs). Microorganism PAH degradation results in CO2 and H2O. Furthermore, redesigning microorganisms for specific purposes can be achieved using synthetic biology tools. A broad spectrum of sequence-specific nucleases, known as gene editing tools, have been applied so microorganisms with new abilities can accomplish a given task. To manipulate any genomic sequence, target DNA double-strand breaks (DSBs) at specific sites and, if needed, repair them by non-homologous end joining (NHEJ) or homologous recombination (HR) to give a specific sequence. Systems biology and gene editing tools used to enhance bioremediation processes are the CRISPR-Cas system, ZFN, and TALEN. While ZFN and TALEN use artificial endonucleases, CRISPR-Cas uses naturally occurring RNA-directed endonucleases. CRISPR stands for clustered regularly interspaced short palindromic repeat DNA sequences with Cas-added protein. There are 3 CRISPR-Cas types and several subtypes defined by the Cas protein. The CRISPR-Cas 9 system consists of Cas9, a DNA endonuclease, and two noncoding RNAs, crRNA and tracrRNA (guide RNA (gRNA)) [57]. CRISPR presents 30-40 bp of repetitive DNA sequences with “spacer” DNA sequences in between, with the repeats being translated into crRNA. CrRNA and Cas protein form crRNP (ribonucleoprotein), which causes a break in the intruder’s DNA/RNA. In the Cas9-gRNA complex, gRNA directs Cas9 to cut both strands of a targeted region of DNA, forming the DBS of a target site [58, 59]. Recent research on prokaryotic CRISPR-Cas systems is leading to models in which inactive and RNA-targeting Cas proteins are coupled to a variety of effector proteins to regulate gene expression and epigenetic modifications, additionally preventing mutations [60]. More advanced tools, TALENs, or transcription activator-like (TAL) effector nucleases, are originally secreted from the pathogenic bacterial genus Xanthomonas. The DNA-binding domains of TALENs form repetitive sequences isolated from Xanthomonas. TALEs combine the carboxy-FokI cleavage domain (C-terminal) and amino-DNA-binding domain (N-terminal). The advantage of TALENs lies in using a ‘protein-DNA code’ that joins DNA-binding TALE repeat domains to target one of the DNA bases at the binding site. Further, Zinc Finger Nucleases (ZFNs) are artificial restriction enzymes that consist of 3-4 domains, each approximately 30 amino acids long, organized in a ββα motif. Like TALEs, they have the carboxy-FokI cleavage domain (C-terminal) isolated from Flavobacterium okeanokoites. These ZFPs have between 9 and 18 bp that can target a wide range of possible DNA sequences and present a zinc-finger DNA-binding domain surrounding the cleavage domain, depending on the target site [60]. All three above-mentioned gene editing tools improved the process of bioremediation.
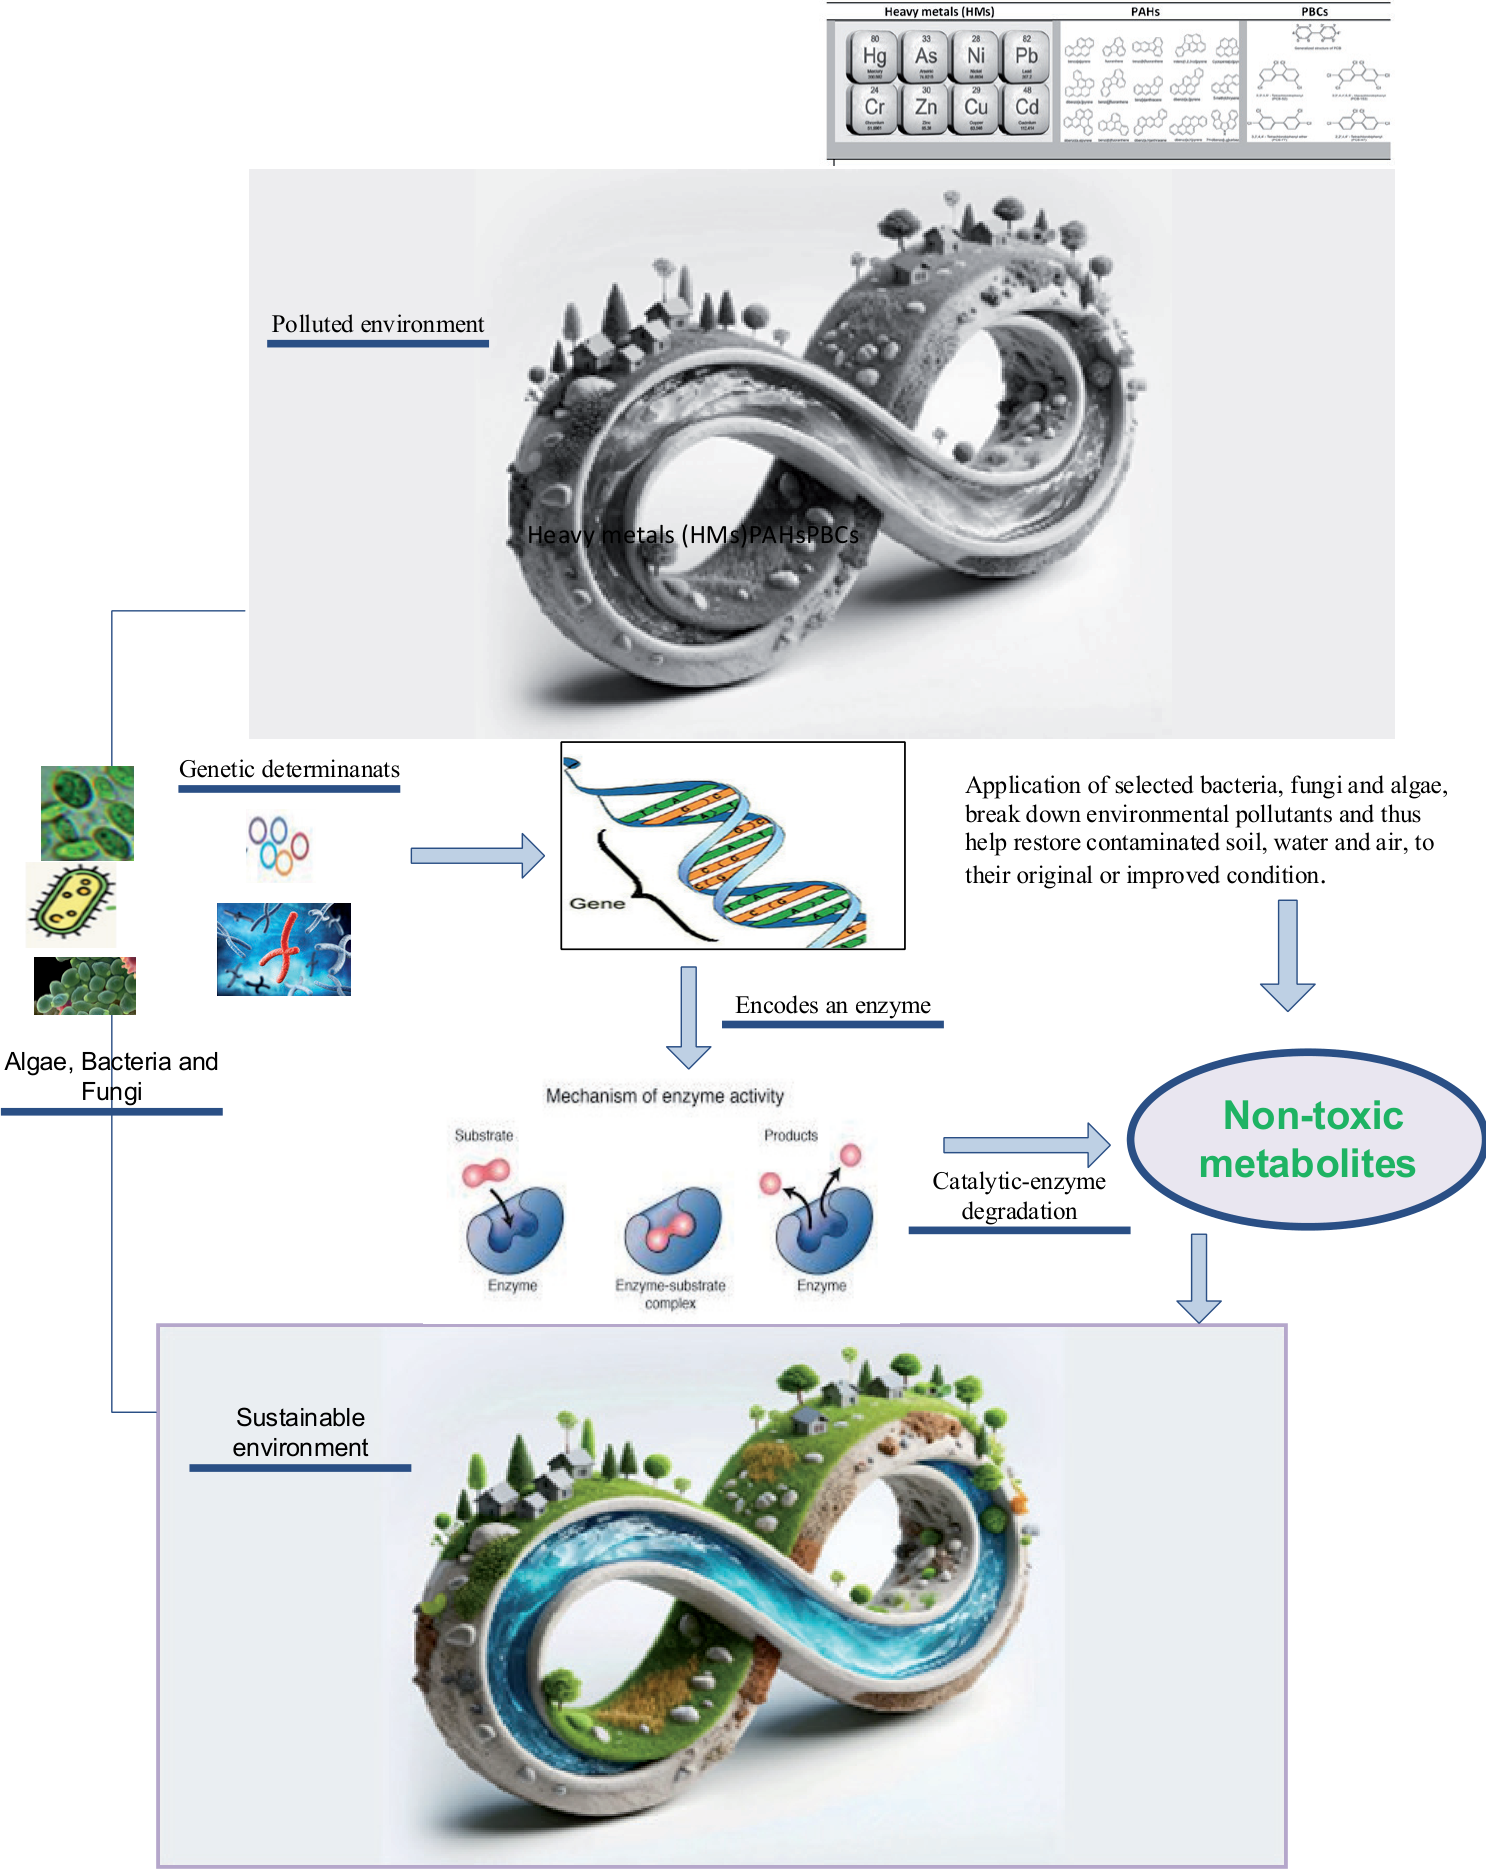
Pesticides are known for their high toxicity, biomagnification, and forming secondary pollutants if bioremediation isn’t completed correctly. Applying CRISPR-Cas, ZFN, and TALEN gene editing tools changes the outcome of bioremediation by improving microorganisms with functional pesticide degradation genes, improving pesticide degradation to less toxic compounds, thus preventing biomagnification [60]. To remediate the environment burdened with harmful compounds, the best thing to do is to heal it with organisms and inhabitants in that environment. If not equipped to break down pollution, researchers can help redesign them. This review aims to summarize the molecular aspects of bioremediation microbes, information gained via genome sequencing, and the phenotype plasticity of some microorganisms. Also, to present redesigned genetically modified microorganisms and specific genes that enabled them to remove target pollution. This data can then be used by researchers and enforced to facilitate decision-making. The natural methods applied in the environmental cleaning and maintenance process promise sustainability in every aspect (Fig. 1).
Results and Discussion
Implication of Microorganisms in Heavy Metal (HM) Removal
The pollution of soil and water caused by the release of heavy metals (HMs), such as chromium (Cr), cadmium (Cd), mercury (Hg), nickel (Ni), zinc (Zn), and lead (Pb), as a consequence, have adverse effects on the environment and human health. The issues related to HM cause nutrient imbalances and reduced soil quality, as well as persistent nature and bioaccumulation. Natural sources, such as minerals, weathering, erosion, and volcanoes, along with anthropogenic sources that include industry, mining, agricultural activities, emissions from improper waste management, etc., make this problem urgent to tackle [7]. Heavy metals affect genetic material because they compete to bind to the same sites on DNA as essential ions. Therefore, they are considered mutagenic and carcinogenic [47]. Microbial living environments play a significant role in their performance. The pH of each habitat and microorganism is specific. Changed pH can affect the activity of enzymes, HM adsorption in microorganisms, and mobility of heavy metal ions in the soil. Most bacteria’s best Pb2+ and Zn2+ removal performances are in the pH values 2.0-5.5 range. Also, it has been shown that pH below and above these values decreases microorganisms’ potential to remove HM. For Cu, Ni, and Pb removal, the best result showed Bacillus sp. and Pseudomonas sp. at pH 5 and Bacillus jeotgali U3 at pH 7 [61]. The temperature also plays a crucial role. Cd2+, Cr2+, and Zn2+ removal best capacity for Thiobacillus ferrooxidans, Thiobacillus acidophilus, and Thiobacillus tepidarius are at 25-27º C, while Sulfolobus solfa-tataricus and Acidianus brierleyi prefer extremely high temperatures. Bacillus licheniformis and Bacillus jeotgali U3 showed greater variations from 25 to 35 C [61]. The best capacity for Cu2+ and Cr6+ removal by Ochrobactrum intermedium LBr and Cupriavidus metallidurans CH34 was in the pH 6-7 range. While the temperatures differ, 37º C and 27ºC, respectively [62].
The substrate concentration is another important factor. Different microorganisms remove Pb at different environmental factors. Temp (ºC), optimal pH, initial Pb concentration (mg L−1), Pb removal rate (%), or max Pb uptake (mg g−1) by Pseudomonas stutzeri, Enterobacter cloacae, Serratia marcescens, Alishewanella sp. WH16-1 Microbacterium oxydans CM3, Pseudomonas sp., and Bacillus subtilis X3 was for 30ºC, pH 7, 103.5 (mg L−1)>99.0%; 30ºC pH 7 ,7.2 (mg L−1), 68%;28ºC, pH7 100 (mg L−1), 97.57%; 37ºC pH, 6 100 (mg L−1) 84.13%; 30ºC, pH7.59, 400 (mg L−1), 58.0%; 20ºC, pH 7, 50 (mg L−1),>90.0%, 37ºC, pH4, 300 (mg L−1) 192.05 mg g−1, respectively. In Fungus Temp (ºC), optimal pH, initial Pb concentration (mg L−1), Pb removal rate (%) or max Pb uptake (mg g−1) were: for Bacillus cereus 30ºC, pH 5, 100 (mg L−1), 75.6%; Penicillium polonicum 30ºC, pH 5, 828.8 (mg L−1), 90.3%; Spergillus tubingensis 30ºC, pH 5, 828.8 (mg L−1), >90.0%; Aspergillus niger 30ºC, pH 5, 828.8 (mg L−1), 97.0%; Trichoderma asperellum 30ºC, pH7, 250 (mg L−1), 18.71%; Rhizopus oryzae 30ºC, pH7, 450 (mg L−1) 33.76%; Mucor irregularis 30ºC, pH 7, 100 (mg L−1) 17.37% [63]. Heavy metals affect genetic material because they compete to bind to the same sites on DNA as essential ions. Therefore, they are considered mutagenic and carcinogenic [64].
Microorganisms are recognized as a powerful tool in bioremediation because of their natural and upgraded ability to degrade environmental pollutants via biochemical pathways. The genetic determinants involved in HM removal are presented in Table 1.
Chromium (Cr) Pollution
Chromium appears in the environment via natural and anthropogenic pathways. The two forms with positive and negative impacts on life forms are Cr(III) and Cr(VI) [65]. Industrial waste is a known source of toxic heavy metals such as hexavalent chromium. Because of high solubility and toxicity, Cr(VI) needs to be transformed into less toxic and soluble Cr(III). Chemically, graphene oxide (GO) polymers are found to be useful not only in absorption but also in reduction from Cr(VI) into Cr(III) [66]. The conversion of the highly toxic compound hexavalent chromium Cr(VI) into immobile and less toxic trivalent chromium Cr(III) can occur naturally. The chemical reduction process demands high energy, while biologically, the reduction process occurs in microorganisms and does not require high energy input [67]. Many species of microorganisms, Arthrobacter, Pseudomonas, Pannonibacter, Escherichia, Enterobacter, Bacillus, Shewanella, etc., have been reported to reduce Cr(VI) [67-69].
Although Cr (VI) enzymes reductase are divided into two classes, class I has a role in chromium reduction, while class II relates to other pathways. Class I is encoded by ChrR and YieF genes, and Class II by NfsA (NiR genes) [69]. Pannonibacter phragmitetus BB tolerates high concentrations of Cr(VI) because of the presence of chrA1 and chrA2 that encode for two chromate transporters (ChrA1 and ChrA2) and nitR genes that encode a chromate reductase [69]. Bacillus sp. SFC 500-1E can tolerate high concentrations of hexavalent chromium, which is related to the chrA gene that encodes for the chromate ion transporter protein (ChrA) belonging to the CHR protein family [70]. It was proven that the modified B. cereus NWUAB01 chromate transport protein encoded by chrA has a role in chromate reduction and transport. Similarly, the Bacillus sp. strain DHS-12(7) chromate reductase (ChrR) gene encodes chromate reductase protein [71]. Also, it has been revealed that the coexpression of ChrA and ChrT in ChrAT-modified bacteria is linked to enzymatic activity via NADH and NADPH [71, 72]. The ChrA transporter, a membrane protein, has also been detected in Bacillus cereus strain TCL [72, 73]. Interestingly, lowering the concentration of Cr(VI) within a cell without changing its valence state (two chromate transporters) only protects cells/microorganisms but does not remove environmental toxins. Therefore, chromate reductase reduces heavy metal pollution in the environment. Also, the locations of genes that encode the chromate reductases (ChrR, YieF, NfsA, NemA, and LpDH) differ. Thus, in the cytoplasm (soluble chromate reductases) and cell membrane (membrane-bound chromate reductases), the reducing conditions are related to aerobic and anaerobic circumstances [69]. Furthermore, chrA genes can be found on plasmids as well as in chromosomal DNA. Therefore, as expected, they are organized in operons with other chr genes; for instance, in C. metallidurans, they are organized in the chrFECAB cluster as a part of a plasmid, or they are a part of the chromosome as the chr2 cluster (chrB2, chrA2, chrF2 genes). Furthermore, chr genes, chrB, chrA, chrC, and chrF found in a transposable operon (the chrBACF cluster) in Ochrobactrum tritici 5bv11 are involved in Cr resistance [5, 75]. A ChrR gene in Stenotrophomonas maltophilia encodes a protein Chromate Reductase (ChrR) homologous to other chromate reductase enzymes [76] that also provides resistance and high Cr(VI) reduction in the natural environment. Furthermore, Stenotrophomonas maltophilia, at the optimum 370C at pH 7, showed the ability to remove and reduce up to 100%, making it a suitable candidate for bioremediation. The Cr(VI) concentration between 10 and 2000 mg/L showed 100% removal at concentrations in the 10-70 to 92% at 500 mg/L [76]. The transportation and reducing ability of transport proteins and Cr-reductase have been used to modify microorganisms that live in contaminated sites but have no Cr (VI) resistance. Transfer of Cr-transporter chrA genes and Cr-reducing yieF genes to Comamonas aquatica makes this genetically modified strain a very effective bioremediation microorganism [77]. Also, in chromium removal, there are enzymes nitrate reductase (NR) and nitrite reductase (NiR), which are related to N cycles in microorganisms. These enzymes, along with the nitrate and nitrogen intake in the NADP and NADPH domains, reduce hexavalent chromium and have been found in different microorganisms. The genes that encode for these enzymes are found in NR (Pisolithus sp. 1 NR) and NiR (Pisolithus sp. 1 NiR) of Pisolithus sp. 1 [78]. The text below contains information about identifying specific genes in Vibrio harveyi KCTC 2720. It also discusses the similarity of the reducing capacity of enzymes encoded by these genes to the nfsA gene found in E. coli [79]. Additionally, it mentions that some genes enable resistance to Cr(VI) in Ralstonia metallidurans chr2, specifically within the chrB2 and chrA2 genes, the niaD gene is the first to encode Cr(VI) reductase, and it has been identified in Vibrio harveyi KCTC 2720. Moreover, enzymes encoded by genes related to Nassimilation showed the same reducing capacity, such as the nfsA gene found in E. coli [79]. Apart from reducing capacity, some genes enable Cr(VI) resistance. In Ralstonia metallidurans, chr2, as a part of chrB2, chrA2, and chrF2 gene clusters, has a role in chromate resistance. The chromate efflux pump encoded by the chrA gene, the previously known chr1, belongs to Ralstonia metallidurans plasmid pMOL28 genes. These genes, along with chrI, chrB1, chrA1, chrC, chrE, chrF1, and chromosomal chr2 cluster (genes chrB2, chrA2, chrF2) enable Cr resistance [80]. Another promising bacterium, Shewanella oneidensis MR1, with chromosomal gene chrASO, encodes the chromate efflux pump protein. Furthermore, there are genetically modified strains, such as modified E. coli (heterologous expression techniques), with increased Cr(VI) reduction [81].
Table 1.
The molecular aspect of HM removal.
Nickel (Ni) Pollution
Nickel is an essential transitional trace heavy metal; being a micronutrient, it is found abundantly in the environment. Anthropogenic activities opposite from natural appearance have led to toxic levels of Ni in ecosystems. Then, it becomes lethal and immunosuppressive instead of essential for the metabolic activities in living cells. Bacillus thuringiensis KUNi1 was found to be resistant to Ni up to 82%. Furthermore, this strain is also resistant to zinc (Zn), copper (Cu), and cobalt (Co) at the same level of resistance, while in cadmium (Cd), small activity is shown [82]. Exploring the potential of E. coli in the Ni resistance mechanism, the nik operon has been identified. The nik operon consists of 6 genes (nikABCDE) that regulate Ni cell concentration. The first five encode for the ABC transporter system (ABC-type nickel permease) linked to translocation, whereas genes nikB and nikC encode for proteins linked to the chemiosmotic gradient, while nikD and nikE encode for proteins that need energy provided by hydrolysis of ATP to translocate Ni. Interestingly, the first gene in the cluster, nikA, relates to chemotaxis [82-84]. Another gene, also part of this cluster, steps out, a nikR gene that can function as a repressor of the nik operon and as an activator of gene transcription [85, 86]. The transporter family encoded by nikABCDE in Bacillus altitudinis MT422188 is regulated by the ABC transporter family, ATP independent or ATP dependent [87], also known as ATP-binding cassette systems encoded by the same nikABCDE operon in Bacillus altitudinis FD48 [88, 89]. Furthermore, in Ni2+ resistance, secondary permeases of the NiCoT family (Ni-tolerant genes) were found in Microbacterium oxydans strains CM3 and CM7 and Microbacterium paraoxydans. The M. paraoxydans Ni resistance is linked to three members of Ni-binding proteins with different locations. The nikABCDE system includes NikA (periplasmic), NikB, and NikC (inner membrane) proteins involved in Ni uptake and bioaccumulation. Additionally, in Microbacterium paraoxydans, genes involved in Zn tolerance include nikC, an inner membrane protein, then znuB integral membrane protein, and zur negative regulator [90]. Engineered Escherichia coli bioaccumulate high concentrations of Ni by its native NikABCDE transporter and a heterologous, cysteine-rich metallothionein from the plant Pisum sativum [91]. Another group of genes (CDSs) showed involvement in Ni resistance. In the Cupriavidus sp. Ni-2 genome, 12 CDS genes encoding CnrA, CnrB, CnrY, NccB, NccC, and NccN proteins related to Ni resistance, and NccX2+ proteins related to Ni transport were identified [92]. In addition, various members of the HoxN-family transporters proteins are involved in the Ni2+ resistance mechanism. For instance, HoxN is found in Ralstonia eutropha and used in Streptomyces sp., which are very effective microorganisms for high Ni concentration removal [93]. Actinomycetes showed a high ability to remove organic and inorganic waste from polluted soil and water. The focus can be directed towards the Rhodococcus genus because of its many-sided ability to produce enzymes and biosurfactants and degrade a wide range of pollutants. Rhodococcus opacus and Rhodococcus rhodochrous were identified as successful in Ni2+ and Pb2+ removal [94].
Lead (Pb) Pollution
Anthropogenic activities related to waste batteries, pigments, lead arsenate insecticides, lead water pipes, manufacturing, mining, industrial wastewater, pesticides, etc., release high amounts of lead into the environment [95]. Lead is widely recognized as a ubiquitous and very toxic metal, proven to inhibit enzyme activity, damage DNA, and disrupt cell membrane permeability in microorganisms. Metallothioneins, cysteine-rich proteins encoded by plasmid genes, are involved in the sequestration or bioaccumulation of toxic metals within the cell [96]. Bacteria exhibit several detoxification mechanisms to reduce the harmful effects of heavy metals and protect themselves [97]. Several lead-resistant bacteria, including Klebsiella michiganensis R19, Providencia rettgeri L2, Raoultella planticola R3, Serratia sp. L30, Pseudomonas pseudoalcaligenes, and Micrococcus luteus are found to be useful in Pb removal [98]. A genetic determinant, a PbrR operon, activates a group of transcriptional regulators called the MerR family that controls heavy metal toxicity. This was found in Cupriavidus metallidurans CH34, which showed resistance in the presence of lead (II) [99, 100]. Another cluster of the pbrABCD genes also codes for Pb, Cd, and Zn transport proteins. Thus, pbrAB genes code for PbrA protein, a type of ATPase protein, and PbrB, an undecaprenyl pyrophosphate phosphate, with a role in lead sequestration. Notably, the Pseudomonas fluorescens M114 pbrA gene is responsible for the cellular response to iron via pseudobactin M114 synthesis [101]. A high-affinity iron-chelating Pseudobactin B10 regulated by the pbrA gene is also identified in Pseudomonas strain B10 and Pseudomonas aeruginosa PAO1, while in Pseudomonas aeruginosa mutant PALS124 (pvdA), it is encoded by the psbA gene [102]. The soil bacterium Pseudomonas sp. NCIMB10586 showed strong iron toxicity resistance, producing two mupirochelin siderophores by NRPS genes mchA and mchB [103]. Apart from bacteria, some microalgae are also known as organisms with high heavy metal tolerance. For instance, the genera Chlorella and Scenedesmus with certain species (C. reinhardtii, C. sorokiniana, and S. obliquus) can tolerate to some extent Pb(II), Cd(II), Cu(II), and Cr(VI), depending on the cell structure and metabolism [104]. Several transport protein groups enable the influx and efflux of different metal ions. This mechanism protects cells from overtoxicity. The study related to the Pb removal taxon, Chlorophyta with Chlamydomonas reinhardtii, Chlorophyta, Chlorella vulgaris, Oedogonium sp., and Pseudochlorococcum typicum has shown promising results [105]. Also, C. acidophila PM01 showed substantial tolerance to high concentrations of Cd, Cu, and Zn, with a promising increase in bioaccumulation of Cu due to metabolic adaptation [106].
Zinc (Zn) Pollution
Zinc is an essential element due to its role in bacterial resistance related to plasmids. The resistant strategy mechanism in Gram-negative and positive bacteria includes efflux, sequestration, chelation, and metal uptake. Bacteria’s main efflux proteins are the Cation Diffusion Facilitator (CDF), Resistance Nodulation Division (RND) efflux pumps, and P-type ATPase families [107]. A Zn resistance can be linked with three structural genes (czcABC) that encode RND (CzcA, CzcB) and CDF (CzcD) efflux pumps for Zn2+ transport out of the cell, and they are found in both Gram-positive and negative bacteria. In Streptococcus agalactiae, resistance is related to czcD genes that encode an efflux protein [108], as well as the CPx-type (CadA) efflux proteins found in Gram-positive bacteria. For the metal Zn uptake, Gram-positive bacteria have P-type ATPase (ZosA), znuABC, and membrane proteins (Yci ABC, Ycd H1). However, the P-type ATPase protein ZntA participates in the efflux of Zn2+ in both Gram-positive and negative bacteria [108]. Furthermore, the ABC transporter family belongs to the high-affinity Zn2+ uptake systems and presents members of the ATP-binding cassette, with znuA, znuC, and znuB proteins encoded by znuABC genes [109]. In sequestration, the role has periplasmic or cytoplasmic proteins [108]. In Escherichia coli, Zn detoxification is related to the CDF family with the ZitB and YiiP proteins, one P-type ATPase, and the ZntA protein. In Staphylococcus aureus, resistance enables the CDF family. In Caulobacter crescentus, czrCBA and nczCBA belong to the RND efflux systems. C. crescentus, CzrCBA system exports Zn ions from the cytoplasm of cells, similar to the CzcABC system, while in Ralstonia metallidurans, there is the CzcS/CzcR two-component regulatory system. In Streptococcus pneumoniae, that is CzcD, a member of the CDF family [109]. Bacteria possessing MTs are an ideal tool for the Zn and Cd bioremediation. The marine cyanobacterium, Synechococcus sp. strain RRIMP N1, and a freshwater Synechococcus TX-20 can remove Zn by MT class II encoded by the smtA gene. Another important gene is the smtB gene, which plays a role in the metal-dependent repression of the smtA gene. In Zn and Cd removal, Escherichia coli, Staphylococcus aureus, and Staphylococcus xylosus SmtB gene and ArsR proteins are also involved. Also, bmtA and bxa1 genes of Oscillatoria brevis encode metallothionein (BmtA) and CPx-ATPase (Bxa1) [110, 111]. A CPx-ATPase-like ABC transporter, CDF families, etc., export metal ions out of cytocells [111]. Thus, these microorganisms are capable of chelating and exporting metal ions, therefore protecting themselves and their habitat.
Mercury (Hg) Pollution
Mercury (Hg), previously known as hydragyrum, stands out with a unique ability to keep liquid form at Earth’s temperature and pressure [112]. Recognized as a metal of great toxicity, its removal from the environment is an issue to be tackled. In addition, Hg has no identified biological role opposite to Fe, Cu, Zn, Se, Cr, etc. [113]. The ability of some microorganisms to live in the presence of high concentrations of Hg includes some chemical or biological change of mercury [114]. One of the mechanisms is the reduction of Hg2+ to Hg0, which is controlled by the mer operon, a cluster of genes involved in Hg transport, reduction, and volatilization [115]. A cytoplasmic enzyme, mercuric reductase, and an organomercurial lyase are encoded by merA and merB genes, respectively [116]. Besides merA and merB genes, identified mer operons with merR, merP, merT, merD, merF, and merC encode for different inner-membrane proteins with different roles in Hg transport and metabolic pathways. Namely, merR encodes for MerR, the metalloregulatory protein, a regulator of the operon transcription in the presence/absence of Hg2+; merD encodes for the MerD regulator protein, a secondary regulator protein with a similar but weaker influence on the operon transcription [116]. Other important genes are merT and merP, and sometimes with merF, help Hg2+ transport into the cytoplasm [117, 118]. Hg+2 to Hg0 reduction mechanism can be independent of the mer system and has been found in Geobacter sulfurreducens PCA, Geothrix fermentans H5, Cupriavidus metallidurans AE104, Shenwella oneidensis MR-1, and Geobacter metallireducens GS- 15. The primary drivers of Hg methylation are the corrinoid protein, HgcA, and a ferredoxin, HgcB, encoded by hgcAB genes [119]. The diversity of HgcAB-encoding microbes is reflected in aerobic nitrite-oxidizing bacteria Nitrospirae and Chloroflexi (Dehalococcoides) [120, 121]. Important methylating genes, hgcAB, are also found in mercury methylators from Deltaproteobacteria, Firmicutes, Chloroflexi, Phycisphaerae, Aminicenantes, Spirochaetes, Elusimicrobia, Lentisphaerae, Bacteroidetes, and Atribacteria [121, 122].
Cd Pollution
Among heavy metals, Cd is classified as highly toxic and difficult to degrade; thus, pollution with this metal is serious. Its effect on living organisms is reflected in reducing the biological activity of soil microorganisms and plants, which can be linked to animal and human poisoning by the food chain [123]. Many microorganisms have developed resistance to this metal, and their activity is not disturbed [123]. Staphylococcus aureus strain is known for its natural Cd resistance; Cupriavidus metallidurans, Escherichia coli, and Bacillus subtilis are also applied to Cd removal. The basis for the resistance mechanism lies in the presence of specific resistance genes or operons like cad, czc, and znt. These genes encode for P-type ATPases and cation diffusion facilitators (CDF) that transport Cd from the cytoplasm to the periplasm and RND-driven efflux systems that transport Cd from the periplasm to outside the cells. Therefore, export from the cell is complete [124]. Some studies indicate the role of strain Y8/ genome with more than 75% guanine-cytosine content (CG). According to Chen et al. (2019a, b), metal-resistant strains such as Cupriavidus, Thiobacillus, Deinococcus, and Comamonas spp. have an average GC content of 66.2%, 62.6%, 67.3%, and 61.5%, respectively [125, 126]. Genes identified as Cd metal resistance genes zntA of E. coli and copA of Legionella pneumophila, along with the ACR3 family genes, encode for two of four heavy-metal transporting P-type (PIB) ATPases [124]. These transporters are essential in cell detoxification, transporting heavy metal ions across cellular membranes [127]. Interestingly, genes czcD of Cellulomonas metallidurans, cutC of Enterococcus faecalis, and znuA of E. coli are homologous to genes identified in Y8 strains [124]. Specifically, zntAY8 and czcDY8 in Y8’s E. coli zntAY8 and the genus Cellulomonas with (zntAY8, copAY8, HMTY8, and czcDY8). Furthermore, the recombinant E. coli metallothionein gene, cloned from Neurospora crasa belonging to the fungal class Ascomycetes, is linked with Cd uptake. Furthermore, cadA and norVW operons encode for CadA and NorVW involved in Cd resistance in the highly cadmium-resistant strain Enterobacter cloacae LPY6/ domesticated E. cloacae Cu6 [128]. Genetically modified Pseudomonas putida KT2440 showed great ability in Cd removal. The PC synthase gene (PCS) originated from the fungus Schizosaccharomyces pombe and was cloned and expressed in P. putida KT2440. The PC synthase gene (PCS) encodes for phytochelatins (PCs), an enzyme rich in the cysteine thiolate groups, the primary binding sites for heavy metals [129, 130]. Pseudomonas putida’s ability to remove copper and cadmium lies in plasmid-encoded heavy metal ion resistance [131]. Furthermore, the Microbacterium paraoxydans gene cadA encodes for Cadmium P-type ATPase [73]. Burkholderia sp. SRB-1 genome contains metal-exporting genes zntA, czcA, czcB, and czcC, and they encode for Zn2+ /Cd2+-exporting ATPase, heavy metal efflux system protein, membrane fusion protein, and outer membrane protein, respectively. Additionally, detoxification dsbA and cysM genes encode for the proteins dithiol oxidoreductase and S-sulfo-L-cysteine synthase, and genes katE, katG, and SOD1 encode for catalase, catalase-peroxidase, and superoxide dismutase, Cu–Zn family involved in Cd antioxidation/antioxidative enzyme activity [132]. The phosphate-mineralizing bacteria Enterobacter sp. shows high Cd resistance via mechanisms that include Cd ion transport and efflux, as well as the reduction and antioxidation potential. Genes identified in PMB bacteria encode proteins of the Cd resistance mechanisms. Enterobacter sp. genes czcD, MgtA, and zntB encode for ion metal transport protein, SOD2, and katE, encode for enzymatic detoxification and dsbA, cysM antioxidant enzymes [133]. Different efflux pump proteins encoded by cadA and cadB plasmid R genes belonging to Staphylococcus aureus also play a crucial role in Cd bacteria resistance [134, 135]. Alcaligenes eutrophus and Ralstonia sp. CH34 are the Cad systems encoded by the CadAB operon for cadmium resistance [135, 136]. The cation diffusion facilitators (CDF family) resistance to Zn2+ and Cd2+, encoded by CzcD from R. metallidurans strain CH34, decreases the concentration of Zn2+ and Cd2+ in the cytoplasm [137]. Similarly, Acidithiobacillus ferrooxidans czcA1 A.f, czcA2 A.f, czcB1 A.f, and czcC1 A.f genes belong to the cadmium efflux system (the Czc system) [138]. Furthermore, in Pseudomonas putida KT2440, Cad and Czc systems are also identified in Cd resistance. Namely, a regulator of the CzcRS3 system activates czcCBA1 and czcCBA2 genes that encode efflux pump protein [139]. Additionally, Cad genes of Bacillus cereus strain NWUAB01. also encode for cadmium transporting ATPase, cadmium efflux system accessory protein, and cadmium resistance transporter, all belonging to Cd transport proteins. Cytoplasmic copper homeostasis proteins encoded by cutC with copper resistance proteins encoded by CopC or CopD have a role in copper resistance transport. CorA genes encode for CorA proteins, CorA-like magnesium transport proteins, and magnesium transport proteins, all of which are involved in Mg transport. The manganese transport protein encoded by the MntH gene has a role in manganese transport. Cobalt–zinc–cadmium resistance protein encoded by the CzcD gene has a role in cadmium, cobalt, and zinc transport. Zinc ABC transporter is a periplasmic binding protein encoded by the ZnuA gene that has a role in zinc transport. Both CzcD and cadA operons are energy-dependent efflux systems [139], similar to the PbrA efflux protein, which gains energy by ATP hydrolysis and is encoded by the PbrA gene [140, 141]. The CzcR and CzcD genes encode for the CzcR and CzcD proteins that are implicated in resistance to Co2+, Zn2+, and Cd2+ (czc system) [142]. Pseudomonas aeruginosa T-3 has plasmid-encoded chromate ion resistance [143, 144], as well as Pseudomonas putida PhCN for Cd ion. Furthermore, Pseudomonas aeruginosa strain ZM130 resistance is linked with the enzyme laccase and nicotinamide adenine dinucleotide (reduced)-dichlorophenolindophenol reductase (NADH-DCIP reductase) [145].
Persistent Organic Pollutants POPs-Polycyclic Aromatic Hydrocarbons (PAHs) and Polychlorinated Biphenyls (PCBs)
Toxic, hydrophobic, and lipophilic persistent organic pollutants (POPs) can be found in soil, water, and air, as well as in living organisms. These include many manufactured chemicals, but the 3 main groups are polycyclic aromatic hydrocarbons (PAHs), polychlorinated biphenyls (PCBs), and pesticides [146]. Besides biochemical pathways, the physicochemical parameters (pH, temperature, and substrate concentration) play a role in microorganism performance. Benzo[a]pyrene (PAHs), degraded by Sphingobacterium spiritivorum and Aspergillus brasiliensis, was performed at pH 6 and 40 mg/L benzo[a]pyrene concentration and pH 6.41 and 38.8 mg/L benzo[a] pyrene concentration, respectively [147]. Although fungi prefer low acidic and high alkaline environments, it was noticed that a soil pH of 7.5 was the most suitable for the degradation of all the PAHs, reaching 50. Penicillium sp. prefers acidic pH, whilst Aspergillus sp. was found to be effective at pH 7.5-8 [148]. In POPs degradation, the best-removing capacity is Pseudomonas putida F6 at 30ºC, optimum pH 7, and Streptomyces cyaneus at 60ºC, optimum pH 4.5. Bacillus sp. showed variations depending on the organic compound in the temperature between 30-37º C, optimum pH 5-8, and at similar conditions, Mycobacterium sp. degrading xenobiotics, Sphingobacterium sp. PLA, and Chromobacterium viscosum PBC [149]. The widespread use of many of them, as a consequence, has led to bioaccumulation and magnification. The problem needs to be addressed in a way that cleans but does not harm ecosystems; therefore, microorganisms present the best solution for POP removal. The genetic determinants involved in POP bioremediation are presented in Table 2.
Implication of Microorganisms in Polycyclic Aromatic Hydrocarbons (PAH) Removal
Polycyclic aromatic hydrocarbons (PAHs) are a class of organic compounds produced by incomplete combustion or pyrolysis of fossil fuels (oil, petroleum, gas, coal, and wood). PAHs are a type of organic compound that consists of 3 benzene rings (low molecular weight PAHs) or more (high molecular weight PAHs) [150, 151]. There are many methods of removing PAHs from the environment. Chemically, enzymes oxidize PAHs to dihydrodiol, then dioxygenase→intradiol and extradiol cleavage→protocatechuates and catechol [152].
Biologically, specialized microorganisms isolated from their natural habitats successfully break down PAHs in the oxidation process. The outcome is the conversion of hydrocarbon molecules into carbon dioxide, biomass, or other harmful molecules. The catabolic pathway includes catabolic genes that encode enzymes involved in mineralizing persistent organic pollutants. Several microbial communities have been applied in PAH removal strategies with nidA/B, pah, and phd genes, designated as important biomarkers of pyrene metabolism [153]. An abundant pollutant, pyrene, belongs to high molecular weight PAHs with a tetracyclic structure, and bacteria can very successfully remove it from different phyla. For instance, Mycobacterium spp. is recognized as highly effective, along with other genera, such as Pseudomonas sp. JPN2, Staphylococcus sp. BJ06, Bacillus vallismortis, Diaphorobacter, and Pseudoxanthomonas. Wang et al. (2023) applied metagenomics analysis to identify the degradation pathways of the identified genes and their contribution to this process. The consortium of bacteria involved in pyrene degradation consists of Proteobacteria (99.8%) and Alphaproteobacteria (96.6%). This analysis revealed that nidA and nidB genes encoding PAH dioxygenase α/β subunits catalyze the first step in the degradation process. Duplication of the dioxygenase nid gene nidA/nidB and nidA3/nidB3 plays a key role in the initial degradation of pyrene. The following steps in degradation involve genes such as phdE, phdF, phdG, phdK (phenanthrene), nidD (pyrene), phtAa/Ab/Ac/Ad, phtB, phtC (phthalate), and pcaG/H [154]. Although highly effective, applying only one genus is yet to be proven to be the best bioremediation/solution. For instance, the PAH-removing consortium of anaerobic bacteria affiliated with Diaphorobacter and Paracoccus genera showed high removal capacity for acenaphthene and phenanthrene. A myriad of different genes, pht2/3, pht4, and pht5 (phthalate degradation) and ligA/ligB (protocatechuate degradation) was detected in Achromobacter spanius Devosia sp., Pusillimonas caeni, Eoetvoesia caeni, and Bradyrhizobium sp., Bosea sp., Afipia sp., Steroidobacter sp., and Mesorhizobium sp. [155]. The newly discovered strain Pseudomonas veronii SM-20 showed remarkable results in degrading/ oxidizing phenanthrene [156]. Apart from genera (genomes) that naturally degrade aromatic compounds, horizontal gene transfer enables the employment of different bacterial consortiums. Similarly, the presence and influence of nidA in Mycobacterium gilvum PYR. GCK [157], the nidA/B gene in Mycobacterium vanbaalenii PYR-1 [158], and the nidA, nidA3, pdoA2, and pcaH genes were identified [159]. The Pseudomonas putida G7 (nahAa, nahAb, nahAc, and nahAd) nahAb gene that encodes for the naphthalene dioxygenase protein is involved in naphthalene degradation and is typical for all nah-type degrading bacteria. The NahR protein is a transcription factor encoded by the NahR gene that activates the NAH7 plasmid operons [160]. Another strain, Pseudomonas fluorescens AH-40 with the nahAcR gene, was involved in naphthalene degradation [161]. P. putida strain G7 (pNAH7), where the naphthalene conversion proceeds via salicylate (nah-operon, nahABCDEF) and catechol is further oxidized by the meta-pathway (sal-operon, nahGHIJKLM). Pseudomonas-like organisms with a nah-like genotype were identified using genetic markers with nah-like sequences. In this way, markers nahAc from P. putida G7, phnAc from Burkholderia RP007, and the GST-encoding gene from Sphingomonas WP01 were revealed. The analogs of the bacterial glutathione S-transferase GST-encoding gene from Sphingomonas paucimobilis EPA505 were also identified [162]. Furthermore, P. putida KT2440 NAH7 plasmid genes are under salicylate control, thus encoding for enzymes involved in naphthalene (PAH) mineralization/degradation [163]. In naphthalene degradation, two NAH7 operons (gene clusters) participate with genes nahA-F and nahG-M that encode enzymes for the metabolism of naphthalene to salicylate and salicylate to tricarboxylic acid, respectively [164]. In Mycobacterium vanbaalenii PYR-1, nidB2 and nidB-nidA gene clusters were similar to phtAa and phtAb. They encode α/β-subunits of phthalate dioxygenase, involved in the catabolic pathway of phenanthrene and pyrene. Furthermore, Mycobacterium sp. 6PY1 pdoA1/ pdoB1 and pdoA2/pdoB2 genes are found to be similar to Mycobacterium vanbaalenii PYR-1 nidA and nidB genes. The gene cluster, including nidB2, nidB/nidA, and nidA3/nidB3, also encodes pyrene/fluoranthene/ naphthalene/phenanthrene ring-hydroxylating dioxygenase α- and β-subunits [165]. In the same fungi, for instance, in Rhodococcus sp., naphthalene degradation participates only in the narAa, narAb, and narB genes, which act as single units not organized in clusters, and they are regulated by narR1, narR2, and GntR-like transcriptional regulators. The narAa and narAb genes code for the α- and β- subunits of the naphthalene dioxygenase (NDO) with no similarity to genes in P. putida that encode NDO subunits [166]. Furthermore, Rhodococcus sp. P14 is mentioned in several polycyclic aromatic hydrocarbons via serine hydrolase, hydratase, alcohol dehydrogenase, protocatechuate 3,4-dioxygenase, β-ketoadipate CoA transferase, and β-ketoadipyl CoA thiolase. These enzymes are linked to ring cleavage, side chains, and central aromatic processes. Eight of the thirteen MAGs were confirmed to carry genes for PAH degradation and were affiliated mainly with Actinobacteriota and Proteobacteria. For instance, genes pcaG and pcaH are encoded for two subunits of protocatechuate 3,4-dioxygenase, the enzyme essential in vanillin catabolism in Pseudomonas strain HR199 [167], and in many xenobiotic metabolic reactions in Stenotrophomonas maltophilia KB2 [168]. These genes were also found in three other retrieved MAGs affiliated with Paracoccus denitrificans, Devosia, Pusillimonas caeni, and Eoetvoesia caeni. Also, genes vdh and vanAB, pbdA, pbdB, and pcaL encoding vanillin dehydrogenase and vanillate O-demethylase participate in vanillin catabolism, and they were found in Rhodococcus jostii RHA1 [169]. In addition to pcaG and pcaH, genes phdI, phdJ, phtAc, and phtC were identified in Mycolicibacterium sp. and Mycobacterium vanbaalenii PYR‐1, encoding for enzymes that participate in pyrene degradation and metabolism [170]. In pyrene degradation and metabolism, several enzymes are encoded by previously mentioned genes isolated from different organisms: oxygenase ferredoxin component (phtAc) from Arthrobacter keyseri 12B, ring cleavage dioxygenase (phdI) from Nocardioides sp. strain KP7, and phdF from Sphingomonas sp. strain KA1, hydratase-aldolase (PhdJ) from Nocardioides sp. strain KP7, and decarboxylase (phtC) from Arthrobacter keyseri 12B [171]. An S. paucimobilis 20006FA shows similarities in the gene clusters bph and xyl with the plasmid pNL1. This plasmid with the target genes is found in the Sphingomonas sp. and Sphingobium sp. C100 [172] and Sphingobium yanoikuyae B1 genomes [173, 174]. Genes belonging to Sphingomonas paucimobilis 20006FA, xylQ, nahE, and bphK encode for enzymes involved in phenanthrene degradation, namely, acetaldehyde dehydrogenase, a putative 2-hydroxy benzyl pyruvate aldolase, and glutathione S-transferase, respectively [175]. Besides, cytochrome P450 monooxygenases (CYPs) and laccase (the MtL enzyme) show a high ability for PAH degradation. The MtL oxidoreductase enzyme belongs to ligninolytic enzymes and is characteristic of fungi. For bioremediation, originally found in Aspergillus oryzae, the laccase MtL gene was transferred to Myceliophthora thermophia [176]. Apart from the classical catabolic nah-like genes, the phn genes are detected on the phn locus in Burkholderia sp. strain RP007. The cluster phnABC was under the regulatory control of PhnR and PhnS, encoding for different enzymes. While Burkholderia sp. strain RP007 phnABC cluster showed selective oxidation of either naphthalene or phenanthrene, transgenic E. coli phn enzymes oxidized both naphthalene and phenanthrene [177]. Furthermore, alkB and alkH, besides C12O and C23O, showed high catabolic potential in PAH removal. Bacterial strains Enterococcus faecalis (MK298392.1), Bacillus albus (MT325971), Enterococcus faecalis (MT325972), Providencia vermicola (MT325973), Enterococcus faecalis (MT345788), Carnobacterium gallinarum (MT350233), and Pseudomonas aeruginosa (JF756593.1) isolated from landfill soil artificially polluted with waste engine oil showed increased alkane hydroxylase gene (alkB) and aldehyde dehydrogenase gene (alkH) activity [178].
Table 2.
The molecular aspect of POPs removal.
Polychlorinated Biphenyls (PCBs)
Polychlorinated biphenyls (PCBs) are aromatic synthetic organochlorine chemicals with widespread use in transformers and electrical capacitors. PCBs are also used as hydraulic fluid, oil additives for paints, etc. As the name indicates, this biphenyl molecule consists of two six-carbon rings linked by a single carbon-carbon bond. In the PCB molecule, hydrogen is substituted with chlorine and bonds to 12 carbon atoms [179]. The phenol degradation pathway includes the activity of estradiol-cleaving and estradiol-cleaving enzymes encoded by plasmid genes. Some of the enzymes involved in PBC degradation are catechol 2, 3-dioxygenase (meta-fission), catechol 1,2-dioxygenase (ortho-fission), and phenol hydroxylase. Namely, phenol hydroxylase encoded by pheA and pheB was found in Klebsiella pneumoniae ΙΙ, Pseudomonas putida, and Bacillus subtilis. Furthermore, when testing enzymatic activity, it was found that the highest performance had phenol hydroxylase compared to both catechol 1, 2-dioxygenase and catechol 2,3-dioxygenase with the same enzyme activity [180]. The degradation pathway of phenol in the Marinobacter HA2 strain pathway of phenol degradation can be described via the 2,3 ortho-cleavage reaction, while the Halomonas HA1 strain involves the 1,2 phenol meta-cleavage mechanism. In bacteria, the major catabolic aerobic pathway of PCBs is an estradiol cleavage of the aromatic ring with the production of chlorobenzoate. Several genes, usually designated as bph, encode the key enzymes involved in this pathway. A mixed microbial population (7 bacteria and 1 fungus) was isolated from a soil sample of an unexposed natural environment. After enrichment and establishment in a defined medium containing p-CB (p-chlorobiphenyl) as the sole carbon source, this synthetic organic compound could be utilized. One of the microorganisms was engineered P. putida (pUC18) with the pUC18-nahH gene with catechol 2,3-dioxygenase (C23O), an enzyme able to eliminate phenanthrene and pyrene compounds [181]. The catechol 2,3-dioxygenase (C23O) gene isolated from Pseudomonas sp. strain ND6 showed high similarity with the Pseudomonas putida G7 NAH 7 plasmid, 93% in amino acid sequence, and with the Pseudomonas stutzeri AN10 chromosome, 89% in amino acid sequence [182]. In the same strain, the C12O genes (catA, catAI, catAII, and catAIII) showed involvement in naphthalene catabolism, with the catechol 1,2-dioxygenase isoenzymes encoded by these genes [183]. Interestingly, some bacteria had no genetic basis for the naphthalene-degradative pathway. The main role in this process was played by primers for 1,2 and 2,3 CTD genes, encoding for catechol dioxygenase, bacterial iron-containing enzymes [183]. Some research indicates the involvement of Pseudomonas sp. dioxygenases enzymes in ortho (intradiol) cleavage and meta (extradiol) cleavage. Also, the catechol 1,2-dioxygenase (C12O) has been identified in Achromobacter xylosoxidans DN002, also involved in the ortho-cleavage pathway [184, 185]. The Pseudomonas stutzeri DJP1 strain degradation system consisted of PHE, seven metabolic intermediates, and PYR. The activity of key enzymes such as ring-cleaving dioxygenase, acetaldehyde dehydrogenase, and catechol 12 dioxygenase encoded by PhdF, NidD, and CatA genes, respectively, was recognized in cometabolic degradation and removal of high molecular weight polycyclic aromatic hydrocarbons (HMW-PAHs). Furthermore, Pseudomonas sp. plasmid encodes not only for catechol 1, 2-dioxygenase but also 3, 4-PCD with the same role, aromatic ring degradation [186]. Similar to lignin degradation, biphenyl-degrading bacteria can be useful and degrade xenobiotics. Located on MGE, horizontal gene transfer enables gene sharing and evolution, so these can be found in different biphenyl-degrading bacteria. Therefore, the Bph genes/degradation gene clusters for biphenyl/PCB are located on plasmids Geobacillus sp. strain JF8 pBt40, Rhodococcus sp. linear pRHL1, Cupriavidus sp. pSK-4, and Pseudomonas putida MB1335 [187]. Furthermore, biphenyl-degrading bacteria can be useful and degrade xenobiotics. The xenobiotics, PCBs with up to 10 chlorine atoms attached to two aromatic benzene rings, are highly hydrophobic and toxic compounds. Furthermore, Cupriavidus basilensis KF708 genome sequencing revealed 207 genes involved in the metabolism of aromatic compounds belonging to the bph gene cluster (bphEGFA1A2A3BCDA4) [188]. In general, bph genes encode dehydrogenases, namely bphA1A2A3A4, for the biphenyl dioxygenase, a multi-component enzyme. The bph genes beside the plasmid have been found on ICEs in C. oxalaticus A5 (formerly Ralstonia sp. strain A5) and Acidovorax sp. KKS102 [189]. Pseudomonas sp. strain shows adaptability, expressing biphenyl and salicylate metabolism gene clusters (the bph-sal element). Transfer of the bph-sal element in Pseudomonas putida KF715 enabled genome plasticity and genetic variability to withstand environmental challenges [190]. The role of biphenyl catabolic enzymes is to co-metabolize certain PCBs into chlorobenzoic acids and to substitute chlorine. Another group of PCB genes is salicylate catabolic sal genes. In 9 of 10 biphenyl/PCB bacterial strains, bph and sal gene clusters were transferred by integrative conjugative elements (ICEs), named ICEbph-sal, in Pseudomonas putida KF715. The SalR protein of Acinetobacter sp. strain ADP1 encoded by salA is a salicylate hydroxylase activator, while the same SalR protein, the Pseudomonas putida S-1 SalR protein encoded by sal genes, regulates salicylate degradation. The Sal gene encodes for LysR-Type Transcriptional Regulator family proteins - SalR is involved in converting alkyl salicylates into catechol [191]. Some cyanobacteria are involved in organic pollutant degradation while exhibiting similar mechanisms. Leptolyngbya sp. can be very effective in phenol degradation, while Spirulina CPCC-695 degrades estrone using it as a carbon source [192, 193]. Besides point mutations and mobile genetic elements, metagenomic tandem repeats have shown potential in engineered organisms. A metagenomic tandem repeat sequence of catechol 2,3-dioxygenase (C23O)-encoding genes in genetically modified E. coli showed increased C23O enzyme activity [194]. A catechol 2,3-dioxygenase (C23O)-encoding gene sequence plays a crucial role in crude oil degradation. The most degrading activity had EAU29438 Ewingella americana 16S rRNA gene and partial sequence (CFfab 14) and Bacillus megaterium strain IV22 16S ribosomal RNA gene, partial sequence (CFfab15) [195].
Benefits and Drawbacks of the Introduction of GMOs/GEMs in Bioremediation
The bioremediation technique is an old/new approach to restoring the harmed environment. New methods include the introduction of genetically modified microorganisms that advance pollution elimination without forming partially degraded intermediates, which are more toxic than the prime pollutants, which is the opposite of conventional procedures. Biological methods are given an advantage because chemical and physical remediation approaches are expensive and have adverse effects on the environment. Thus, there has been a rise in the use of genetically modified/ engineered microorganisms and biotechnology or genetic engineering. GMOs refer to all genetically changed living organisms. In contrast, the term “genetically engineered organisms” (GEMs) refers to microorganisms (bacteria, fungus, and yeast, among others) genetically modified by synthetic biology (removing and inserting genes) [38]. These designed organisms are tailored to have specific metabolic activities, the ability to overcome environmental stress, adjust to different abiotic and biotic factors, stronger structuring of genetic variation retained in the population, and the tracing of GEMs with molecular techniques. The enzyme 2,3-dihydroxy biphenyl 1,2-dioxygenase is encoded by genes bphC and etbC present in the Rhodococcus strain transferred from R. corallines to Rhodococcus TE1. Thus, the recombinant Rhodococcus achieved complete degradation of atrazine to cyanuric acid [196]. Manipulation with genetic material brings environmental concern.
In nature, trait variation within bacteria happens through gene transfer (conjugation, transformation, and transduction). Laboratory-engineered microorganisms can, with target genes, introduce unwanted genes into the environment (antibiotic resistance genes). However, the risk can be avoided by introducing technical safeguards and innovation and applying laws and regulations that are characteristic of every region. The plasmids as cloning vectors can be substituted with mini-transposons to clone and insert target sequences with non-antibiotic resistance genes [197]. The natural ability of bacteria’s toxin-antitoxin system to program their suicide under stress can also be activated after the depletion of pollutants. The interaction between microorganisms in environmental conditions is always uncertain; thus, field experiments must be conducted. Furthermore, remediation of the environmental damage caused by pollution and the potential threats of introducing GEMs can be leveled by a detailed risk assessment and constant monitoring after release. Also, the manipulation of genetic material must have boundaries by strict regulations, standards, and laws [198]. Thus, the introduction of rDNA into ecosystems is still restricted.
Other Aspects of Bioremediation without Using Biological Agents
The bioremediation technique that uses biological agents is not without any limitations. There are high molecular PAHs that cannot be broken down using a microorganism or incomplete degradation of pollutants, creating bigger treated, more toxic, and mobile intermediates. Also, unlike chemical and physical techniques, bioremediation takes a lot of time to remove toxins. Remediation with superoxidation products is based on known Fenton and Haber-Weiss reactions. The EU-funded HAC project targets the application of this advanced technique to clean water and soil from a wide range of contaminants such as PAHs, PBCs, CHCs, benzene, toluene, petroleum hydrocarbons, crude oil, and pesticides [199].
PAH degradation by chemical oxidation was shown to be very successful, namely modified Fenton’s reagent, hydrogen peroxide, and potassium permanganate degradation up to 95% light PAHs up to 1600 mg/kgSS, heavy PAHs up to 1200 mg/kgSS, residues were below 100 mg/kgSS. [200]. Some other studies compared benzo[a]pyrene (BaP) degradation by potassium permanganate (KMnO4), Fe2+ +sodium persulfate (Fe2+ + PS), Fenton’s reagent (Fe2+ + H2O2), and hydrogen peroxide (H2O2). The higher removal efficiency showed KMnO4 [201]. Oxidation with persulfate (PS) was less effective (up to 43%) than with permanganate (PM) (up to 98%), while treatment with modified Fenton’s reagent (F) showed the lowest degradation yields [202]. Stabilization/solidification (S/S) is considered the most appropriate method of converting contaminants into their least soluble/toxic. This is a physical and chemical reaction during which insoluble metal hydroxides are formed. Carbon binders are applied to chemically change and mechanically encapsulate the material. The calcined clay (CC) and limestone (LS) showed good Zn, As, and Pb immobilization results. Additionally, limestone-calcined clay cement (LC3) is also an effective Zn and Pb binder. Further, the reaction between LS and CC results in primary and secondary hydration products such as calcium silicate hydrate, calcium hydrate, calcium aluminate silicate hydrate, and calcium aluminate hydrate [203]. Geopolymer binders are considered eco-friendly and safer than conventional ones. Alkaline-activated silica-rich wastes, 3 aluminum silicate hydrates, showed binding capacity with 20% fly ash, 15% silica fume, and rice husk ash, all three with 6.5 molars of NaOH [204].
Advanced oxidation processes include chemical processes (Fenton’s process and Peroxone process), photochemical (photodecomposition (H2O2 ∕UV/ O3∕UV), photocatalysis (TiO2, ZnO), Photo- Fenton process (H2O2∕Fe2+∕UV), electrochemical (Electro- Fenton process) [205]. POP degradation using the TiO2/ UV photocatalytic system in wastewater was efficient. Namely, the TiO2/UV photocatalytic system effectively degraded pollutants with constant hydrogen production. In addition, toxic compounds from textiles, such as suspended solids, toxic chemicals, azo dyes, color, etc., can be degraded by adding strong oxidants such as hydroxyl radical (⋅OH) [206]. The introduction of novel materials that can photocatalytically degrade malachite green (MG) under the UV or VIS has promising results. Zinc oxide nanocomposites impregnated with nickel (Ni/ ZnO) were used in malachite green (MG) degradation. The highest removal efficiency of 99.4% was detected at pH 8 under VIS light [207].
Nanomaterials (nanomembranes, nanotubes, and nanoparticles) have wide use in drinking water purification, being able to remove toxic heavy metals such as Ni(II), Pb(II), Hg(II), Cu(II), and Zn(II) and POPs such as antibiotics, pesticides, and biological toxic components such as viruses, organic materials, bacteria, micronutrients, and algae [208]. Immobilization of pollutants can be achieved using carbon-based nanomaterials. Engineered nanomaterials carbon nanotube (CNT)-FexOy composites and (CFs) carbon nanotubes absorb and immobilize Cd and As. Also, Fe nanoparticles showed 88.34% successful Total Petroleum Hydrocarbons (TPHs) removal capacity from water. NM-based modifications act via reductive reactions, affecting the mobility of pollutants. Sodium carboxymethyl cellulose-stabilized nZVI could remove about 80% of Cr (VI) from soil. However, using these techniques has side effects, such as photocatalysis, the high level of mineralization, toxic postreaction products, photocorrosion, and light irradiation, which cause photocatalysts to become unstable; advanced oxidation processes (AOP) change pollutants, affect oxidants and catalysts, change configuration and form, and release toxic byproducts; the Fenton process applied at large treatments mishandles hydrogen peroxide [209]. Nitrogen and ammonium nitrogen in wastewater can be removed by algal and bacterial consortiums [210]. The problem of drinking water can be resolved through electro-bioremediation. Research funded by the European Union’s Horizon 2020 project ELECTRA within the EU Bioremediation cluster NYMPHE project showed high nitrate removal and chlorine production at the same time, eliminating pathogens such as T. coliforms, E. coli, and Enterococcus from drinking water. The project was conducted per the European Directive 2020/2184 [211]. Also, under this project, research conducted in a real environment showed that electro-bioremediation associated with Pseudomonadaceae, Rhizobiaceae, Gallionellaceae, and Xanthomonadaceae completely reduced NO3- to N2 [212]. Therefore, further research in this direction is needed.
Conclusions
Soil remediation methods may be applied in situ, on-site, and off-site. Within R&I, on-site methods in situ occur directly at the field, leaving the soil intact and clean. Therefore, on-site applied green techniques are within biological methods, bioremediation, and phytoremediation, or their combination. For instance, bacteria like Bacillus, Enterobacter, Escherichia, and Pseudomonas help remediate metal-contaminated soil.
Water pollution can be addressed by the algal-bacteria consortium that applies to the bioremediation of wastewater. This application is climate-friendly because the algae produce oxygen for the bacteria while using carbon dioxide in photosynthesis [213].
According to this review, the implementation of successful bioremediation requires a functional genomics approach. Being capable of adjusting to any environment, microorganisms’ tolerance detoxifies and removes the toxic metals. Mechanisms developed include different efflux transporter proteins. This review presents a variety of proteins, including P-type ATPases, RND (resistance, nodulation, cell division), CDF (cation diffusion facilitator), ATP binding cassette (ABC) transporters, MFS family proteins, and more, encoded by revealed and new genes and operons [214]. Therefore, many different biodegradation pathways have been discovered; however, new genes, their relationships, and their positions in the genome are receiving increasing attention. In addition to native genes, genetic engineering enabled various microorganisms to remove highly toxic substances from the environment. Numerous chemical, physical, and biological factors influence the course and success of bioremediation: pH, temperature, water content, nutrient availability, redox potential, and concentration of pollutants. Namely, many microorganisms synthesize enzymes only in the presence of a high concentration of toxic substances. Therefore, significance is in the novel gene sequence that will enable removal at low pollutant concentrations. Throughout this review, many native and engineered microorganisms showed a high ability to deal with both inorganic and organic pollution. Moreover, the same bacterial strain genome exhibits HMs and POPs removal potential. For instance, Pseudomonas sp., Cupriavidus sp., Burkholderia sp., and fungi Rhodococcus sp. were shown to be useful in the removal of a wide variety of pollutants. In general, microorganisms are handy tools for removing pollution, and offering insight into the mechanism of their action is essential for all researchers. Lab-scale biodegradation studies to field-scale trial insights can be conducted and evaluated by applying Molecular Biological Tools (MBTs) such as qPCR and 16S gene amplicon rRNA sequencing. Thus, the site abundance of functional genes can be detected. In enhanced bioremediation field trials, microbial consortia linked to PHC and TCE biodegradation include Dechloromonas, Deinococcus, Clostridium, Geobacillus, Syntrophus, Enterobacter, as well as Methanobacteria and Methanomicrobia that degrade HC to CH4 [215]. Furthermore, in situ bioremediation reduced TPHs in soil by about 88%, introducing Bacillus sp., Pseudomonas sp., Arthrobacter sp., Acinetobacter sp., Alcaligenes sp., and Brevibacillus sp. Some other strains, Acinetobacter baumannii (S19, S26, S30), Burkholderia cepacia (P20), and Pseudomonas sp. (S24), also showed the in situ potential of 90% in TPH removal. Further, Bacillus, B. thuringiensis B3, and B. cereus B6 reduced diesel and crude oil from crude oil-polluted fields between 28% and 84% [216]. In field trials for removing heavy metals from water and soil, Bacillus sp. and Pseudomonas sp. applications have shown promising results. Furthermore, Geobacteraceae species were found to be the most abundant in reducing Fe3+. Intrinsic bioremediation, bioaugmentation, and engineered bioremediation help restore polluted soil. However, from lab to field applications, there need to be considerations for monitoring after the release of GEMs and factors such as environmental challenges, including temperature, humidity, pH, rather than the competition and survival of microorganisms [216, 217]. The findings of these studies imply that microorganisms are suitable candidates for the bio-removal of a wide range of inorganic and organic pollutants, as shown in Table 1. and Table 2. Research results are vital in implementing preventive and protective actions. The implication of this research is to identify genetic determinants of prime bioremediation as an advantage in target pollutant removal and to direct researchers toward genetically modified organisms of interest with respect to potential drawbacks. Therefore, this research is relevant for researchers and stakeholders participating in living labs and lighthouses as open innovation platforms for the transdisciplinary conservation, environmental, and agricultural sustainability fields.
Further Perspectives
The greatest worry in recent years is human-induced climate change. As a consequence, contaminant release, pathways, and bioavailability in terrestrial and aquatic environments are changed [218]. To tackle this issue, future bioremediation focus must be shifted to genetic microbial community modeling, pollutant-microbe interaction modeling, and site-specific pollution forecasting. The biggest problem in successful bioremediation is environmental changes that occur faster than wanted. Therefore, predictive ecology must tailor effective microbial consortia.
Future research should focus on forecasting models based on improved genomic techniques and developing precision-designed synthetic microbial communities that won’t be affected by environmental factors. Consequently, climate reshapes the landscape and influences environmental factors and biocenosis; therefore, advancing genetic encoding of pollutant pathways via DNA nanotechnology can give precise pollutant-microbe interaction patterns in changed ecosystems.
The limiting factor is the concentration of the contaminants in the environment. Toxins in high and low concentrations directly affect microbial activity. Thus, lower pollution concentration decreases degrading enzyme production by microorganisms. Therefore, the decomposition rate of catabolic enzymes can be increased by genetic engineering methods, increasing the yield of enzyme production. For instance, organophosphate degrading enzymes were overexpressed in recombinant E. coli BL21, increasing the pesticide degrading rate compared with non-recombinant enzymes [219]. The aspects of assisted evolution also should be considered as a new direction.

